return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CSNH2 (Ethanethioamide)

using model chemistry: CCD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/6-31G(2df,p)
 hartrees
Energy at 0K-531.344539
Energy at 298.15K 
HF Energy-530.635467
Nuclear repulsion energy155.406099
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3800 3599 53.84      
2 A 3663 3470 97.09      
3 A 3239 3068 0.87      
4 A 3164 2997 6.13      
5 A 3093 2930 10.33      
6 A 1674 1585 193.57      
7 A 1519 1439 8.47      
8 A 1518 1438 26.53      
9 A 1445 1369 155.99      
10 A 1425 1350 200.11      
11 A 1364 1292 30.92      
12 A 1056 1001 0.03      
13 A 1044 988 36.50      
14 A 1021 967 9.83      
15 A 755 716 9.97      
16 A 635 601 8.00      
17 A 529 501 2.79      
18 A 434 411 1.01      
19 A 383 363 2.81      
20 A 314 297 177.72      
21 A 48 45 0.30      

Unscaled Zero Point Vibrational Energy (zpe) 16061.0 cm-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 15212.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G(2df,p)
ABC
0.32718 0.16587 0.11238

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.272 0.061 0.000
S2 -1.363 -0.118 0.000
C3 1.238 -1.101 0.000
N4 0.872 1.262 -0.001
H5 0.695 -2.042 0.007
H6 1.875 -1.058 -0.888
H7 1.885 -1.051 0.880
H8 1.872 1.353 0.003
H9 0.309 2.093 0.001

Atom - Atom Distances (Å)
  C1 S2 C3 N4 H5 H6 H7 H8 H9
C11.64491.51121.34212.14502.14702.14762.05592.0321
S21.64492.78002.62682.81633.48603.49163.55342.7719
C31.51122.78002.39141.08571.09371.09372.53543.3269
N41.34212.62682.39143.30832.67872.67451.00351.0043
H52.14502.81631.08573.30831.77751.77773.59334.1527
H62.14703.48601.09372.67871.77751.76852.57133.6297
H72.14763.49161.09372.67451.77771.76852.55973.6255
H82.05593.55342.53541.00353.59332.57132.55971.7291
H92.03212.77193.32691.00434.15273.62973.62551.7291

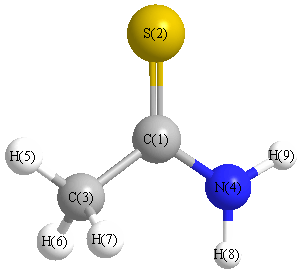
picture of Ethanethioamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H5 110.293 C1 C3 H6 109.983
C1 C3 H7 110.031 C1 N4 H8 121.780
C1 N4 H9 119.308 S2 C1 C3 123.426
S2 C1 N4 122.820 C3 C1 N4 113.754
H5 C3 H6 109.291 H5 C3 H7 109.303
H6 C3 H7 107.897 H8 N4 H9 118.910
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability