Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -553.223373 |
| Energy at 298.15K | -553.228586 |
| Nuclear repulsion energy | 199.307987 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3319 |
3135 |
1.56 |
|
|
|
| 2 |
A |
3238 |
3058 |
2.08 |
|
|
|
| 3 |
A |
3214 |
3036 |
0.30 |
|
|
|
| 4 |
A |
1655 |
1563 |
3.27 |
|
|
|
| 5 |
A |
1433 |
1353 |
8.11 |
|
|
|
| 6 |
A |
1308 |
1235 |
0.95 |
|
|
|
| 7 |
A |
1064 |
1005 |
0.29 |
|
|
|
| 8 |
A |
1014 |
957 |
4.76 |
|
|
|
| 9 |
A |
926 |
875 |
14.08 |
|
|
|
| 10 |
A |
752 |
711 |
0.03 |
|
|
|
| 11 |
A |
635 |
600 |
8.08 |
|
|
|
| 12 |
A |
386 |
364 |
0.27 |
|
|
|
| 13 |
A |
184 |
174 |
0.00 |
|
|
|
| 14 |
A |
67 |
63 |
0.12 |
|
|
|
| 15 |
A |
3319 |
3135 |
1.34 |
|
|
|
| 16 |
A |
3235 |
3055 |
8.71 |
|
|
|
| 17 |
A |
3214 |
3036 |
0.95 |
|
|
|
| 18 |
A |
1639 |
1548 |
55.25 |
|
|
|
| 19 |
A |
1427 |
1348 |
3.42 |
|
|
|
| 20 |
A |
1294 |
1222 |
17.25 |
|
|
|
| 21 |
A |
1044 |
986 |
8.26 |
|
|
|
| 22 |
A |
1004 |
948 |
62.36 |
|
|
|
| 23 |
A |
926 |
874 |
40.45 |
|
|
|
| 24 |
A |
756 |
714 |
17.09 |
|
|
|
| 25 |
A |
604 |
571 |
13.45 |
|
|
|
| 26 |
A |
393 |
371 |
1.73 |
|
|
|
| 27 |
A |
64 |
60 |
2.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19055.3 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 17997.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C2
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -553.225518 |
| Energy at 298.15K | -553.231050 |
| HF Energy | -552.444125 |
| Nuclear repulsion energy | 202.954138 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3322 |
3138 |
1.72 |
|
|
|
| 2 |
A |
3318 |
3134 |
1.23 |
|
|
|
| 3 |
A |
3242 |
3062 |
3.28 |
|
|
|
| 4 |
A |
3229 |
3050 |
3.67 |
|
|
|
| 5 |
A |
3219 |
3040 |
0.45 |
|
|
|
| 6 |
A |
3212 |
3033 |
0.23 |
|
|
|
| 7 |
A |
1650 |
1559 |
23.78 |
|
|
|
| 8 |
A |
1640 |
1549 |
19.18 |
|
|
|
| 9 |
A |
1434 |
1354 |
4.24 |
|
|
|
| 10 |
A |
1430 |
1351 |
5.43 |
|
|
|
| 11 |
A |
1309 |
1236 |
3.16 |
|
|
|
| 12 |
A |
1296 |
1225 |
10.02 |
|
|
|
| 13 |
A |
1067 |
1008 |
4.55 |
|
|
|
| 14 |
A |
1039 |
981 |
16.93 |
|
|
|
| 15 |
A |
1020 |
963 |
21.76 |
|
|
|
| 16 |
A |
1004 |
948 |
25.59 |
|
|
|
| 17 |
A |
942 |
890 |
26.22 |
|
|
|
| 18 |
A |
902 |
852 |
31.04 |
|
|
|
| 19 |
A |
765 |
723 |
8.73 |
|
|
|
| 20 |
A |
718 |
678 |
4.65 |
|
|
|
| 21 |
A |
643 |
607 |
13.14 |
|
|
|
| 22 |
A |
618 |
583 |
9.80 |
|
|
|
| 23 |
A |
460 |
434 |
0.33 |
|
|
|
| 24 |
A |
370 |
349 |
0.53 |
|
|
|
| 25 |
A |
238 |
225 |
1.25 |
|
|
|
| 26 |
A |
153 |
145 |
4.27 |
|
|
|
| 27 |
A |
88 |
84 |
1.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19163.3 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 18099.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.009 |
-0.990 |
0.001 |
| C2 |
1.061 |
0.362 |
0.384 |
| C3 |
-1.552 |
-0.190 |
-0.030 |
| C4 |
2.234 |
0.558 |
-0.225 |
| C5 |
-1.776 |
1.126 |
-0.127 |
| H6 |
0.728 |
0.986 |
1.208 |
| H7 |
-2.374 |
-0.894 |
0.049 |
| H8 |
2.904 |
1.339 |
0.112 |
| H9 |
2.553 |
-0.061 |
-1.054 |
| H10 |
-0.972 |
1.841 |
-0.237 |
| H11 |
-2.789 |
1.504 |
-0.109 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7552 | 1.7551 | 2.7200 | 2.7715 | 2.4248 | 2.3854 | 3.7170 | 2.9065 | 3.0061 | 3.7504 |
C2 | 1.7552 | | 2.7025 | 1.3368 | 2.9819 | 1.0856 | 3.6721 | 2.1039 | 2.1153 | 2.5897 | 4.0461 | C3 | 1.7551 | 2.7025 | | 3.8647 | 1.3380 | 2.8489 | 1.0846 | 4.7134 | 4.2334 | 2.1224 | 2.0988 | C4 | 2.7200 | 1.3368 | 3.8647 | | 4.0513 | 2.1224 | 4.8389 | 1.0827 | 1.0823 | 3.4535 | 5.1132 | C5 | 2.7715 | 2.9819 | 1.3380 | 4.0513 | | 2.8416 | 2.1135 | 4.6908 | 4.5834 | 1.0817 | 1.0819 | H6 | 2.4248 | 1.0856 | 2.8489 | 2.1224 | 2.8416 | | 3.8080 | 2.4614 | 3.0889 | 2.3899 | 3.7919 | H7 | 2.3854 | 3.6721 | 1.0846 | 4.8389 | 2.1135 | 3.8080 | | 5.7308 | 5.1169 | 3.0865 | 2.4385 | H8 | 3.7170 | 2.1039 | 4.7134 | 1.0827 | 4.6908 | 2.4614 | 5.7308 | | 1.8552 | 3.9240 | 5.7000 | H9 | 2.9065 | 2.1153 | 4.2334 | 1.0823 | 4.5834 | 3.0889 | 5.1169 | 1.8552 | | 4.0880 | 5.6466 | H10 | 3.0061 | 2.5897 | 2.1224 | 3.4535 | 1.0817 | 2.3899 | 3.0865 | 3.9240 | 4.0880 | | 1.8528 | H11 | 3.7504 | 4.0461 | 2.0988 | 5.1132 | 1.0819 | 3.7919 | 2.4385 | 5.7000 | 5.6466 | 1.8528 | |
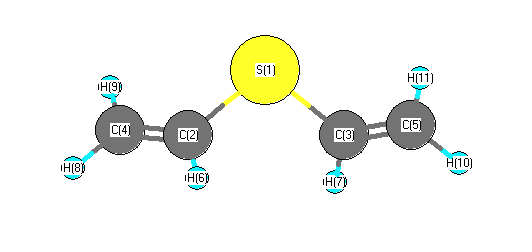 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.634 |
|
S1 |
C2 |
H6 |
115.171 |
| S1 |
C3 |
C5 |
126.751 |
|
S1 |
C3 |
H7 |
112.117 |
| C2 |
S1 |
C3 |
100.688 |
|
C2 |
C4 |
H8 |
120.453 |
| C2 |
C4 |
H9 |
121.595 |
|
C3 |
C5 |
H10 |
122.243 |
| C3 |
C5 |
H11 |
119.925 |
|
C4 |
C2 |
H6 |
122.025 |
| C5 |
C3 |
H7 |
121.131 |
|
H8 |
C4 |
H9 |
117.945 |
| H10 |
C5 |
H11 |
117.828 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
