return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H7NO (Ethoxyacetonitrile)

using model chemistry: QCISD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G(2df,p)
 hartrees
Energy at 0K-285.936117
Energy at 298.15K-285.943955
HF Energy-284.861353
Nuclear repulsion energy216.502053
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3180 3013 15.95      
2 A' 3096 2934 10.03      
3 A' 3043 2883 34.93      
4 A' 3030 2871 23.91      
5 A' 2364 2240 0.77      
6 A' 1566 1484 3.35      
7 A' 1536 1455 2.65      
8 A' 1531 1450 3.92      
9 A' 1482 1404 7.56      
10 A' 1438 1362 20.98      
11 A' 1410 1336 59.92      
12 A' 1209 1145 184.25      
13 A' 1176 1114 18.79      
14 A' 1082 1025 14.01      
15 A' 967 916 9.93      
16 A' 921 872 2.70      
17 A' 559 530 1.43      
18 A' 429 406 1.04      
19 A' 306 290 2.60      
20 A' 135 128 3.49      
21 A" 3186 3019 16.15      
22 A" 3076 2915 8.30      
23 A" 3067 2906 51.73      
24 A" 1514 1435 4.73      
25 A" 1325 1255 3.11      
26 A" 1287 1219 2.96      
27 A" 1212 1148 5.82      
28 A" 1057 1001 1.28      
29 A" 834 790 0.48      
30 A" 377 357 2.56      
31 A" 252 239 0.35      
32 A" 114 108 6.01      
33 A" 78 73 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 23918.3 cm-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 22660.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G(2df,p)
ABC
0.51820 0.05022 0.04699

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.948 2.529 0.000
C2 -1.242 1.042 0.000
O3 0.000 0.358 0.000
C4 -0.183 -1.029 0.000
C5 1.139 -1.677 0.000
N6 2.163 -2.219 0.000
H7 -1.879 3.099 0.000
H8 -0.370 2.800 0.886
H9 -0.370 2.800 -0.886
H10 -1.826 0.757 -0.887
H11 -1.826 0.757 0.887
H12 -0.737 -1.368 -0.887
H13 -0.737 -1.368 0.887

Atom - Atom Distances (Å)
  C1 C2 O3 C4 C5 N6 H7 H8 H9 H10 H11 H12 H13
C11.51612.36873.63914.69505.67651.09221.09141.09142.16762.16764.00224.0022
C21.51611.41742.32543.61374.71452.15382.15292.15291.09951.09952.61692.6169
O32.36871.41741.39912.33203.36483.32332.62392.62392.06852.06852.07572.0757
C43.63912.32541.39911.47272.63124.46273.93453.93452.58312.58311.09901.0990
C54.69503.61372.33201.47271.15885.64974.80674.80673.93693.93692.09802.0980
N65.67654.71453.36482.63121.15886.68025.69175.69175.05515.05513.14983.1498
H71.09222.15383.32334.46275.64976.68021.77501.77502.50512.50514.69534.6953
H81.09142.15292.62393.93454.80675.69171.77501.77163.07162.50854.54414.1841
H91.09142.15292.62393.93454.80675.69171.77501.77162.50853.07164.18414.5441
H102.16761.09952.06852.58313.93695.05512.50513.07162.50851.77372.38752.9743
H112.16761.09952.06852.58313.93695.05512.50512.50853.07161.77372.97432.3875
H124.00222.61692.07571.09902.09803.14984.69534.54414.18412.38752.97431.7736
H134.00222.61692.07571.09902.09803.14984.69534.18414.54412.97432.38751.7736

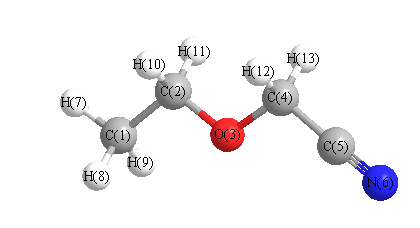
picture of Ethoxyacetonitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 107.655 C1 C2 H10 110.934
C1 C2 H11 110.934 C2 C1 H7 110.272
C2 C1 H8 110.249 C2 C1 H9 110.249
C2 O3 C4 111.309 O3 C2 H10 109.897
O3 C2 H11 109.897 O3 C4 C5 108.565
O3 C4 H12 111.833 O3 C4 H13 111.833
C4 C5 N6 178.183 C5 C4 H12 108.456
C5 C4 H13 108.456 H7 C1 H8 108.759
H7 C1 H9 108.759 H8 C1 H9 108.505
H10 C2 H11 107.533 H12 C4 H13 107.593
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability