Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -749.953842 |
| Energy at 298.15K | |
| HF Energy | -748.037656 |
| Nuclear repulsion energy | 564.622174 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1863 |
1759 |
0.00 |
|
|
|
| 2 |
Ag |
1471 |
1389 |
0.00 |
|
|
|
| 3 |
Ag |
1382 |
1305 |
0.00 |
|
|
|
| 4 |
Ag |
1221 |
1154 |
0.00 |
|
|
|
| 5 |
Ag |
727 |
687 |
0.00 |
|
|
|
| 6 |
Ag |
596 |
563 |
0.00 |
|
|
|
| 7 |
Ag |
383 |
362 |
0.00 |
|
|
|
| 8 |
Ag |
353 |
333 |
0.00 |
|
|
|
| 9 |
Ag |
222 |
210 |
0.00 |
|
|
|
| 10 |
Au |
557 |
526 |
3.01 |
|
|
|
| 11 |
Au |
349 |
329 |
3.40 |
|
|
|
| 12 |
Au |
110 |
104 |
0.05 |
|
|
|
| 13 |
Au |
17i |
16i |
0.09 |
|
|
|
| 14 |
Bg |
614 |
580 |
0.00 |
|
|
|
| 15 |
Bg |
484 |
457 |
0.00 |
|
|
|
| 16 |
Bg |
184 |
174 |
0.00 |
|
|
|
| 17 |
Bu |
1803 |
1703 |
326.37 |
|
|
|
| 18 |
Bu |
1399 |
1322 |
374.87 |
|
|
|
| 19 |
Bu |
1259 |
1189 |
249.77 |
|
|
|
| 20 |
Bu |
979 |
924 |
228.96 |
|
|
|
| 21 |
Bu |
629 |
594 |
5.68 |
|
|
|
| 22 |
Bu |
503 |
475 |
3.25 |
|
|
|
| 23 |
Bu |
284 |
268 |
5.16 |
|
|
|
| 24 |
Bu |
139 |
131 |
0.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8746.2 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 8260.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.156 |
1.849 |
0.000 |
| C2 |
0.471 |
0.549 |
0.000 |
| C3 |
-0.471 |
-0.549 |
0.000 |
| C4 |
-0.156 |
-1.849 |
0.000 |
| F5 |
1.061 |
2.794 |
0.000 |
| F6 |
-1.061 |
2.322 |
0.000 |
| F7 |
1.765 |
0.218 |
0.000 |
| F8 |
-1.765 |
-0.218 |
0.000 |
| F9 |
1.061 |
-2.322 |
0.000 |
| F10 |
-1.061 |
-2.794 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3377 | 2.4793 | 3.7118 | 1.3079 | 1.3058 | 2.2909 | 2.8228 | 4.2685 | 4.8003 |
C2 | 1.3377 | | 1.4476 | 2.4793 | 2.3210 | 2.3434 | 1.3355 | 2.3648 | 2.9313 | 3.6777 | C3 | 2.4793 | 1.4476 | | 1.3377 | 3.6777 | 2.9313 | 2.3648 | 1.3355 | 2.3434 | 2.3210 | C4 | 3.7118 | 2.4793 | 1.3377 | | 4.8003 | 4.2685 | 2.8228 | 2.2909 | 1.3058 | 1.3079 | F5 | 1.3079 | 2.3210 | 3.6777 | 4.8003 | | 2.1734 | 2.6703 | 4.1306 | 5.1164 | 5.9774 | F6 | 1.3058 | 2.3434 | 2.9313 | 4.2685 | 2.1734 | | 3.5232 | 2.6366 | 5.1062 | 5.1164 | F7 | 2.2909 | 1.3355 | 2.3648 | 2.8228 | 2.6703 | 3.5232 | | 3.5575 | 2.6366 | 4.1306 | F8 | 2.8228 | 2.3648 | 1.3355 | 2.2909 | 4.1306 | 2.6366 | 3.5575 | | 3.5232 | 2.6703 | F9 | 4.2685 | 2.9313 | 2.3434 | 1.3058 | 5.1164 | 5.1062 | 2.6366 | 3.5232 | | 2.1734 | F10 | 4.8003 | 3.6777 | 2.3210 | 1.3079 | 5.9774 | 5.1164 | 4.1306 | 2.6703 | 2.1734 | |
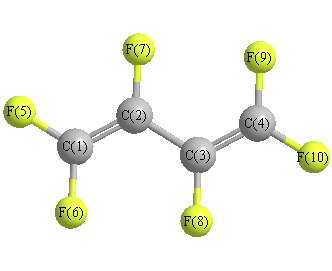 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.737 |
|
C1 |
C2 |
F7 |
117.960 |
| C2 |
C1 |
F5 |
122.630 |
|
C2 |
C1 |
F6 |
124.858 |
| C2 |
C3 |
C4 |
125.737 |
|
C2 |
C3 |
F8 |
116.303 |
| C3 |
C2 |
F7 |
116.303 |
|
C3 |
C4 |
F9 |
124.858 |
| C3 |
C4 |
F10 |
122.630 |
|
C4 |
C3 |
F8 |
117.960 |
| F5 |
C1 |
F6 |
112.512 |
|
F9 |
C4 |
F10 |
112.512 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -749.955497 |
| Energy at 298.15K | -749.956717 |
| HF Energy | -748.038665 |
| Nuclear repulsion energy | 570.177046 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
1852 |
1750 |
57.10 |
|
|
|
| 2 |
A |
1444 |
1364 |
22.68 |
|
|
|
| 3 |
A |
1400 |
1323 |
157.87 |
|
|
|
| 4 |
A |
1182 |
1116 |
237.30 |
|
|
|
| 5 |
A |
726 |
685 |
0.25 |
|
|
|
| 6 |
A |
649 |
613 |
1.77 |
|
|
|
| 7 |
A |
544 |
514 |
0.01 |
|
|
|
| 8 |
A |
481 |
454 |
0.14 |
|
|
|
| 9 |
A |
384 |
363 |
0.78 |
|
|
|
| 10 |
A |
251 |
237 |
0.28 |
|
|
|
| 11 |
A |
194 |
183 |
0.32 |
|
|
|
| 12 |
A |
100 |
95 |
0.14 |
|
|
|
| 13 |
A |
63 |
59 |
0.00 |
|
|
|
| 14 |
B |
1829 |
1728 |
220.80 |
|
|
|
| 15 |
B |
1392 |
1315 |
254.17 |
|
|
|
| 16 |
B |
1252 |
1182 |
80.92 |
|
|
|
| 17 |
B |
1002 |
946 |
148.32 |
|
|
|
| 18 |
B |
633 |
598 |
5.56 |
|
|
|
| 19 |
B |
614 |
580 |
2.17 |
|
|
|
| 20 |
B |
571 |
539 |
3.40 |
|
|
|
| 21 |
B |
417 |
394 |
3.39 |
|
|
|
| 22 |
B |
298 |
281 |
3.64 |
|
|
|
| 23 |
B |
208 |
197 |
3.29 |
|
|
|
| 24 |
B |
108 |
102 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8797.3 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 8309.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.085 |
1.538 |
-0.379 |
| C2 |
0.256 |
0.676 |
0.625 |
| C3 |
-0.256 |
-0.676 |
0.625 |
| C4 |
-0.085 |
-1.538 |
-0.379 |
| F5 |
0.598 |
2.736 |
-0.411 |
| F6 |
-0.598 |
1.256 |
-1.458 |
| F7 |
0.944 |
1.066 |
1.704 |
| F8 |
-0.944 |
-1.066 |
1.704 |
| F9 |
0.598 |
-1.256 |
-1.458 |
| F10 |
-0.598 |
-2.736 |
-0.411 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3344 | 2.4542 | 3.0798 | 1.3042 | 1.3078 | 2.3021 | 3.4898 | 3.0385 | 4.3281 |
C2 | 1.3344 | | 1.4450 | 2.4542 | 2.3317 | 2.3252 | 1.3380 | 2.3744 | 2.8618 | 3.6666 | C3 | 2.4542 | 1.4450 | | 1.3344 | 3.6666 | 2.8618 | 2.3744 | 1.3380 | 2.3252 | 2.3317 | C4 | 3.0798 | 2.4542 | 1.3344 | | 4.3281 | 3.0385 | 3.4898 | 2.3021 | 1.3078 | 1.3042 | F5 | 1.3042 | 2.3317 | 3.6666 | 4.3281 | | 2.1719 | 2.7172 | 4.6163 | 4.1275 | 5.6016 | F6 | 1.3078 | 2.3252 | 2.8618 | 3.0385 | 2.1719 | | 3.5234 | 3.9386 | 2.7828 | 4.1275 | F7 | 2.3021 | 1.3380 | 2.3744 | 3.4898 | 2.7172 | 3.5234 | | 2.8483 | 3.9386 | 4.6163 | F8 | 3.4898 | 2.3744 | 1.3380 | 2.3021 | 4.6163 | 3.9386 | 2.8483 | | 3.5234 | 2.7172 | F9 | 3.0385 | 2.8618 | 2.3252 | 1.3078 | 4.1275 | 2.7828 | 3.9386 | 3.5234 | | 2.1719 | F10 | 4.3281 | 3.6666 | 2.3317 | 1.3042 | 5.6016 | 4.1275 | 4.6163 | 2.7172 | 2.1719 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.970 |
|
C1 |
C2 |
F7 |
118.958 |
| C2 |
C1 |
F5 |
124.184 |
|
C2 |
C1 |
F6 |
123.291 |
| C2 |
C3 |
C4 |
123.970 |
|
C2 |
C3 |
F8 |
117.071 |
| C3 |
C2 |
F7 |
117.071 |
|
C3 |
C4 |
F9 |
123.291 |
| C3 |
C4 |
F10 |
124.184 |
|
C4 |
C3 |
F8 |
118.958 |
| F5 |
C1 |
F6 |
112.510 |
|
F9 |
C4 |
F10 |
112.510 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
