return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,4-Hexadiene, (E)-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-233.957417
Energy at 298.15K-233.967078
HF Energy-233.014134
Nuclear repulsion energy215.278696
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3313 3129 7.30      
2 A 3220 3041 7.47      
3 A 3207 3029 27.06      
4 A 3205 3027 13.15      
5 A 3198 3021 4.45      
6 A 3192 3015 2.83      
7 A 3171 2995 11.56      
8 A 3123 2949 9.89      
9 A 3094 2923 19.23      
10 A 3076 2905 18.14      
11 A 1738 1642 0.84      
12 A 1703 1608 8.67      
13 A 1523 1439 8.30      
14 A 1510 1427 5.25      
15 A 1493 1410 8.25      
16 A 1455 1374 3.62      
17 A 1431 1352 2.57      
18 A 1376 1300 0.63      
19 A 1342 1268 1.33      
20 A 1332 1258 3.80      
21 A 1323 1249 0.99      
22 A 1233 1164 1.06      
23 A 1145 1082 3.67      
24 A 1128 1065 4.13      
25 A 1075 1016 0.85      
26 A 1060 1001 7.75      
27 A 1049 991 7.10      
28 A 1026 969 23.83      
29 A 969 915 9.38      
30 A 950 897 23.78      
31 A 940 888 1.89      
32 A 898 848 1.93      
33 A 787 743 0.23      
34 A 603 570 4.74      
35 A 547 516 5.07      
36 A 468 442 0.21      
37 A 323 305 0.37      
38 A 283 267 1.26      
39 A 212 201 0.97      
40 A 158 149 0.49      
41 A 134 126 0.06      
42 A 75 71 0.15      

Unscaled Zero Point Vibrational Energy (zpe) 31543.6 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 29792.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.27264 0.05853 0.05134

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.729 0.462 0.033
H2 2.950 1.291 -0.644
H3 2.661 0.859 1.047
H4 3.583 -0.218 -0.010
C5 1.463 -0.234 -0.360
H6 1.430 -0.663 -1.360
C7 0.390 -0.364 0.427
H8 0.411 0.070 1.425
C9 -0.876 -1.058 0.031
H10 -0.737 -1.568 -0.929
H11 -1.108 -1.847 0.757
C12 -2.067 -0.138 -0.063
H13 -3.036 -0.623 -0.143
C14 -2.005 1.196 -0.067
H15 -2.900 1.798 -0.146
H16 -1.057 1.716 0.001

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 C7 H8 C9 H10 H11 C12 H13 C14 H15 H16
C11.09301.09081.09301.49762.21222.51142.73163.91224.13024.53574.83375.86844.79115.78813.9884
H21.09301.76921.75492.14922.57703.23103.49544.54004.67415.31725.24846.30434.98935.89354.0814
H31.09081.76921.76892.14773.10252.65312.41424.14964.61994.64884.95756.00564.80885.76483.9566
H41.09301.75491.76892.14932.57983.22633.49344.53774.61855.02445.65066.63255.76436.79115.0274
C51.49762.14922.14772.14931.08801.33682.09382.51012.63453.23333.54304.52033.76214.81783.2067
H62.21222.57703.10252.57981.08802.08863.05402.72152.38713.50973.76604.62854.11385.12663.7010
C72.51143.23102.65313.22631.33682.08861.08811.49702.13512.13332.51513.48242.90023.97842.5691
H82.73163.49542.41423.49342.09383.05401.08812.20693.08882.53502.89733.84943.05434.05202.6251
C93.91224.54004.14964.53772.51012.72151.49702.20691.09611.09671.50772.20992.52243.50522.7796
H104.13024.67414.61994.61852.63452.38712.13513.08881.09611.74882.13612.60653.16014.07693.4274
H114.53575.31724.64885.02443.23333.50972.13332.53501.09671.74882.12442.45453.27734.16113.6420
C124.83375.24844.95755.65063.54303.76602.51512.89731.50772.13612.12441.08691.33482.10942.1111
H135.86846.30436.00566.63254.52034.62853.48243.84942.20992.60652.45451.08692.09212.42513.0666
C144.79114.98934.80885.76433.76214.11382.90023.05432.52243.16013.27731.33482.09211.08221.0828
H155.78815.89355.76486.79114.81785.12663.97844.05203.50524.07694.16112.10942.42511.08221.8505
H163.98844.08143.95665.02743.20673.70102.56912.62512.77963.42743.64202.11113.06661.08281.8505

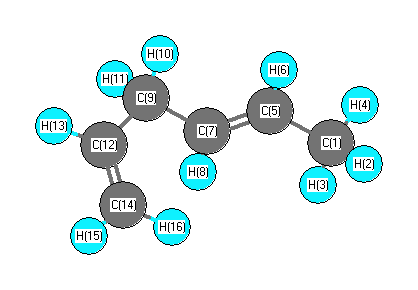
picture of 1,4-Hexadiene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 116.755 C1 C5 C7 124.664
H2 C1 H3 108.221 H2 C1 H4 106.788
H2 C1 C5 111.153 H3 C1 H4 108.184
H3 C1 C5 111.165 H4 C1 C5 111.158
C5 C7 H8 119.058 C5 C7 C9 124.601
H6 C5 C7 118.581 C7 C9 H10 109.882
C7 C9 H11 109.704 C7 C9 C12 113.658
H8 C7 C9 116.335 C9 C12 H13 115.853
C9 C12 C14 124.983 H10 C9 H11 105.789
H10 C9 C12 109.221 H11 C9 C12 108.276
C12 C14 H15 121.203 C12 C14 H16 121.328
H13 C12 C14 119.160 H15 C14 H16 117.468
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability