Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -79.583902 |
| Energy at 298.15K | -79.589836 |
| HF Energy | -79.241645 |
| Nuclear repulsion energy | 42.283735 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3112 |
2940 |
0.00 |
186.98 |
0.01 |
0.02 |
| 2 |
A1g |
1448 |
1368 |
0.00 |
3.96 |
0.75 |
0.86 |
| 3 |
A1g |
1035 |
978 |
0.00 |
9.68 |
0.25 |
0.41 |
| 4 |
A1u |
320 |
303 |
0.00 |
0.00 |
0.00 |
0.00 |
| 5 |
A2u |
3115 |
2942 |
37.50 |
0.00 |
0.01 |
0.02 |
| 6 |
A2u |
1427 |
1348 |
1.09 |
0.00 |
0.00 |
0.00 |
| 7 |
Eg |
3193 |
3016 |
0.00 |
109.56 |
0.75 |
0.86 |
| 7 |
Eg |
3193 |
3016 |
0.00 |
109.56 |
0.75 |
0.86 |
| 8 |
Eg |
1538 |
1453 |
0.00 |
28.75 |
0.75 |
0.86 |
| 8 |
Eg |
1538 |
1453 |
0.00 |
28.75 |
0.75 |
0.86 |
| 9 |
Eg |
1239 |
1170 |
0.00 |
2.01 |
0.75 |
0.86 |
| 9 |
Eg |
1239 |
1170 |
0.00 |
2.01 |
0.75 |
0.86 |
| 10 |
Eu |
3214 |
3036 |
37.85 |
0.00 |
0.00 |
0.00 |
| 10 |
Eu |
3214 |
3036 |
37.85 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1540 |
1455 |
7.42 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1540 |
1455 |
7.42 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
832 |
785 |
4.63 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
832 |
785 |
4.63 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 16784.5 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 15853.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
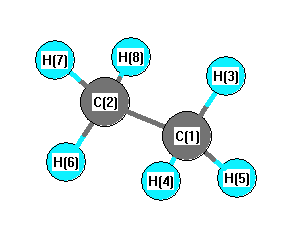
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.763 |
| C2 |
0.000 |
0.000 |
-0.763 |
| H3 |
0.000 |
1.017 |
1.158 |
| H4 |
-0.880 |
-0.508 |
1.158 |
| H5 |
0.880 |
-0.508 |
1.158 |
| H6 |
0.000 |
-1.017 |
-1.158 |
| H7 |
-0.880 |
0.508 |
-1.158 |
| H8 |
0.880 |
0.508 |
-1.158 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5264 | 1.0907 | 1.0907 | 1.0907 | 2.1738 | 2.1738 | 2.1738 |
C2 | 1.5264 | | 2.1738 | 2.1738 | 2.1738 | 1.0907 | 1.0907 | 1.0907 | H3 | 1.0907 | 2.1738 | | 1.7609 | 1.7609 | 3.0822 | 2.5297 | 2.5297 | H4 | 1.0907 | 2.1738 | 1.7609 | | 1.7609 | 2.5297 | 2.5297 | 3.0822 | H5 | 1.0907 | 2.1738 | 1.7609 | 1.7609 | | 2.5297 | 3.0822 | 2.5297 | H6 | 2.1738 | 1.0907 | 3.0822 | 2.5297 | 2.5297 | | 1.7609 | 1.7609 | H7 | 2.1738 | 1.0907 | 2.5297 | 2.5297 | 3.0822 | 1.7609 | | 1.7609 | H8 | 2.1738 | 1.0907 | 2.5297 | 3.0822 | 2.5297 | 1.7609 | 1.7609 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.234 |
|
C1 |
C2 |
H7 |
111.234 |
| C1 |
C2 |
H8 |
111.234 |
|
C2 |
C1 |
H3 |
111.234 |
| C2 |
C1 |
H4 |
111.234 |
|
C2 |
C1 |
H5 |
111.234 |
| H3 |
C1 |
H4 |
107.652 |
|
H3 |
C1 |
H5 |
107.652 |
| H4 |
C1 |
H5 |
107.652 |
|
H6 |
C2 |
H7 |
107.652 |
| H6 |
C2 |
H8 |
107.652 |
|
H7 |
C2 |
H8 |
107.652 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -79.579243 |
| Energy at 298.15K | |
| HF Energy | -79.236734 |
| Nuclear repulsion energy | 42.115953 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3127 |
2953 |
0.00 |
|
|
|
| 2 |
A1' |
1467 |
1385 |
0.00 |
|
|
|
| 3 |
A1' |
1033 |
976 |
0.00 |
|
|
|
| 4 |
A1" |
311i |
294i |
0.00 |
|
|
|
| 5 |
A2" |
3122 |
2949 |
38.53 |
|
|
|
| 6 |
A2" |
1426 |
1347 |
0.67 |
|
|
|
| 7 |
E' |
3220 |
3041 |
31.39 |
|
|
|
| 7 |
E' |
3220 |
3041 |
31.39 |
|
|
|
| 8 |
E' |
1545 |
1459 |
8.75 |
|
|
|
| 8 |
E' |
1545 |
1459 |
8.75 |
|
|
|
| 9 |
E' |
911 |
860 |
3.78 |
|
|
|
| 9 |
E' |
911 |
860 |
3.78 |
|
|
|
| 10 |
E" |
3202 |
3024 |
0.00 |
|
|
|
| 10 |
E" |
3202 |
3024 |
0.00 |
|
|
|
| 11 |
E" |
1537 |
1452 |
0.00 |
|
|
|
| 11 |
E" |
1537 |
1452 |
0.00 |
|
|
|
| 12 |
E" |
1190 |
1124 |
0.00 |
|
|
|
| 12 |
E" |
1190 |
1124 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16536.5 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 15618.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.770 |
| C2 |
0.000 |
0.000 |
-0.770 |
| H3 |
0.000 |
1.012 |
1.173 |
| H4 |
-0.877 |
-0.506 |
1.173 |
| H5 |
0.877 |
-0.506 |
1.173 |
| H6 |
0.000 |
1.012 |
-1.173 |
| H7 |
0.877 |
-0.506 |
-1.173 |
| H8 |
-0.877 |
-0.506 |
-1.173 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5404 | 1.0896 | 1.0896 | 1.0896 | 2.1911 | 2.1911 | 2.1911 |
C2 | 1.5404 | | 2.1911 | 2.1911 | 2.1911 | 1.0896 | 1.0896 | 1.0896 | H3 | 1.0896 | 2.1911 | | 1.7536 | 1.7536 | 2.3459 | 2.9289 | 2.9289 | H4 | 1.0896 | 2.1911 | 1.7536 | | 1.7536 | 2.9289 | 2.9289 | 2.3459 | H5 | 1.0896 | 2.1911 | 1.7536 | 1.7536 | | 2.9289 | 2.3459 | 2.9289 | H6 | 2.1911 | 1.0896 | 2.3459 | 2.9289 | 2.9289 | | 1.7536 | 1.7536 | H7 | 2.1911 | 1.0896 | 2.9289 | 2.9289 | 2.3459 | 1.7536 | | 1.7536 | H8 | 2.1911 | 1.0896 | 2.9289 | 2.3459 | 2.9289 | 1.7536 | 1.7536 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.691 |
|
C1 |
C2 |
H7 |
111.691 |
| C1 |
C2 |
H8 |
111.691 |
|
C2 |
C1 |
H3 |
111.691 |
| C2 |
C1 |
H4 |
111.691 |
|
C2 |
C1 |
H5 |
111.691 |
| H3 |
C1 |
H4 |
107.163 |
|
H3 |
C1 |
H5 |
107.163 |
| H4 |
C1 |
H5 |
107.163 |
|
H6 |
C2 |
H7 |
107.163 |
| H6 |
C2 |
H8 |
107.163 |
|
H7 |
C2 |
H8 |
107.163 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
