Jump to
S1C2
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -169.529499 |
| Energy at 298.15K | |
| HF Energy | -168.952005 |
| Nuclear repulsion energy | 71.478144 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3830 |
3627 |
55.82 |
|
|
|
| 2 |
A' |
3686 |
3492 |
56.82 |
|
|
|
| 3 |
A' |
3032 |
2872 |
77.50 |
|
|
|
| 4 |
A' |
1891 |
1791 |
375.23 |
|
|
|
| 5 |
A' |
1648 |
1561 |
73.42 |
|
|
|
| 6 |
A' |
1467 |
1390 |
4.04 |
|
|
|
| 7 |
A' |
1300 |
1232 |
116.40 |
|
|
|
| 8 |
A' |
1078 |
1022 |
5.12 |
|
|
|
| 9 |
A' |
573 |
543 |
12.18 |
|
|
|
| 10 |
A" |
1092 |
1035 |
0.14 |
|
|
|
| 11 |
A" |
645 |
611 |
15.69 |
|
|
|
| 12 |
A" |
124i |
117i |
230.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10059.5 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 9528.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.418 |
0.000 |
| O2 |
1.191 |
0.242 |
0.000 |
| N3 |
-0.933 |
-0.568 |
0.000 |
| H4 |
-0.459 |
1.421 |
0.000 |
| H5 |
-0.633 |
-1.526 |
0.000 |
| H6 |
-1.912 |
-0.363 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2042 | 1.3574 | 1.1035 | 2.0446 | 2.0652 |
O2 | 1.2042 | | 2.2735 | 2.0283 | 2.5408 | 3.1618 | N3 | 1.3574 | 2.2735 | | 2.0454 | 1.0039 | 1.0008 | H4 | 1.1035 | 2.0283 | 2.0454 | | 2.9529 | 2.3009 | H5 | 2.0446 | 2.5408 | 1.0039 | 2.9529 | | 1.7295 | H6 | 2.0652 | 3.1618 | 1.0008 | 2.3009 | 1.7295 | |
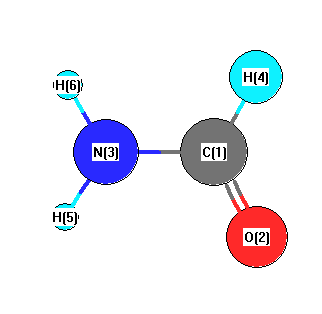 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.217 |
|
C1 |
N3 |
H6 |
121.534 |
| O2 |
C1 |
N3 |
125.019 |
|
O2 |
C1 |
H4 |
122.959 |
| N3 |
C1 |
H4 |
112.022 |
|
H5 |
N3 |
H6 |
119.249 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -169.529515 |
| Energy at 298.15K | -169.533158 |
| HF Energy | -168.951956 |
| Nuclear repulsion energy | 71.456061 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3820 |
3618 |
52.63 |
|
|
|
| 2 |
A |
3679 |
3485 |
52.65 |
|
|
|
| 3 |
A |
3034 |
2873 |
76.88 |
|
|
|
| 4 |
A |
1891 |
1791 |
365.50 |
|
|
|
| 5 |
A |
1649 |
1562 |
70.27 |
|
|
|
| 6 |
A |
1467 |
1390 |
3.79 |
|
|
|
| 7 |
A |
1300 |
1231 |
113.43 |
|
|
|
| 8 |
A |
1096 |
1038 |
1.76 |
|
|
|
| 9 |
A |
1078 |
1021 |
4.09 |
|
|
|
| 10 |
A |
639 |
605 |
22.68 |
|
|
|
| 11 |
A |
573 |
542 |
11.40 |
|
|
|
| 12 |
A |
178 |
168 |
241.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10201.2 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 9662.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.164 |
0.385 |
0.001 |
| O2 |
1.190 |
-0.245 |
0.005 |
| N3 |
-1.083 |
-0.156 |
-0.031 |
| H4 |
0.138 |
1.488 |
0.001 |
| H5 |
-1.175 |
-1.152 |
0.064 |
| H6 |
-1.893 |
0.415 |
0.108 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2041 | 1.3596 | 1.1034 | 2.0391 | 2.0601 |
O2 | 1.2041 | | 2.2751 | 2.0275 | 2.5336 | 3.1550 | N3 | 1.3596 | 2.2751 | | 2.0479 | 1.0045 | 1.0015 | H4 | 1.1034 | 2.0275 | 2.0479 | | 2.9488 | 2.2990 | H5 | 2.0391 | 2.5336 | 1.0045 | 2.9488 | | 1.7249 | H6 | 2.0601 | 3.1550 | 1.0015 | 2.2990 | 1.7249 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.434 |
|
C1 |
N3 |
H6 |
120.760 |
| O2 |
C1 |
N3 |
124.998 |
|
O2 |
C1 |
H4 |
122.907 |
| N3 |
C1 |
H4 |
112.083 |
|
H5 |
N3 |
H6 |
118.605 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
