return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(CH3)CH3 (Isobutane)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3V 1A1
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-157.992713
Energy at 298.15K-158.003452
HF Energy-157.321091
Nuclear repulsion energy134.936940
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3189 3012 56.94      
2 A1 3097 2925 16.34      
3 A1 3093 2921 2.99      
4 A1 1545 1459 15.21      
5 A1 1446 1366 3.53      
6 A1 1227 1159 0.44      
7 A1 826 780 0.66      
8 A1 430 407 0.26      
9 A2 3189 3012 0.00      
10 A2 1511 1427 0.00      
11 A2 963 910 0.00      
12 A2 219 207 0.00      
13 E 3192 3015 32.40      
14 E 3182 3006 4.44      
14 E 3182 3006 4.44      
15 E 3093 2921 22.46      
16 E 1537 1452 3.07      
16 E 1537 1452 3.07      
17 E 1517 1433 0.36      
17 E 1517 1433 0.36      
18 E 1419 1341 7.36      
18 E 1419 1341 7.36      
19 E 1376 1300 2.40      
19 E 1376 1300 2.40      
20 E 1217 1149 1.87      
20 E 1217 1149 1.87      
21 E 1003 947 0.32      
21 E 1003 947 0.32      
22 E 933 881 1.47      
22 E 933 881 1.47      
23 E 363 343 0.00      
23 E 363 343 0.00      
24 E 273 258 0.02      
24 E 273 258 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 29473.0 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 27837.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.26175 0.26175 0.15204

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C3v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.382
H2 0.000 0.000 1.478
C3 0.000 1.449 -0.097
C4 1.255 -0.725 -0.097
C5 -1.255 -0.725 -0.097
H6 0.000 1.490 -1.189
H7 1.290 -0.745 -1.189
H8 -1.290 -0.745 -1.189
H9 0.883 1.985 0.257
H10 -0.883 1.985 0.257
H11 1.277 -1.757 0.257
H12 2.161 -0.227 0.257
H13 -2.161 -0.227 0.257
H14 -1.277 -1.757 0.257

Atom - Atom Distances (Å)
  C1 H2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14
C11.09601.52661.52661.52662.16572.16572.16572.17612.17612.17612.17612.17612.1761
H21.09602.14052.14052.14053.05563.05563.05562.49242.49242.49242.49242.49242.4924
C31.52662.14052.51042.51041.09332.77012.77011.09181.09183.46992.75762.75763.4699
C41.52662.14052.51042.51042.77011.09332.77012.75763.46991.09181.09183.46992.7576
C51.52662.14052.51042.51042.77012.77011.09333.46992.75762.75763.46991.09181.0918
H62.16573.05561.09332.77012.77012.58042.58041.76551.76553.77723.11573.11573.7772
H72.16573.05562.77011.09332.77012.58042.58043.11573.77721.76551.76553.77723.1157
H82.16573.05562.77012.77011.09332.58042.58043.77723.11573.11573.77721.76551.7655
H92.17612.49241.09182.75763.46991.76553.11573.77721.76693.76292.55423.76294.3211
H102.17612.49241.09183.46992.75761.76553.77723.11571.76694.32113.76292.55423.7629
H112.17612.49243.46991.09182.75763.77721.76553.11573.76294.32111.76693.76292.5542
H122.17612.49242.75761.09183.46993.11571.76553.77722.55423.76291.76694.32113.7629
H132.17612.49242.75763.46991.09183.11573.77721.76553.76292.55423.76294.32111.7669
H142.17612.49243.46992.75761.09183.77723.11571.76554.32113.76292.55423.76291.7669

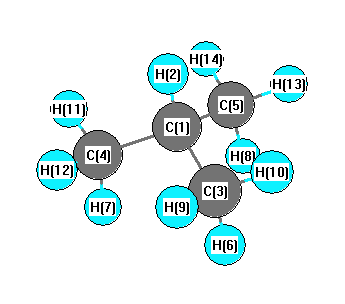
picture of Isobutane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H6 110.414 C1 C3 H9 111.338
C1 C3 H10 111.338 C1 C4 H7 110.414
C1 C4 H11 111.338 C1 C4 H12 111.338
C1 C5 H8 110.414 C1 C5 H13 111.338
C1 C5 H14 111.338 H2 C1 C3 108.296
H2 C1 C4 108.296 H2 C1 C5 108.296
C3 C1 C4 110.621 C3 C1 C5 110.621
C4 C1 C5 110.621 H6 C3 H9 107.788
H6 C3 H10 107.788 H7 C4 H11 107.788
H7 C4 H12 107.788 H8 C5 H13 107.788
H8 C5 H14 107.788 H9 C3 H10 108.024
H11 C4 H12 108.024 H13 C5 H14 108.024
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability