return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(NH2)CH3 (2-Propanamine)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-174.015898
Energy at 298.15K-174.026462
HF Energy-173.311906
Nuclear repulsion energy136.013916
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3539 3343 0.16      
2 A' 3207 3029 30.14      
3 A' 3190 3013 27.34      
4 A' 3100 2928 3.51      
5 A' 3023 2855 57.09      
6 A' 1675 1582 32.17      
7 A' 1540 1454 5.47      
8 A' 1524 1440 3.43      
9 A' 1434 1354 15.85      
10 A' 1399 1322 15.61      
11 A' 1227 1159 5.15      
12 A' 1180 1115 20.04      
13 A' 1028 971 5.73      
14 A' 895 846 111.14      
15 A' 844 797 3.92      
16 A' 474 447 10.57      
17 A' 364 344 0.14      
18 A' 281 266 0.32      
19 A" 3636 3434 0.63      
20 A" 3203 3026 0.39      
21 A" 3188 3011 14.19      
22 A" 3098 2926 24.84      
23 A" 1518 1434 0.14      
24 A" 1514 1430 0.44      
25 A" 1438 1358 15.49      
26 A" 1402 1324 0.26      
27 A" 1285 1214 0.00      
28 A" 1064 1005 0.86      
29 A" 977 923 0.65      
30 A" 938 886 0.45      
31 A" 404 382 6.59      
32 A" 308 291 32.57      
33 A" 227 214 5.94      

Unscaled Zero Point Vibrational Energy (zpe) 27062.0 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 25560.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.28080 0.26798 0.15712

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.296 0.227 0.000
N2 -0.921 1.035 0.000
H3 1.216 0.833 0.000
C4 0.296 -0.635 1.253
C5 0.296 -0.635 -1.253
H6 -0.930 1.642 -0.812
H7 -0.930 1.642 0.812
H8 -0.594 -1.267 1.269
H9 -0.594 -1.267 -1.269
H10 1.183 -1.271 1.287
H11 1.183 -1.271 -1.287
H12 0.291 -0.016 2.154
H13 0.291 -0.016 -2.154

Atom - Atom Distances (Å)
  C1 N2 H3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.46141.10151.52131.52132.04032.04032.15292.15292.16512.16512.16772.1677
N21.46142.14742.41732.41731.01341.01342.64882.64883.37693.37692.68602.6860
H31.10152.14742.13832.13832.43272.43273.04893.04892.46672.46672.49332.4933
C41.52132.41732.13832.50683.30932.62341.09152.74871.09152.76521.09273.4631
C51.52132.41732.13832.50682.62343.30932.74871.09152.76521.09153.46311.0927
H62.04031.01342.43273.30932.62341.62353.59202.96334.16573.62943.61022.4576
H72.04031.01342.43272.62343.30931.62352.96333.59203.62944.16572.45763.6102
H82.15292.64883.04891.09152.74873.59202.96332.53851.77683.11351.76903.7503
H92.15292.64883.04892.74871.09152.96333.59202.53853.11351.77683.75031.7690
H102.16513.37692.46671.09152.76524.16573.62941.77683.11352.57501.76663.7701
H112.16513.37692.46672.76521.09153.62944.16573.11351.77682.57503.77011.7666
H122.16772.68602.49331.09273.46313.61022.45761.76903.75031.76663.77014.3079
H132.16772.68602.49333.46311.09272.45763.61023.75031.76903.77011.76664.3079

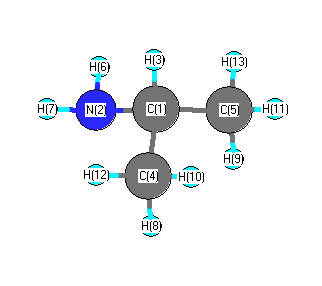
picture of 2-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H6 109.732 C1 N2 H7 109.732
C1 C4 H8 109.870 C1 C4 H10 110.846
C1 C4 H12 110.974 C1 C5 H9 109.870
C1 C5 H11 110.846 C1 C5 H13 110.974
N2 C1 H3 113.078 N2 C1 C4 108.260
N2 C1 C5 108.260 H3 C1 C4 108.159
H3 C1 C5 108.159 C4 C1 C5 110.950
H6 N2 H7 106.445 H8 C4 H10 108.959
H8 C4 H12 108.163 H9 C5 H11 108.959
H9 C5 H13 108.163 H10 C4 H12 107.951
H11 C5 H13 107.951
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability