return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(NH2)CH3 (2-Propanamine)

using model chemistry: QCISD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G(2df,p)
 hartrees
Energy at 0K-174.067178
Energy at 298.15K-174.077735
HF Energy-173.311894
Nuclear repulsion energy135.691732
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3534 3348 0.42      
2 A' 3171 3004 43.15      
3 A' 3153 2988 37.38      
4 A' 3079 2917 4.05      
5 A' 3000 2843 59.81      
6 A' 1693 1604 31.55      
7 A' 1541 1460 4.45      
8 A' 1527 1447 2.97      
9 A' 1448 1371 14.14      
10 A' 1411 1337 15.46      
11 A' 1231 1166 4.57      
12 A' 1188 1126 21.53      
13 A' 1028 974 6.49      
14 A' 909 861 110.64      
15 A' 840 796 1.68      
16 A' 475 450 10.07      
17 A' 366 346 0.13      
18 A' 275 260 0.29      
19 A" 3617 3427 0.00      
20 A" 3165 2999 0.54      
21 A" 3150 2985 17.34      
22 A" 3076 2914 30.34      
23 A" 1522 1442 0.07      
24 A" 1518 1438 0.25      
25 A" 1454 1377 14.33      
26 A" 1414 1340 0.02      
27 A" 1292 1224 0.00      
28 A" 1065 1009 1.09      
29 A" 980 929 0.49      
30 A" 942 893 0.23      
31 A" 405 384 6.07      
32 A" 303 287 33.34      
33 A" 223 211 5.80      

Unscaled Zero Point Vibrational Energy (zpe) 26998.4 cm-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 25578.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G(2df,p)
ABC
0.27929 0.26647 0.15608

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.295 0.225 0.000
N2 -0.919 1.045 0.000
H3 1.217 0.829 0.000
C4 0.295 -0.639 1.259
C5 0.295 -0.639 -1.259
H6 -0.922 1.652 -0.811
H7 -0.922 1.652 0.811
H8 -0.597 -1.270 1.279
H9 -0.597 -1.270 -1.279
H10 1.181 -1.278 1.293
H11 1.181 -1.278 -1.293
H12 0.293 -0.018 2.160
H13 0.293 -0.018 -2.160

Atom - Atom Distances (Å)
  C1 N2 H3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.46471.10241.52621.52622.04392.04392.16022.16022.17102.17102.17342.1734
N21.46472.14642.42712.42711.01321.01322.66462.66463.38743.38742.69482.6948
H31.10242.14642.14182.14182.43162.43163.05503.05502.47172.47172.49702.4970
C41.52622.42712.14182.51713.31862.63231.09322.76331.09322.77541.09443.4743
C51.52622.42712.14182.51712.63233.31862.76331.09322.77541.09323.47431.0944
H62.04391.01322.43163.31862.63231.62263.60772.97724.17553.63863.61842.4666
H72.04391.01322.43162.63233.31861.62262.97723.60773.63864.17552.46663.6184
H82.16022.66463.05501.09322.76333.60772.97722.55871.77843.12701.77093.7668
H92.16022.66463.05502.76331.09322.97723.60772.55873.12701.77843.76681.7709
H102.17103.38742.47171.09322.77544.17553.63861.77843.12702.58551.76853.7810
H112.17103.38742.47172.77541.09323.63864.17553.12701.77842.58553.78101.7685
H122.17342.69482.49701.09443.47433.61842.46661.77093.76681.76853.78104.3196
H132.17342.69482.49703.47431.09442.46663.61843.76681.77093.78101.76854.3196

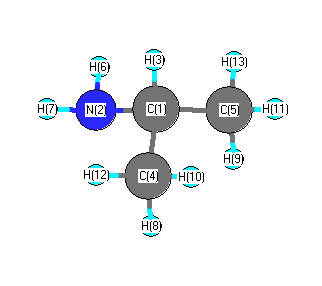
picture of 2-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H6 109.803 C1 N2 H7 109.803
C1 C4 H8 110.024 C1 C4 H10 110.879
C1 C4 H12 110.993 C1 C5 H9 110.024
C1 C5 H11 110.879 C1 C5 H13 110.993
N2 C1 H3 112.701 N2 C1 C4 108.473
N2 C1 C5 108.473 H3 C1 C4 108.057
H3 C1 C5 108.057 C4 C1 C5 111.110
H6 N2 H7 106.396 H8 C4 H10 108.867
H8 C4 H12 108.102 H9 C5 H11 108.867
H9 C5 H13 108.102 H10 C4 H12 107.891
H11 C5 H13 107.891
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability