return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3SH (2-Propanethiol, 2-methyl-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-555.673305
Energy at 298.15K-555.684188
HF Energy-554.836840
Nuclear repulsion energy245.669709
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3203 3026 14.68      
2 A' 3189 3012 15.82      
3 A' 3180 3004 24.56      
4 A' 3095 2923 11.50      
5 A' 3090 2918 20.05      
6 A' 2771 2617 1.88      
7 A' 1540 1454 6.68      
8 A' 1525 1441 8.89      
9 A' 1510 1426 0.22      
10 A' 1443 1363 0.87      
11 A' 1414 1335 9.59      
12 A' 1280 1209 0.38      
13 A' 1219 1151 39.26      
14 A' 1073 1013 1.90      
15 A' 965 911 0.63      
16 A' 889 839 3.89      
17 A' 857 809 1.42      
18 A' 615 581 3.13      
19 A' 391 369 0.86      
20 A' 369 348 0.17      
21 A' 311 294 0.07      
22 A' 288 272 0.91      
23 A" 3202 3024 10.41      
24 A" 3197 3020 4.22      
25 A" 3176 3000 0.31      
26 A" 3087 2916 11.19      
27 A" 1527 1442 7.50      
28 A" 1506 1423 0.22      
29 A" 1500 1416 0.00      
30 A" 1414 1336 9.52      
31 A" 1267 1197 2.80      
32 A" 1063 1004 0.19      
33 A" 974 920 0.02      
34 A" 956 903 0.48      
35 A" 398 376 0.41      
36 A" 309 292 2.69      
37 A" 294 278 0.15      
38 A" 264 249 6.38      
39 A" 213 201 10.70      

Unscaled Zero Point Vibrational Energy (zpe) 29278.9 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 27653.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.15055 0.10064 0.09973

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.340 -0.010 0.000
S2 -1.491 0.094 0.000
C3 0.828 1.438 0.000
C4 0.828 -0.730 1.254
C5 0.828 -0.730 -1.254
H6 -1.717 -1.226 0.000
H7 1.921 1.454 0.000
H8 0.478 1.971 -0.886
H9 0.478 1.971 0.886
H10 1.922 -0.753 1.267
H11 1.922 -0.753 -1.267
H12 0.479 -0.224 2.155
H13 0.470 -1.761 1.279
H14 0.479 -0.224 -2.155
H15 0.470 -1.761 -1.279

Atom - Atom Distances (Å)
  C1 S2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.83431.52861.52631.52632.39002.15482.17512.17512.15872.15872.16982.17272.16982.1727
S21.83432.68102.76242.76241.33863.67342.86172.86173.73813.73812.93662.98702.93662.9870
C31.52862.68102.50522.50523.68491.09291.09171.09172.75752.75752.74403.46482.74403.4648
C41.52632.76242.50522.50832.88092.74573.46442.74901.09432.74871.09071.09213.46402.7589
C51.52632.76242.50522.50832.88092.74572.74903.46442.74871.09433.46402.75891.09071.0921
H62.39001.33863.68492.88092.88094.51903.97843.97843.88303.88303.23552.58983.23552.5898
H72.15483.67341.09292.74572.74574.51901.77051.77052.54492.54493.08883.75303.08883.7530
H82.17512.86171.09173.46442.74903.97841.77051.77243.76073.10663.75074.31562.53573.7535
H92.17512.86171.09172.74903.46443.97841.77051.77243.10663.76072.53573.75353.75074.3156
H102.15873.73812.75751.09432.74873.88302.54493.76073.10662.53471.77511.76873.75163.1007
H112.15873.73812.75752.74871.09433.88302.54493.10663.76072.53473.75163.10071.77511.7687
H122.16982.93662.74401.09073.46403.23553.08883.75072.53571.77513.75161.76904.30953.7626
H132.17272.98703.46481.09212.75892.58983.75304.31563.75351.76873.10071.76903.76262.5590
H142.16982.93662.74403.46401.09073.23553.08882.53573.75073.75161.77514.30953.76261.7690
H152.17272.98703.46482.75891.09212.58983.75303.75354.31563.10071.76873.76262.55901.7690

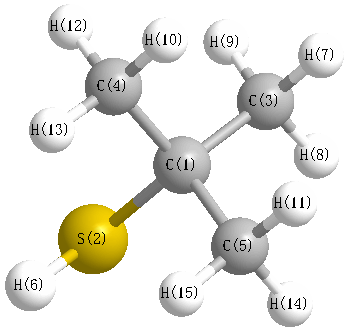
picture of 2-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 H6 96.493 C1 C3 H7 109.443
C1 C3 H8 111.118 C1 C3 H9 111.118
C1 C4 H10 109.829 C1 C4 H12 110.921
C1 C4 H13 111.070 C1 C5 H11 109.829
C1 C5 H14 110.921 C1 C5 H15 111.070
S2 C1 C3 105.369 S2 C1 C4 110.239
S2 C1 C5 110.239 C3 C1 C4 110.184
C3 C1 C5 110.184 C4 C1 C5 110.511
H7 C3 H8 108.272 H7 C3 H9 108.272
H8 C3 H9 108.530 H10 C4 H12 108.667
H10 C4 H13 107.989 H11 C5 H14 108.667
H11 C5 H15 107.989 H12 C4 H13 108.278
H14 C5 H15 108.278
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability