Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -637.718051 |
| Energy at 298.15K | -637.722980 |
| HF Energy | -636.995548 |
| Nuclear repulsion energy | 157.768283 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3161 |
2985 |
10.24 |
|
|
|
| 2 |
A' |
3121 |
2948 |
14.85 |
|
|
|
| 3 |
A' |
1552 |
1466 |
1.33 |
|
|
|
| 4 |
A' |
1518 |
1434 |
5.37 |
|
|
|
| 5 |
A' |
1448 |
1368 |
5.77 |
|
|
|
| 6 |
A' |
1296 |
1224 |
9.10 |
|
|
|
| 7 |
A' |
1117 |
1055 |
91.28 |
|
|
|
| 8 |
A' |
1098 |
1037 |
10.86 |
|
|
|
| 9 |
A' |
820 |
775 |
37.63 |
|
|
|
| 10 |
A' |
391 |
369 |
2.11 |
|
|
|
| 11 |
A' |
242 |
228 |
9.99 |
|
|
|
| 12 |
A" |
3238 |
3058 |
3.44 |
|
|
|
| 13 |
A" |
3187 |
3010 |
14.73 |
|
|
|
| 14 |
A" |
1322 |
1249 |
0.02 |
|
|
|
| 15 |
A" |
1244 |
1175 |
1.31 |
|
|
|
| 16 |
A" |
1082 |
1022 |
2.06 |
|
|
|
| 17 |
A" |
802 |
758 |
0.63 |
|
|
|
| 18 |
A" |
137 |
130 |
8.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13388.3 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 12645.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.617 |
0.000 |
| C2 |
0.985 |
-0.535 |
0.000 |
| Cl3 |
-1.656 |
-0.036 |
0.000 |
| F4 |
2.255 |
-0.005 |
0.000 |
| H5 |
0.122 |
1.232 |
0.889 |
| H6 |
0.122 |
1.232 |
-0.889 |
| H7 |
0.852 |
-1.153 |
0.890 |
| H8 |
0.852 |
-1.153 |
-0.890 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5154 | 1.7802 | 2.3393 | 1.0882 | 1.0882 | 2.1568 | 2.1568 |
C2 | 1.5154 | | 2.6879 | 1.3761 | 2.1580 | 2.1580 | 1.0919 | 1.0919 | Cl3 | 1.7802 | 2.6879 | | 3.9114 | 2.3581 | 2.3581 | 2.8867 | 2.8867 | F4 | 2.3393 | 1.3761 | 3.9114 | | 2.6214 | 2.6214 | 2.0195 | 2.0195 | H5 | 1.0882 | 2.1580 | 2.3581 | 2.6214 | | 1.7787 | 2.4946 | 3.0641 | H6 | 1.0882 | 2.1580 | 2.3581 | 2.6214 | 1.7787 | | 3.0641 | 2.4946 | H7 | 2.1568 | 1.0919 | 2.8867 | 2.0195 | 2.4946 | 3.0641 | | 1.7798 | H8 | 2.1568 | 1.0919 | 2.8867 | 2.0195 | 3.0641 | 2.4946 | 1.7798 | |
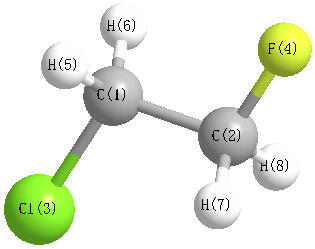 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.907 |
|
C1 |
C2 |
H7 |
110.578 |
| C1 |
C2 |
H8 |
110.578 |
|
C2 |
C1 |
Cl3 |
109.029 |
| C2 |
C1 |
H5 |
110.897 |
|
C2 |
C1 |
H6 |
110.897 |
| Cl3 |
C1 |
H5 |
108.153 |
|
Cl3 |
C1 |
H6 |
108.153 |
| F4 |
C2 |
H7 |
109.284 |
|
F4 |
C2 |
H8 |
109.284 |
| H5 |
C1 |
H6 |
109.626 |
|
H7 |
C2 |
H8 |
109.182 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -637.717777 |
| Energy at 298.15K | -637.722817 |
| HF Energy | -636.994550 |
| Nuclear repulsion energy | 162.358298 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3225 |
3046 |
2.09 |
|
|
|
| 2 |
A |
3172 |
2996 |
19.48 |
|
|
|
| 3 |
A |
3149 |
2974 |
7.15 |
|
|
|
| 4 |
A |
3099 |
2927 |
24.17 |
|
|
|
| 5 |
A |
1536 |
1451 |
2.47 |
|
|
|
| 6 |
A |
1490 |
1408 |
8.49 |
|
|
|
| 7 |
A |
1453 |
1373 |
12.91 |
|
|
|
| 8 |
A |
1354 |
1279 |
25.69 |
|
|
|
| 9 |
A |
1297 |
1225 |
0.65 |
|
|
|
| 10 |
A |
1244 |
1175 |
4.22 |
|
|
|
| 11 |
A |
1153 |
1089 |
57.58 |
|
|
|
| 12 |
A |
1088 |
1028 |
12.45 |
|
|
|
| 13 |
A |
996 |
941 |
4.71 |
|
|
|
| 14 |
A |
875 |
826 |
5.64 |
|
|
|
| 15 |
A |
725 |
685 |
24.18 |
|
|
|
| 16 |
A |
472 |
446 |
10.83 |
|
|
|
| 17 |
A |
287 |
271 |
0.84 |
|
|
|
| 18 |
A |
138 |
130 |
1.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13377.4 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 12635.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.087 |
0.860 |
-0.288 |
| C2 |
1.204 |
0.411 |
0.352 |
| Cl3 |
-1.405 |
-0.284 |
0.066 |
| F4 |
1.611 |
-0.775 |
-0.202 |
| H5 |
-0.381 |
1.837 |
0.094 |
| H6 |
0.021 |
0.907 |
-1.370 |
| H7 |
1.974 |
1.169 |
0.172 |
| H8 |
1.075 |
0.275 |
1.428 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5093 | 1.7808 | 2.3586 | 1.0896 | 1.0889 | 2.1340 | 2.1533 |
C2 | 1.5093 | | 2.7153 | 1.3705 | 2.1477 | 2.1473 | 1.0951 | 1.0922 | Cl3 | 1.7808 | 2.7153 | | 3.0674 | 2.3561 | 2.3487 | 3.6796 | 2.8845 | F4 | 2.3586 | 1.3705 | 3.0674 | | 3.2980 | 2.5923 | 2.0121 | 2.0118 | H5 | 1.0896 | 2.1477 | 2.3561 | 3.2980 | | 1.7807 | 2.4488 | 2.5177 | H6 | 1.0889 | 2.1473 | 2.3487 | 2.5923 | 1.7807 | | 2.5018 | 3.0562 | H7 | 2.1340 | 1.0951 | 3.6796 | 2.0121 | 2.4488 | 2.5018 | | 1.7842 | H8 | 2.1533 | 1.0922 | 2.8845 | 2.0118 | 2.5177 | 3.0562 | 1.7842 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.875 |
|
C1 |
C2 |
H7 |
109.010 |
| C1 |
C2 |
H8 |
110.704 |
|
C2 |
C1 |
Cl3 |
110.966 |
| C2 |
C1 |
H5 |
110.416 |
|
C2 |
C1 |
H6 |
110.431 |
| Cl3 |
C1 |
H5 |
107.895 |
|
Cl3 |
C1 |
H6 |
107.397 |
| F4 |
C2 |
H7 |
108.879 |
|
F4 |
C2 |
H8 |
109.026 |
| H5 |
C1 |
H6 |
109.651 |
|
H7 |
C2 |
H8 |
109.318 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
