return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7N (Cyclopropylamine)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-172.788017
Energy at 298.15K-172.796498
HF Energy-172.106578
Nuclear repulsion energy125.519282
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3552 3355 0.33      
2 A' 3324 3139 5.53      
3 A' 3216 3038 3.04      
4 A' 3161 2985 33.47      
5 A' 1675 1582 15.20      
6 A' 1524 1439 16.16      
7 A' 1433 1354 16.91      
8 A' 1265 1195 0.21      
9 A' 1196 1130 6.86      
10 A' 1068 1008 14.78      
11 A' 1041 983 9.48      
12 A' 878 830 113.57      
13 A' 843 796 25.16      
14 A' 782 738 2.65      
15 A' 406 384 5.74      
16 A" 3648 3445 3.80      
17 A" 3313 3129 0.31      
18 A" 3211 3033 8.56      
19 A" 1469 1387 1.68      
20 A" 1288 1216 0.96      
21 A" 1211 1144 0.11      
22 A" 1150 1086 0.60      
23 A" 1079 1019 1.49      
24 A" 932 880 8.80      
25 A" 854 806 1.69      
26 A" 406 383 8.71      
27 A" 281 266 31.28      

Unscaled Zero Point Vibrational Energy (zpe) 22101.6 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 20874.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.54600 0.22636 0.19542

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.311 0.411 0.000
H2 -1.266 0.936 0.000
N3 0.891 1.193 0.000
C4 -0.311 -0.887 0.751
C5 -0.311 -0.887 -0.751
H6 0.926 1.786 0.819
H7 0.926 1.786 -0.819
H8 -1.213 -1.174 1.275
H9 -1.213 -1.174 -1.275
H10 0.600 -1.165 1.261
H11 0.600 -1.165 -1.261

Atom - Atom Distances (Å)
  C1 H2 N3 C4 C5 H6 H7 H8 H9 H10 H11
C11.09001.43451.50001.50002.02272.02272.22532.22532.21532.2153
H21.09002.17312.19112.19112.49012.49012.46592.46593.08093.0809
N31.43452.17312.51772.51771.01111.01113.41453.41452.69052.6905
C41.50002.19112.51771.50262.94633.33781.08182.23661.08102.2269
C51.50002.19112.51771.50263.33782.94632.23661.08182.22691.0810
H62.02272.49011.01112.94633.33781.63703.68054.20973.00193.6251
H72.02272.49011.01113.33782.94631.63704.20973.68053.62513.0019
H82.22532.46593.41451.08182.23663.68054.20972.55041.81383.1183
H92.22532.46593.41452.23661.08184.20973.68052.55043.11831.8138
H102.21533.08092.69051.08102.22693.00193.62511.81383.11832.5227
H112.21533.08092.69052.22691.08103.62513.00193.11831.81382.5227

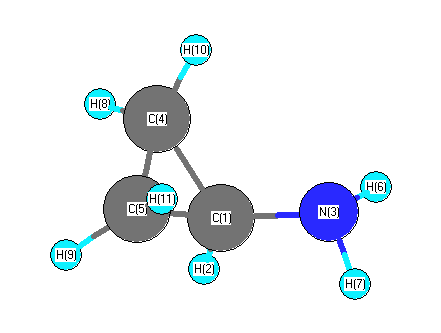
picture of Cyclopropylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 H6 110.401 C1 N3 H7 110.401
C1 C4 C5 59.945 C1 C4 H8 118.159
C1 C4 H10 117.324 C1 C5 C4 59.945
C1 C5 H9 118.159 C1 C5 H11 117.324
H2 C1 N3 118.173 H2 C1 C4 114.632
H2 C1 C5 114.632 N3 C1 C4 118.160
N3 C1 C5 118.160 C4 C1 C5 60.111
C4 C5 H9 118.965 C4 C5 H11 118.155
C5 C4 H8 118.965 C5 C4 H10 118.155
H6 N3 H7 108.107 H8 C4 H10 113.983
H9 C5 H11 113.983
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability