return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7N (Cyclopropylamine)

using model chemistry: QCISD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at QCISD/6-31G(2df,p)
 hartrees
Energy at 0K-172.828309
Energy at 298.15K-172.836800
HF Energy-172.106540
Nuclear repulsion energy125.273477
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3546 3359 0.26      
2 A' 3291 3118 11.84      
3 A' 3196 3027 4.69      
4 A' 3141 2976 37.13      
5 A' 1694 1604 15.29      
6 A' 1537 1457 15.31      
7 A' 1443 1367 16.90      
8 A' 1263 1197 0.29      
9 A' 1201 1138 8.43      
10 A' 1085 1028 12.70      
11 A' 1041 986 7.91      
12 A' 894 847 121.29      
13 A' 839 795 14.57      
14 A' 783 742 2.37      
15 A' 407 386 5.79      
16 A" 3628 3437 1.13      
17 A" 3279 3107 0.23      
18 A" 3188 3021 13.59      
19 A" 1478 1400 1.15      
20 A" 1301 1233 0.98      
21 A" 1219 1155 0.11      
22 A" 1160 1099 0.35      
23 A" 1100 1042 0.93      
24 A" 934 885 9.33      
25 A" 857 812 2.09      
26 A" 404 383 8.12      
27 A" 278 264 31.50      

Unscaled Zero Point Vibrational Energy (zpe) 22093.8 cm-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 20931.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G(2df,p)
ABC
0.54508 0.22501 0.19416

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.311 0.410 0.000
H2 -1.266 0.938 0.000
N3 0.891 1.199 0.000
C4 -0.311 -0.890 0.752
C5 -0.311 -0.890 -0.752
H6 0.924 1.792 0.818
H7 0.924 1.792 -0.818
H8 -1.214 -1.178 1.277
H9 -1.214 -1.178 -1.277
H10 0.601 -1.170 1.265
H11 0.601 -1.170 -1.265

Atom - Atom Distances (Å)
  C1 H2 N3 C4 C5 H6 H7 H8 H9 H10 H11
C11.09091.43771.50241.50242.02582.02582.22942.22942.21992.2199
H21.09092.17332.19502.19502.48952.48952.47202.47203.08633.0863
N31.43772.17332.52472.52471.01091.01093.42253.42252.70082.7008
C41.50242.19502.52471.50502.95363.34441.08332.23991.08242.2313
C51.50242.19502.52471.50503.34442.95362.23991.08332.23131.0824
H62.02582.48951.01092.95363.34441.63553.68884.21713.01303.6353
H72.02582.48951.01093.34442.95361.63554.21713.68883.63533.0130
H82.22942.47203.42251.08332.23993.68884.21712.55401.81443.1229
H92.22942.47203.42252.23991.08334.21713.68882.55403.12291.8144
H102.21993.08632.70081.08242.23133.01303.63531.81443.12292.5296
H112.21993.08632.70082.23131.08243.63533.01303.12291.81442.5296

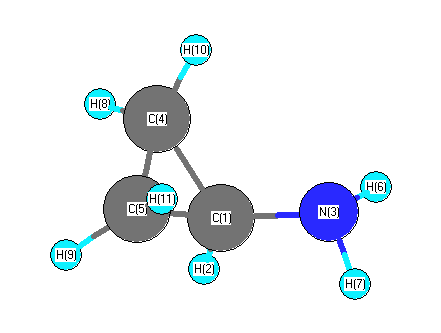
picture of Cyclopropylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 H6 110.435 C1 N3 H7 110.435
C1 C4 C5 59.943 C1 C4 H8 118.228
C1 C4 H10 117.449 C1 C5 C4 59.943
C1 C5 H9 118.228 C1 C5 H11 117.449
H2 C1 N3 117.860 H2 C1 C4 114.721
H2 C1 C5 114.721 N3 C1 C4 118.328
N3 C1 C5 118.328 C4 C1 C5 60.115
C4 C5 H9 118.959 C4 C5 H11 118.250
C5 C4 H8 118.959 C5 C4 H10 118.250
H6 N3 H7 107.979 H8 C4 H10 113.814
H9 C5 H11 113.814
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability