Jump to
S1C2
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Geometric Data calculated at MP2/6-31G(2df,p)
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -235.177110 |
| Energy at 298.15K | -235.189373 |
| HF Energy | -234.206661 |
| Nuclear repulsion energy | 231.055924 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3222 |
3043 |
35.35 |
|
|
|
| 2 |
A |
3210 |
3032 |
4.91 |
|
|
|
| 3 |
A |
3196 |
3019 |
21.54 |
|
|
|
| 4 |
A |
3195 |
3017 |
34.71 |
|
|
|
| 5 |
A |
3191 |
3014 |
0.86 |
|
|
|
| 6 |
A |
3170 |
2994 |
11.39 |
|
|
|
| 7 |
A |
3165 |
2990 |
14.68 |
|
|
|
| 8 |
A |
3147 |
2972 |
1.35 |
|
|
|
| 9 |
A |
3104 |
2932 |
35.26 |
|
|
|
| 10 |
A |
3101 |
2929 |
13.53 |
|
|
|
| 11 |
A |
3099 |
2927 |
12.86 |
|
|
|
| 12 |
A |
3090 |
2918 |
10.24 |
|
|
|
| 13 |
A |
1730 |
1634 |
1.35 |
|
|
|
| 14 |
A |
1538 |
1453 |
5.14 |
|
|
|
| 15 |
A |
1530 |
1445 |
6.58 |
|
|
|
| 16 |
A |
1527 |
1442 |
4.72 |
|
|
|
| 17 |
A |
1520 |
1435 |
0.17 |
|
|
|
| 18 |
A |
1513 |
1429 |
4.73 |
|
|
|
| 19 |
A |
1506 |
1423 |
0.53 |
|
|
|
| 20 |
A |
1449 |
1369 |
7.58 |
|
|
|
| 21 |
A |
1430 |
1351 |
1.90 |
|
|
|
| 22 |
A |
1419 |
1340 |
4.23 |
|
|
|
| 23 |
A |
1387 |
1310 |
0.36 |
|
|
|
| 24 |
A |
1339 |
1265 |
0.02 |
|
|
|
| 25 |
A |
1308 |
1235 |
0.48 |
|
|
|
| 26 |
A |
1301 |
1229 |
2.98 |
|
|
|
| 27 |
A |
1267 |
1196 |
0.08 |
|
|
|
| 28 |
A |
1180 |
1115 |
0.63 |
|
|
|
| 29 |
A |
1131 |
1068 |
1.51 |
|
|
|
| 30 |
A |
1085 |
1024 |
1.36 |
|
|
|
| 31 |
A |
1072 |
1012 |
0.76 |
|
|
|
| 32 |
A |
1069 |
1010 |
0.69 |
|
|
|
| 33 |
A |
1023 |
967 |
5.12 |
|
|
|
| 34 |
A |
1015 |
959 |
0.25 |
|
|
|
| 35 |
A |
933 |
881 |
3.20 |
|
|
|
| 36 |
A |
913 |
862 |
5.45 |
|
|
|
| 37 |
A |
879 |
830 |
3.00 |
|
|
|
| 38 |
A |
755 |
713 |
1.85 |
|
|
|
| 39 |
A |
734 |
693 |
26.40 |
|
|
|
| 40 |
A |
566 |
534 |
6.83 |
|
|
|
| 41 |
A |
479 |
452 |
0.79 |
|
|
|
| 42 |
A |
365 |
345 |
0.03 |
|
|
|
| 43 |
A |
296 |
280 |
0.06 |
|
|
|
| 44 |
A |
251 |
237 |
0.00 |
|
|
|
| 45 |
A |
201 |
190 |
0.02 |
|
|
|
| 46 |
A |
121 |
115 |
0.26 |
|
|
|
| 47 |
A |
94 |
89 |
0.01 |
|
|
|
| 48 |
A |
40 |
38 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36927.2 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 34877.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.409 |
0.979 |
-0.040 |
| C2 |
1.980 |
-0.452 |
-0.167 |
| C3 |
0.775 |
-0.966 |
0.112 |
| C4 |
-0.443 |
-0.234 |
0.588 |
| C5 |
-1.531 |
-0.182 |
-0.491 |
| C6 |
-2.802 |
0.502 |
0.003 |
| H7 |
1.605 |
1.632 |
0.297 |
| H8 |
3.239 |
1.074 |
0.666 |
| H9 |
2.769 |
1.358 |
-1.000 |
| H10 |
2.745 |
-1.136 |
-0.525 |
| H11 |
0.637 |
-2.033 |
-0.050 |
| H12 |
-0.199 |
0.783 |
0.902 |
| H13 |
-0.850 |
-0.742 |
1.470 |
| H14 |
-1.135 |
0.342 |
-1.366 |
| H15 |
-1.760 |
-1.201 |
-0.820 |
| H16 |
-3.567 |
0.537 |
-0.775 |
| H17 |
-3.220 |
-0.027 |
0.861 |
| H18 |
-2.595 |
1.528 |
0.314 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4989 | 2.5451 | 3.1620 | 4.1321 | 5.2328 | 1.0888 | 1.0932 | 1.0931 | 2.1953 | 3.4946 | 2.7796 | 3.9825 | 3.8372 | 4.7687 | 6.0371 | 5.7891 | 5.0470 |
C2 | 1.4989 | | 1.3398 | 2.5469 | 3.5358 | 4.8788 | 2.1668 | 2.1456 | 2.1426 | 1.0868 | 2.0780 | 2.7229 | 3.2818 | 3.4307 | 3.8697 | 5.6667 | 5.3179 | 5.0084 | C3 | 2.5451 | 1.3398 | | 1.4983 | 2.5089 | 3.8679 | 2.7336 | 3.2461 | 3.2581 | 2.0774 | 1.0877 | 2.1523 | 2.1289 | 2.7466 | 2.7109 | 4.6792 | 4.1718 | 4.1978 | C4 | 3.1620 | 2.5469 | 1.4983 | | 1.5333 | 2.5395 | 2.7856 | 3.9075 | 3.9204 | 3.4945 | 2.1928 | 1.0923 | 1.0960 | 2.1511 | 2.1566 | 3.4942 | 2.7986 | 2.7952 | C5 | 4.1321 | 3.5358 | 2.5089 | 1.5333 | | 1.5259 | 3.7076 | 5.0659 | 4.5953 | 4.3805 | 2.8838 | 2.1560 | 2.1500 | 1.0937 | 1.0943 | 2.1778 | 2.1699 | 2.1700 | C6 | 5.2328 | 4.8788 | 3.8679 | 2.5395 | 1.5259 | | 4.5590 | 6.1036 | 5.7244 | 5.8073 | 4.2722 | 2.7683 | 2.7404 | 2.1625 | 2.1591 | 1.0912 | 1.0920 | 1.0921 | H7 | 1.0888 | 2.1668 | 2.7336 | 2.7856 | 3.7076 | 4.5590 | | 1.7652 | 1.7635 | 3.1034 | 3.8061 | 2.0833 | 3.6106 | 3.4548 | 4.5381 | 5.3942 | 5.1338 | 4.2020 | H8 | 1.0932 | 2.1456 | 3.2461 | 3.9075 | 5.0659 | 6.1036 | 1.7652 | | 1.7539 | 2.5577 | 4.1148 | 3.4578 | 4.5451 | 4.8776 | 5.6891 | 6.9769 | 6.5550 | 5.8623 | H9 | 1.0931 | 2.1426 | 3.2581 | 3.9204 | 4.5953 | 5.7244 | 1.7635 | 1.7539 | | 2.5386 | 4.1165 | 3.5712 | 4.8582 | 4.0503 | 5.2048 | 6.3925 | 6.4227 | 5.5255 | H10 | 2.1953 | 1.0868 | 2.0774 | 3.4945 | 4.3805 | 5.8073 | 3.1034 | 2.5577 | 2.5386 | | 2.3397 | 3.7924 | 4.1298 | 4.2359 | 4.5148 | 6.5341 | 6.2235 | 6.0264 | H11 | 3.4946 | 2.0780 | 1.0877 | 2.1928 | 2.8838 | 4.2722 | 3.8061 | 4.1148 | 4.1165 | 2.3397 | | 3.0879 | 2.4873 | 3.2418 | 2.6511 | 4.9796 | 4.4419 | 4.8228 | H12 | 2.7796 | 2.7229 | 2.1523 | 1.0923 | 2.1560 | 2.7683 | 2.0833 | 3.4578 | 3.5712 | 3.7924 | 3.0879 | | 1.7528 | 2.4927 | 3.0559 | 3.7704 | 3.1286 | 2.5775 | H13 | 3.9825 | 3.2818 | 2.1289 | 1.0960 | 2.1500 | 2.7404 | 3.6106 | 4.5451 | 4.8582 | 4.1298 | 2.4873 | 1.7528 | | 3.0489 | 2.5060 | 3.7488 | 2.5495 | 3.0877 | H14 | 3.8372 | 3.4307 | 2.7466 | 2.1511 | 1.0937 | 2.1625 | 3.4548 | 4.8776 | 4.0503 | 4.2359 | 3.2418 | 2.4927 | 3.0489 | | 1.7520 | 2.5101 | 3.0732 | 2.5224 | H15 | 4.7687 | 3.8697 | 2.7109 | 2.1566 | 1.0943 | 2.1591 | 4.5381 | 5.6891 | 5.2048 | 4.5148 | 2.6511 | 3.0559 | 2.5060 | 1.7520 | | 2.5071 | 2.5171 | 3.0711 | H16 | 6.0371 | 5.6667 | 4.6792 | 3.4942 | 2.1778 | 1.0912 | 5.3942 | 6.9769 | 6.3925 | 6.5341 | 4.9796 | 3.7704 | 3.7488 | 2.5101 | 2.5071 | | 1.7648 | 1.7644 | H17 | 5.7891 | 5.3179 | 4.1718 | 2.7986 | 2.1699 | 1.0920 | 5.1338 | 6.5550 | 6.4227 | 6.2235 | 4.4419 | 3.1286 | 2.5495 | 3.0732 | 2.5171 | 1.7648 | | 1.7630 | H18 | 5.0470 | 5.0084 | 4.1978 | 2.7952 | 2.1700 | 1.0921 | 4.2020 | 5.8623 | 5.5255 | 6.0264 | 4.8228 | 2.5775 | 3.0877 | 2.5224 | 3.0711 | 1.7644 | 1.7630 | |
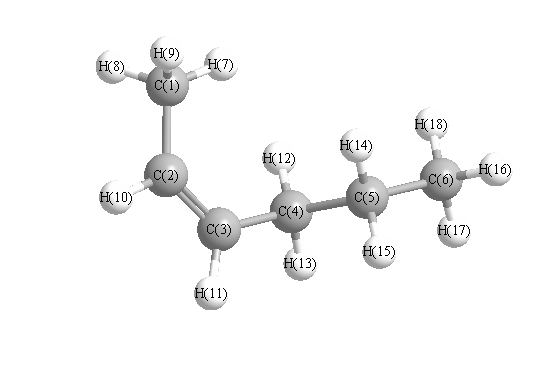 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.335 |
|
C1 |
C2 |
H10 |
115.285 |
| C2 |
C1 |
H7 |
112.762 |
|
C2 |
C1 |
H8 |
110.765 |
| C2 |
C1 |
H9 |
110.535 |
|
C2 |
C3 |
C4 |
127.542 |
| C2 |
C3 |
H11 |
117.369 |
|
C3 |
C2 |
H10 |
117.380 |
| C3 |
C4 |
C5 |
111.698 |
|
C3 |
C4 |
H12 |
111.400 |
| C3 |
C4 |
H13 |
109.310 |
|
C4 |
C3 |
H11 |
115.056 |
| C4 |
C5 |
C6 |
112.225 |
|
C4 |
C5 |
H14 |
108.788 |
| C4 |
C5 |
H15 |
109.178 |
|
C5 |
C4 |
H12 |
109.247 |
| C5 |
C4 |
H13 |
108.566 |
|
C5 |
C6 |
H16 |
111.564 |
| C5 |
C6 |
H17 |
110.883 |
|
C5 |
C6 |
H18 |
110.886 |
| C6 |
C5 |
H14 |
110.188 |
|
C6 |
C5 |
H15 |
109.881 |
| H7 |
C1 |
H8 |
107.992 |
|
H7 |
C1 |
H9 |
107.855 |
| H8 |
C1 |
H9 |
106.689 |
|
H12 |
C4 |
H13 |
106.445 |
| H14 |
C5 |
H16 |
94.356 |
|
H16 |
C6 |
H17 |
107.868 |
| H16 |
C6 |
H18 |
107.832 |
|
H17 |
C6 |
H18 |
107.645 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
