return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for Si3H8 (trisilane)

using model chemistry: MP2=FULL/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31G(2df,p)
 hartrees
Energy at 0K-871.826918
Energy at 298.15K-871.834674
HF Energy-871.414809
Nuclear repulsion energy193.610381
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 2311 2172 120.28      
2 A1 2300 2161 12.03      
3 A1 2281 2144 60.54      
4 A1 989 930 69.75      
5 A1 962 904 0.78      
6 A1 923 867 194.98      
7 A1 585 549 5.16      
8 A1 404 380 0.43      
9 A1 96 90 1.27      
10 A2 2308 2170 0.00      
11 A2 982 923 0.00      
12 A2 729 686 0.00      
13 A2 425 399 0.00      
14 A2 86 81 0.00      
15 B1 2313 2174 199.66      
16 B1 2291 2153 21.83      
17 B1 986 927 76.24      
18 B1 615 578 8.92      
19 B1 316 297 17.38      
20 B1 106 100 0.04      
21 B2 2309 2171 59.57      
22 B2 2296 2158 96.71      
23 B2 982 923 38.54      
24 B2 909 854 325.95      
25 B2 737 693 261.75      
26 B2 480 452 5.75      
27 B2 444 417 18.26      

Unscaled Zero Point Vibrational Energy (zpe) 15081.1 cm-1
Scaled (by 0.9399) Zero Point Vibrational Energy (zpe) 14174.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G(2df,p)
ABC
0.30395 0.06766 0.05869

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G(2df,p)

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Si1 0.000 0.000 0.911
Si2 0.000 1.917 -0.427
Si3 0.000 -1.917 -0.427
H4 1.198 0.000 1.778
H5 -1.198 0.000 1.778
H6 0.000 3.143 0.396
H7 0.000 -3.143 0.396
H8 1.199 1.922 -1.288
H9 -1.199 1.922 -1.288
H10 -1.199 -1.922 -1.288
H11 1.199 -1.922 -1.288

Atom - Atom Distances (Å)
  Si1 Si2 Si3 H4 H5 H6 H7 H8 H9 H10 H11
Si12.33762.33761.47911.47913.18473.18473.15743.15743.15743.1574
Si22.33763.83423.15783.15781.47645.12641.47651.47654.11354.1135
Si32.33763.83423.15783.15785.12641.47644.11354.11351.47651.4765
H41.47913.15783.15782.39673.63623.63623.61894.34104.34103.6189
H51.47913.15783.15782.39673.63623.63624.34103.61893.61894.3410
H63.18471.47645.12643.63623.63626.28572.40112.40115.47095.4709
H73.18475.12641.47643.63623.63626.28575.47095.47092.40112.4011
H83.15741.47654.11353.61894.34102.40115.47092.39804.53123.8446
H93.15741.47654.11354.34103.61892.40115.47092.39803.84464.5312
H103.15744.11351.47654.34103.61895.47092.40114.53123.84462.3980
H113.15744.11351.47653.61894.34105.47092.40113.84464.53122.3980

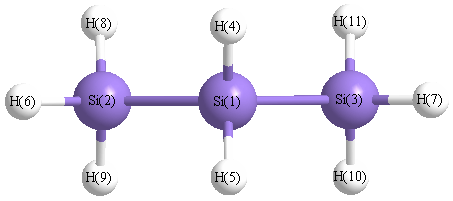
picture of trisilane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 H6 111.218 S1 S2 H8 109.683
S1 S2 H9 109.683 S1 S3 H7 111.218
S1 S3 H10 109.683 S1 S3 H11 109.683
S2 S1 S3 110.192 S2 S1 H4 109.597
S2 S1 H5 109.597 S3 S1 H4 109.597
S3 S1 H5 109.597 H4 S1 H5 108.229
H6 S2 H8 108.805 H6 S2 H9 108.805
H7 S3 H10 108.805 H7 S3 H11 108.805
H8 S2 H9 108.597 H10 S3 H11 108.597
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability