return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(CH3)CN (Propanenitrile, 2-methyl-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-210.838429
Energy at 298.15K-210.845713
HF Energy-210.016906
Nuclear repulsion energy159.076329
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3214 3036 9.92      
2 A' 3209 3031 18.13      
3 A' 3119 2946 3.59      
4 A' 3110 2937 8.04      
5 A' 2209 2086 0.05      
6 A' 1538 1453 9.52      
7 A' 1528 1444 7.05      
8 A' 1440 1360 3.26      
9 A' 1365 1289 6.03      
10 A' 1211 1144 2.39      
11 A' 1138 1075 4.61      
12 A' 956 903 1.12      
13 A' 789 745 0.44      
14 A' 546 516 1.04      
15 A' 353 333 0.21      
16 A' 295 278 0.65      
17 A' 222 210 2.59      
18 A" 3213 3034 6.64      
19 A" 3206 3028 0.09      
20 A" 3110 2937 9.64      
21 A" 1517 1432 2.95      
22 A" 1514 1430 0.58      
23 A" 1419 1340 3.90      
24 A" 1339 1265 0.39      
25 A" 1157 1093 1.81      
26 A" 992 937 0.46      
27 A" 946 893 2.19      
28 A" 563 532 0.00      
29 A" 236 223 0.01      
30 A" 185 174 4.54      

Unscaled Zero Point Vibrational Energy (zpe) 22818.4 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 21552.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.26670 0.13200 0.09694

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.150 -2.215 0.000
C2 -0.107 -1.070 0.000
C3 -0.410 0.362 0.000
C4 0.150 1.016 1.268
C5 0.150 1.016 -1.268
H6 -1.500 0.460 0.000
H7 -0.105 2.077 1.277
H8 -0.257 0.552 2.166
H9 1.237 0.922 1.295
H10 -0.105 2.077 -1.277
H11 -0.257 0.552 -2.166
H12 1.237 0.922 -1.295

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.17402.63753.47083.47083.14324.48513.53723.56304.48513.53723.5630
C21.17401.46352.45412.45412.06893.39562.70962.72873.39562.70962.7287
C32.63751.46351.53241.53241.09432.15962.17962.16802.15962.17962.1680
C43.47082.45411.53242.53552.15381.09101.09001.09102.76893.48892.7848
C53.47082.45411.53242.53552.15382.76893.48892.78481.09101.09001.0910
H63.14322.06891.09432.15382.15382.48822.49893.06242.48822.49893.0624
H74.48513.39562.15961.09102.76892.48821.77181.77042.55443.76893.1221
H83.53722.70962.17961.09003.48892.49891.77181.76853.76894.33203.7873
H93.56302.72872.16801.09102.78483.06241.77041.76853.12213.78732.5891
H104.48513.39562.15962.76891.09102.48822.55443.76893.12211.77181.7704
H113.53722.70962.17963.48891.09002.49893.76894.33203.78731.77181.7685
H123.56302.72872.16802.78481.09103.06243.12213.78732.58911.77041.7685

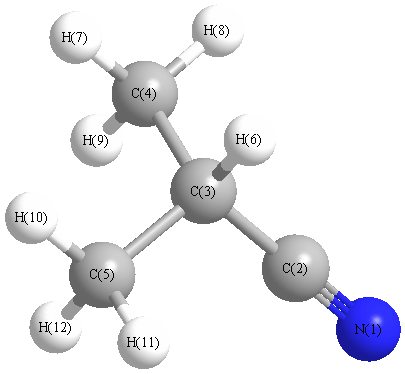
picture of Propanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 179.298 C2 C3 C4 109.976
C2 C3 C5 109.976 C2 C3 H6 107.079
C3 C4 H7 109.663 C3 C4 H8 111.315
C3 C4 H9 110.331 C3 C5 H10 109.663
C3 C5 H11 111.315 C3 C5 H12 110.331
C4 C3 C5 111.648 C4 C3 H6 109.021
C5 C3 H6 109.021 H7 C4 H8 108.654
H7 C4 H9 108.452 H8 C4 H9 108.357
H10 C5 H11 108.654 H10 C5 H12 108.452
H11 C5 H12 108.357
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability