Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -216.627791 |
| Energy at 298.15K | |
| HF Energy | -215.936417 |
| Nuclear repulsion energy | 116.992959 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3331 |
3146 |
1.82 |
41.52 |
0.72 |
0.84 |
| 2 |
A' |
3237 |
3057 |
4.42 |
136.03 |
0.14 |
0.25 |
| 3 |
A' |
3222 |
3044 |
2.64 |
21.81 |
0.65 |
0.79 |
| 4 |
A' |
3084 |
2913 |
31.45 |
127.83 |
0.10 |
0.18 |
| 5 |
A' |
1715 |
1620 |
2.32 |
5.30 |
0.08 |
0.14 |
| 6 |
A' |
1537 |
1452 |
2.47 |
13.76 |
0.61 |
0.75 |
| 7 |
A' |
1461 |
1380 |
11.97 |
10.34 |
0.54 |
0.70 |
| 8 |
A' |
1444 |
1364 |
8.76 |
4.23 |
0.74 |
0.85 |
| 9 |
A' |
1320 |
1247 |
0.19 |
12.40 |
0.37 |
0.53 |
| 10 |
A' |
1169 |
1104 |
50.22 |
1.88 |
0.66 |
0.80 |
| 11 |
A' |
1025 |
968 |
23.53 |
4.17 |
0.75 |
0.85 |
| 12 |
A' |
934 |
882 |
1.27 |
4.64 |
0.12 |
0.22 |
| 13 |
A' |
609 |
575 |
4.35 |
1.12 |
0.74 |
0.85 |
| 14 |
A' |
279 |
264 |
2.27 |
0.70 |
0.47 |
0.64 |
| 15 |
A" |
3133 |
2959 |
27.18 |
75.56 |
0.75 |
0.86 |
| 16 |
A" |
1293 |
1221 |
0.10 |
7.92 |
0.75 |
0.86 |
| 17 |
A" |
1063 |
1004 |
10.83 |
0.15 |
0.75 |
0.86 |
| 18 |
A" |
1042 |
985 |
10.22 |
0.01 |
0.75 |
0.86 |
| 19 |
A" |
959 |
906 |
32.71 |
0.14 |
0.75 |
0.86 |
| 20 |
A" |
566 |
535 |
7.42 |
7.87 |
0.75 |
0.86 |
| 21 |
A" |
178 |
168 |
2.40 |
3.73 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16300.7 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 15396.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.945 |
-0.201 |
0.000 |
| C2 |
0.000 |
0.956 |
0.000 |
| C3 |
1.325 |
0.819 |
0.000 |
| F4 |
-0.266 |
-1.396 |
0.000 |
| H5 |
1.979 |
1.680 |
0.000 |
| H6 |
1.779 |
-0.162 |
0.000 |
| H7 |
-0.462 |
1.939 |
0.000 |
| H8 |
-1.591 |
-0.170 |
0.884 |
| H9 |
-1.591 |
-0.170 |
-0.884 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4947 | 2.4894 | 1.3741 | 3.4776 | 2.7244 | 2.1937 | 1.0955 | 1.0955 |
C2 | 1.4947 | | 1.3323 | 2.3670 | 2.1074 | 2.1012 | 1.0856 | 2.1411 | 2.1411 | C3 | 2.4894 | 1.3323 | | 2.7269 | 1.0813 | 1.0811 | 2.1092 | 3.2043 | 3.2043 | F4 | 1.3741 | 2.3670 | 2.7269 | | 3.8080 | 2.3877 | 3.3400 | 2.0101 | 2.0101 | H5 | 3.4776 | 2.1074 | 1.0813 | 3.8080 | | 1.8533 | 2.4553 | 4.1178 | 4.1178 | H6 | 2.7244 | 2.1012 | 1.0811 | 2.3877 | 1.8533 | | 3.0718 | 3.4842 | 3.4842 | H7 | 2.1937 | 1.0856 | 2.1092 | 3.3400 | 2.4553 | 3.0718 | | 2.5503 | 2.5503 | H8 | 1.0955 | 2.1411 | 3.2043 | 2.0101 | 4.1178 | 3.4842 | 2.5503 | | 1.7684 | H9 | 1.0955 | 2.1411 | 3.2043 | 2.0101 | 4.1178 | 3.4842 | 2.5503 | 1.7684 | |
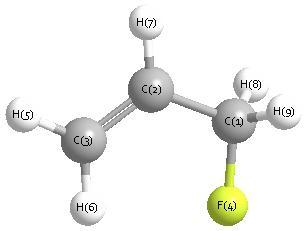 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.321 |
|
C1 |
C2 |
H7 |
115.551 |
| C2 |
C1 |
F4 |
111.121 |
|
C2 |
C1 |
H8 |
110.556 |
| C2 |
C1 |
H9 |
110.556 |
|
C2 |
C3 |
H5 |
121.303 |
| C2 |
C3 |
H6 |
120.717 |
|
C3 |
C2 |
H7 |
121.128 |
| F4 |
C1 |
H8 |
108.433 |
|
F4 |
C1 |
H9 |
108.433 |
| H5 |
C3 |
H6 |
117.980 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -216.626261 |
| Energy at 298.15K | |
| HF Energy | -215.935478 |
| Nuclear repulsion energy | 114.488300 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3311 |
3128 |
5.41 |
48.30 |
0.63 |
0.78 |
| 2 |
A |
3236 |
3056 |
3.38 |
95.59 |
0.23 |
0.38 |
| 3 |
A |
3211 |
3033 |
3.28 |
55.55 |
0.12 |
0.22 |
| 4 |
A |
3155 |
2980 |
24.86 |
55.45 |
0.75 |
0.86 |
| 5 |
A |
3094 |
2922 |
30.41 |
97.08 |
0.09 |
0.17 |
| 6 |
A |
1713 |
1618 |
0.35 |
5.66 |
0.09 |
0.16 |
| 7 |
A |
1539 |
1454 |
1.15 |
7.52 |
0.70 |
0.83 |
| 8 |
A |
1482 |
1400 |
21.67 |
8.31 |
0.56 |
0.72 |
| 9 |
A |
1421 |
1342 |
12.09 |
3.60 |
0.59 |
0.74 |
| 10 |
A |
1322 |
1249 |
0.11 |
11.14 |
0.48 |
0.64 |
| 11 |
A |
1295 |
1223 |
3.67 |
9.89 |
0.74 |
0.85 |
| 12 |
A |
1199 |
1132 |
2.91 |
1.92 |
0.64 |
0.78 |
| 13 |
A |
1106 |
1044 |
93.76 |
4.92 |
0.46 |
0.63 |
| 14 |
A |
1047 |
988 |
24.05 |
0.38 |
0.64 |
0.78 |
| 15 |
A |
1005 |
949 |
5.04 |
1.73 |
0.14 |
0.24 |
| 16 |
A |
971 |
917 |
31.51 |
0.15 |
0.47 |
0.64 |
| 17 |
A |
938 |
886 |
2.26 |
2.42 |
0.16 |
0.28 |
| 18 |
A |
665 |
628 |
5.65 |
4.62 |
0.64 |
0.78 |
| 19 |
A |
436 |
412 |
1.69 |
3.26 |
0.49 |
0.66 |
| 20 |
A |
328 |
310 |
5.93 |
3.04 |
0.75 |
0.86 |
| 21 |
A |
114 |
108 |
0.89 |
4.38 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16293.4 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 15389.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.611 |
0.428 |
0.317 |
| C2 |
0.640 |
-0.383 |
0.248 |
| C3 |
1.796 |
0.094 |
-0.214 |
| F4 |
-1.633 |
-0.224 |
-0.346 |
| H5 |
2.695 |
-0.508 |
-0.227 |
| H6 |
1.877 |
1.108 |
-0.589 |
| H7 |
0.568 |
-1.402 |
0.614 |
| H8 |
-0.929 |
0.572 |
1.354 |
| H9 |
-0.463 |
1.410 |
-0.142 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4928 | 2.4882 | 1.3817 | 3.4796 | 2.7334 | 2.1974 | 1.0946 | 1.0934 |
C2 | 1.4928 | | 1.3335 | 2.3545 | 2.1134 | 2.1097 | 1.0852 | 2.1443 | 2.1405 | C3 | 2.4882 | 1.3335 | | 3.4468 | 1.0819 | 1.0837 | 2.1054 | 3.1807 | 2.6157 | F4 | 1.3817 | 2.3545 | 3.4468 | | 4.3394 | 3.7619 | 2.6744 | 2.0052 | 2.0198 | H5 | 3.4796 | 2.1134 | 1.0819 | 4.3394 | | 1.8470 | 2.4563 | 4.0993 | 3.6960 | H6 | 2.7334 | 2.1097 | 1.0837 | 3.7619 | 1.8470 | | 3.0753 | 3.4545 | 2.4011 | H7 | 2.1974 | 1.0852 | 2.1054 | 2.6744 | 2.4563 | 3.0753 | | 2.5857 | 3.0886 | H8 | 1.0946 | 2.1443 | 3.1807 | 2.0052 | 4.0993 | 3.4545 | 2.5857 | | 1.7772 | H9 | 1.0934 | 2.1405 | 2.6157 | 2.0198 | 3.6960 | 2.4011 | 3.0886 | 1.7772 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.278 |
|
C1 |
C2 |
H7 |
116.042 |
| C2 |
C1 |
F4 |
109.933 |
|
C2 |
C1 |
H8 |
111.013 |
| C2 |
C1 |
H9 |
110.771 |
|
C2 |
C3 |
H5 |
121.732 |
| C2 |
C3 |
H6 |
121.222 |
|
C3 |
C2 |
H7 |
120.674 |
| F4 |
C1 |
H8 |
107.585 |
|
F4 |
C1 |
H9 |
108.821 |
| H5 |
C3 |
H6 |
117.045 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
