return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (2-methylbutyraldehyde)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-271.077078
Energy at 298.15K-271.087892
HF Energy-270.049967
Nuclear repulsion energy240.686770
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3217 3038 11.13      
2 A 3208 3030 14.39      
3 A 3206 3028 15.06      
4 A 3196 3019 17.76      
5 A 3144 2970 13.39      
6 A 3115 2942 9.83      
7 A 3107 2935 14.81      
8 A 3091 2919 17.85      
9 A 3067 2896 10.54      
10 A 2945 2782 101.36      
11 A 1788 1689 104.07      
12 A 1540 1455 3.17      
13 A 1531 1446 6.92      
14 A 1524 1439 8.18      
15 A 1520 1436 7.11      
16 A 1509 1425 0.52      
17 A 1447 1367 7.15      
18 A 1433 1353 2.30      
19 A 1421 1342 4.58      
20 A 1387 1310 5.77      
21 A 1377 1300 0.97      
22 A 1324 1251 1.51      
23 A 1285 1213 0.66      
24 A 1209 1142 2.17      
25 A 1187 1121 2.65      
26 A 1133 1070 2.88      
27 A 1068 1009 0.11      
28 A 1034 976 13.22      
29 A 985 931 6.83      
30 A 954 901 6.92      
31 A 929 878 12.09      
32 A 808 763 4.84      
33 A 791 747 2.75      
34 A 667 630 7.71      
35 A 408 385 0.78      
36 A 387 366 0.49      
37 A 301 284 3.68      
38 A 264 250 0.44      
39 A 243 229 3.37      
40 A 203 192 0.39      
41 A 98 93 1.92      
42 A 78 74 3.53      

Unscaled Zero Point Vibrational Energy (zpe) 31563.7 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 29812.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.23507 0.07081 0.05924

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.369 -0.689 0.229
C2 -0.081 0.074 0.422
C3 1.039 -0.697 -0.293
C4 2.430 -0.137 -0.010
C5 -0.224 1.516 -0.043
O6 -2.373 -0.241 -0.282
H7 -1.336 -1.742 0.582
H8 0.134 0.040 1.500
H9 1.004 -1.747 0.015
H10 0.843 -0.681 -1.370
H11 3.199 -0.749 -0.483
H12 2.539 0.880 -0.389
H13 2.629 -0.118 1.064
H14 -0.380 1.552 -1.122
H15 -1.087 1.988 0.426
H16 0.664 2.100 0.199

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.50952.46433.84692.49991.21281.11102.09912.60702.73004.62394.25604.12442.79772.69943.4518
C21.50951.53692.55731.52162.41822.21381.09952.15882.15403.50152.85852.79232.15842.16192.1702
C32.46431.53691.52632.56013.44282.73902.13961.09501.09512.16902.17772.16912.78563.49962.8649
C43.84692.55731.52633.12674.81244.13732.75392.15122.16041.09101.09011.09223.46254.13232.8582
C52.49991.52162.56013.12672.78633.49932.16453.48682.78024.12822.85573.46911.09151.09011.0897
O61.21282.41823.44284.81242.78632.01893.08873.71003.42415.59935.03905.18192.80902.66893.8645
H71.11102.21382.73904.13733.49932.01892.48672.40823.11254.76364.77844.31263.83013.74194.3489
H82.09911.09952.13962.75392.16453.08872.48672.48103.04353.73513.17092.53813.07002.53692.4930
H92.60702.15881.09502.15123.48683.71002.40822.48101.75552.46203.06862.52853.75414.30023.8666
H102.73002.15401.09512.16042.78023.42413.11253.04351.75552.51872.50473.07152.55863.75243.1992
H114.62393.50152.16901.09104.12825.59934.76363.73512.46202.51871.76041.76514.30345.16643.8750
H124.25602.85852.17771.09012.85575.03904.77843.17093.06862.50471.76041.76513.08373.87792.3134
H134.12442.79232.16911.09223.46915.18194.31262.53812.52853.07151.76511.76514.07764.31893.0875
H142.79772.15842.78563.46251.09152.80903.83013.07003.75412.55864.30343.08374.07761.75741.7714
H152.69942.16193.49964.13231.09012.66893.74192.53694.30023.75245.16643.87794.31891.75741.7688
H163.45182.17022.86492.85821.08973.86454.34892.49303.86663.19923.87502.31343.08751.77141.7688

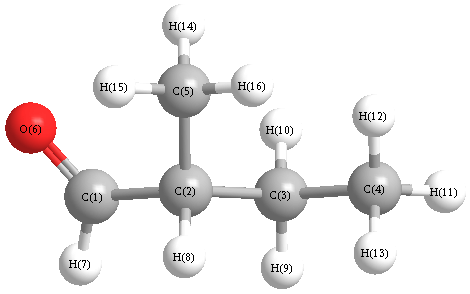
picture of 2-methylbutyraldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 107.979 C1 C2 C5 111.130
C1 C2 H8 106.076 C2 C1 O6 124.961
C2 C1 H7 114.448 C2 C3 C4 113.204
C2 C3 H9 109.063 C2 C3 H10 108.684
C2 C5 H14 110.293 C2 C5 H15 110.656
C2 C5 H16 111.348 C3 C2 C5 113.661
C3 C2 H8 107.337 C3 C4 H11 110.840
C3 C4 H12 111.588 C3 C4 H13 110.783
C4 C3 H9 109.194 C4 C3 H10 109.916
C5 C2 H8 110.303 O6 C1 H7 120.581
H9 C3 H10 106.561 H11 C4 H12 107.620
H11 C4 H13 107.896 H12 C4 H13 107.960
H14 C5 H15 107.332 H14 C5 H16 108.609
H15 C5 H16 108.484
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability