Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -342.568429 |
| Energy at 298.15K | -342.573259 |
| HF Energy | -341.383875 |
| Nuclear repulsion energy | 271.894560 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3336 |
3151 |
0.37 |
145.35 |
0.11 |
0.21 |
| 2 |
A' |
3316 |
3132 |
1.00 |
35.69 |
0.58 |
0.74 |
| 3 |
A' |
3306 |
3122 |
0.55 |
73.59 |
0.61 |
0.76 |
| 4 |
A' |
2980 |
2814 |
88.79 |
111.60 |
0.29 |
0.45 |
| 5 |
A' |
1766 |
1668 |
190.40 |
120.92 |
0.29 |
0.45 |
| 6 |
A' |
1590 |
1501 |
6.85 |
0.38 |
0.74 |
0.85 |
| 7 |
A' |
1514 |
1430 |
30.73 |
66.68 |
0.33 |
0.49 |
| 8 |
A' |
1445 |
1365 |
26.35 |
42.91 |
0.32 |
0.49 |
| 9 |
A' |
1409 |
1331 |
0.74 |
14.55 |
0.12 |
0.22 |
| 10 |
A' |
1344 |
1269 |
18.59 |
5.90 |
0.34 |
0.51 |
| 11 |
A' |
1258 |
1188 |
14.60 |
4.72 |
0.52 |
0.68 |
| 12 |
A' |
1223 |
1155 |
12.35 |
5.27 |
0.15 |
0.26 |
| 13 |
A' |
1114 |
1052 |
6.41 |
7.87 |
0.33 |
0.49 |
| 14 |
A' |
1055 |
996 |
33.92 |
3.76 |
0.34 |
0.51 |
| 15 |
A' |
955 |
902 |
15.28 |
4.38 |
0.10 |
0.18 |
| 16 |
A' |
893 |
843 |
8.17 |
5.91 |
0.71 |
0.83 |
| 17 |
A' |
770 |
727 |
60.61 |
1.50 |
0.75 |
0.85 |
| 18 |
A' |
499 |
471 |
1.03 |
6.07 |
0.27 |
0.43 |
| 19 |
A' |
195 |
184 |
5.15 |
0.84 |
0.62 |
0.77 |
| 20 |
A" |
1020 |
964 |
0.01 |
3.74 |
0.75 |
0.86 |
| 21 |
A" |
860 |
812 |
0.15 |
0.43 |
0.75 |
0.86 |
| 22 |
A" |
814 |
768 |
14.62 |
0.37 |
0.75 |
0.86 |
| 23 |
A" |
777 |
733 |
48.47 |
0.40 |
0.75 |
0.86 |
| 24 |
A" |
662 |
625 |
1.37 |
2.09 |
0.75 |
0.86 |
| 25 |
A" |
608 |
575 |
4.09 |
0.09 |
0.75 |
0.86 |
| 26 |
A" |
291 |
275 |
10.19 |
0.43 |
0.75 |
0.86 |
| 27 |
A" |
134 |
127 |
1.52 |
1.50 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17565.4 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 16590.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.500 |
-0.892 |
0.000 |
| C2 |
-0.558 |
-1.730 |
0.000 |
| C3 |
-1.739 |
-1.033 |
0.000 |
| C4 |
0.000 |
0.369 |
0.000 |
| C5 |
-1.378 |
0.336 |
0.000 |
| C6 |
0.929 |
1.488 |
0.000 |
| O7 |
2.142 |
1.390 |
0.000 |
| H8 |
-0.318 |
-2.780 |
0.000 |
| H9 |
-2.729 |
-1.458 |
0.000 |
| H10 |
-2.030 |
1.195 |
0.000 |
| H11 |
0.419 |
2.472 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3499 | 2.2433 | 1.3560 | 2.2432 | 2.4184 | 2.8108 | 2.0578 | 3.2781 | 3.2792 | 3.3641 |
C2 | 1.3499 | | 1.3709 | 2.1718 | 2.2224 | 3.5453 | 4.1258 | 1.0773 | 2.1875 | 3.2742 | 4.3135 | C3 | 2.2433 | 1.3709 | | 2.2340 | 1.4163 | 3.6714 | 4.5752 | 2.2518 | 1.0773 | 2.2473 | 4.1160 | C4 | 1.3560 | 2.1718 | 2.2340 | | 1.3780 | 1.4549 | 2.3726 | 3.1649 | 3.2843 | 2.1914 | 2.1439 | C5 | 2.2432 | 2.2224 | 1.4163 | 1.3780 | | 2.5787 | 3.6737 | 3.2913 | 2.2464 | 1.0785 | 2.7908 | C6 | 2.4184 | 3.5453 | 3.6714 | 1.4549 | 2.5787 | | 1.2164 | 4.4468 | 4.6975 | 2.9735 | 1.1077 | O7 | 2.8108 | 4.1258 | 4.5752 | 2.3726 | 3.6737 | 1.2164 | | 4.8411 | 5.6424 | 4.1760 | 2.0342 | H8 | 2.0578 | 1.0773 | 2.2518 | 3.1649 | 3.2913 | 4.4468 | 4.8411 | | 2.7494 | 4.3280 | 5.3030 | H9 | 3.2781 | 2.1875 | 1.0773 | 3.2843 | 2.2464 | 4.6975 | 5.6424 | 2.7494 | | 2.7442 | 5.0354 | H10 | 3.2792 | 3.2742 | 2.2473 | 2.1914 | 1.0785 | 2.9735 | 4.1760 | 4.3280 | 2.7442 | | 2.7615 | H11 | 3.3641 | 4.3135 | 4.1160 | 2.1439 | 2.7908 | 1.1077 | 2.0342 | 5.3030 | 5.0354 | 2.7615 | |
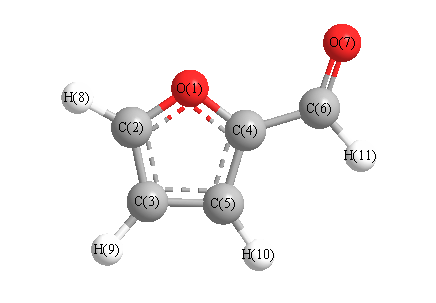 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.079 |
|
O1 |
C2 |
H8 |
115.492 |
| O1 |
C4 |
C5 |
110.259 |
|
O1 |
C4 |
C6 |
118.673 |
| C2 |
O1 |
C4 |
106.759 |
|
C2 |
C3 |
C5 |
105.749 |
| C2 |
C3 |
H9 |
126.220 |
|
C3 |
C2 |
H8 |
133.429 |
| C3 |
C5 |
C4 |
106.155 |
|
C3 |
C5 |
H10 |
128.013 |
| C4 |
C5 |
H10 |
125.832 |
|
C4 |
C6 |
O7 |
125.051 |
| C4 |
C6 |
H11 |
112.872 |
|
C5 |
C3 |
H9 |
128.031 |
| C5 |
C4 |
C6 |
131.068 |
|
O7 |
C6 |
H11 |
122.077 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -342.569771 |
| Energy at 298.15K | -342.574589 |
| HF Energy | -341.385331 |
| Nuclear repulsion energy | 271.052508 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3336 |
3151 |
0.35 |
152.21 |
0.12 |
0.21 |
| 2 |
A' |
3322 |
3137 |
2.59 |
20.02 |
0.42 |
0.59 |
| 3 |
A' |
3310 |
3126 |
0.15 |
77.14 |
0.75 |
0.86 |
| 4 |
A' |
2998 |
2832 |
80.07 |
118.20 |
0.29 |
0.45 |
| 5 |
A' |
1759 |
1661 |
163.51 |
109.49 |
0.31 |
0.48 |
| 6 |
A' |
1600 |
1511 |
41.18 |
14.32 |
0.30 |
0.46 |
| 7 |
A' |
1508 |
1424 |
60.66 |
131.80 |
0.29 |
0.45 |
| 8 |
A' |
1459 |
1378 |
2.29 |
4.06 |
0.71 |
0.83 |
| 9 |
A' |
1407 |
1329 |
10.92 |
26.20 |
0.40 |
0.57 |
| 10 |
A' |
1297 |
1225 |
7.42 |
6.99 |
0.14 |
0.25 |
| 11 |
A' |
1271 |
1201 |
25.84 |
2.68 |
0.73 |
0.84 |
| 12 |
A' |
1207 |
1140 |
7.32 |
4.03 |
0.12 |
0.22 |
| 13 |
A' |
1118 |
1056 |
13.82 |
10.23 |
0.33 |
0.50 |
| 14 |
A' |
1046 |
988 |
34.25 |
1.90 |
0.23 |
0.38 |
| 15 |
A' |
967 |
913 |
7.14 |
4.20 |
0.12 |
0.21 |
| 16 |
A' |
893 |
844 |
4.89 |
4.78 |
0.75 |
0.86 |
| 17 |
A' |
757 |
715 |
63.60 |
2.22 |
0.74 |
0.85 |
| 18 |
A' |
498 |
470 |
0.54 |
5.26 |
0.32 |
0.48 |
| 19 |
A' |
196 |
185 |
6.90 |
0.28 |
0.35 |
0.52 |
| 20 |
A" |
1030 |
972 |
0.04 |
4.44 |
0.75 |
0.86 |
| 21 |
A" |
872 |
823 |
0.75 |
0.46 |
0.75 |
0.86 |
| 22 |
A" |
824 |
778 |
20.23 |
0.59 |
0.75 |
0.86 |
| 23 |
A" |
767 |
725 |
41.36 |
0.38 |
0.75 |
0.86 |
| 24 |
A" |
654 |
618 |
1.40 |
0.94 |
0.75 |
0.86 |
| 25 |
A" |
611 |
577 |
5.53 |
0.24 |
0.75 |
0.86 |
| 26 |
A" |
256 |
241 |
10.74 |
1.41 |
0.75 |
0.86 |
| 27 |
A" |
157 |
148 |
2.80 |
1.19 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17559.1 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 16584.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.249 |
-0.254 |
0.000 |
| C2 |
1.095 |
-1.595 |
0.000 |
| C3 |
-0.234 |
-1.933 |
0.000 |
| C4 |
0.000 |
0.285 |
0.000 |
| C5 |
-0.949 |
-0.712 |
0.000 |
| C6 |
-0.090 |
1.737 |
0.000 |
| O7 |
-1.146 |
2.343 |
0.000 |
| H8 |
2.007 |
-2.169 |
0.000 |
| H9 |
-0.634 |
-2.934 |
0.000 |
| H10 |
-2.015 |
-0.554 |
0.000 |
| H11 |
0.889 |
2.254 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3505 | 2.2408 | 1.3600 | 2.2455 | 2.3988 | 3.5325 | 2.0597 | 3.2755 | 3.2782 | 2.5330 |
C2 | 1.3505 | | 1.3719 | 2.1758 | 2.2274 | 3.5367 | 4.5315 | 1.0771 | 2.1866 | 3.2806 | 3.8545 | C3 | 2.2408 | 1.3719 | | 2.2302 | 1.4152 | 3.6728 | 4.3722 | 2.2539 | 1.0775 | 2.2528 | 4.3347 | C4 | 1.3600 | 2.1758 | 2.2302 | | 1.3763 | 1.4550 | 2.3559 | 3.1698 | 3.2804 | 2.1828 | 2.1602 | C5 | 2.2455 | 2.2274 | 1.4152 | 1.3763 | | 2.5951 | 3.0610 | 3.2960 | 2.2445 | 1.0778 | 3.4887 | C6 | 2.3988 | 3.5367 | 3.6728 | 1.4550 | 2.5951 | | 1.2179 | 4.4329 | 4.7024 | 2.9924 | 1.1064 | O7 | 3.5325 | 4.5315 | 4.3722 | 2.3559 | 3.0610 | 1.2179 | | 5.5044 | 5.3017 | 3.0242 | 2.0366 | H8 | 2.0597 | 1.0771 | 2.2539 | 3.1698 | 3.2960 | 4.4329 | 5.5044 | | 2.7495 | 4.3347 | 4.5616 | H9 | 3.2755 | 2.1866 | 1.0775 | 3.2804 | 2.2445 | 4.7024 | 5.3017 | 2.7495 | | 2.7523 | 5.4063 | H10 | 3.2782 | 3.2806 | 2.2528 | 2.1828 | 1.0778 | 2.9924 | 3.0242 | 4.3347 | 2.7523 | | 4.0390 | H11 | 2.5330 | 3.8545 | 4.3347 | 2.1602 | 3.4887 | 1.1064 | 2.0366 | 4.5616 | 5.4063 | 4.0390 | |
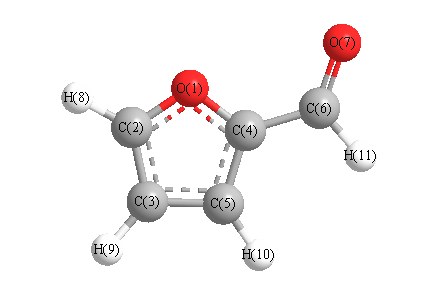 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
110.785 |
|
O1 |
C2 |
H8 |
115.627 |
| O1 |
C4 |
C5 |
110.300 |
|
O1 |
C4 |
C6 |
116.852 |
| C2 |
O1 |
C4 |
106.781 |
|
C2 |
C3 |
C5 |
106.090 |
| C2 |
C3 |
H9 |
126.005 |
|
C3 |
C2 |
H8 |
133.588 |
| C3 |
C5 |
C4 |
106.043 |
|
C3 |
C5 |
H10 |
128.781 |
| C4 |
C5 |
H10 |
125.175 |
|
C4 |
C6 |
O7 |
123.384 |
| C4 |
C6 |
H11 |
114.305 |
|
C5 |
C3 |
H9 |
127.905 |
| C5 |
C4 |
C6 |
132.848 |
|
O7 |
C6 |
H11 |
122.311 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
