return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3COC6H5 (acetophenone)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-383.939693
Energy at 298.15K-383.948012
HF Energy-382.510881
Nuclear repulsion energy401.698694
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3259 3078 6.03      
2 A' 3255 3075 4.84      
3 A' 3247 3067 6.04      
4 A' 3239 3059 3.79      
5 A' 3233 3053 6.10      
6 A' 3228 3049 0.19      
7 A' 3107 2935 1.05      
8 A' 1761 1663 98.05      
9 A' 1643 1552 9.31      
10 A' 1627 1537 5.58      
11 A' 1523 1438 1.44      
12 A' 1495 1412 8.24      
13 A' 1479 1397 15.73      
14 A' 1469 1387 5.09      
15 A' 1398 1321 59.35      
16 A' 1335 1261 7.07      
17 A' 1305 1233 148.28      
18 A' 1207 1140 8.37      
19 A' 1189 1123 0.49      
20 A' 1111 1049 5.30      
21 A' 1100 1039 0.92      
22 A' 1049 991 4.48      
23 A' 1007 951 0.73      
24 A' 968 914 19.65      
25 A' 740 699 2.00      
26 A' 615 581 0.63      
27 A' 589 557 24.63      
28 A' 460 434 0.39      
29 A' 364 344 0.77      
30 A' 219 207 4.62      
31 A" 3196 3019 2.55      
32 A" 1504 1420 9.18      
33 A" 1048 990 1.02      
34 A" 985 930 0.44      
35 A" 962 909 0.42      
36 A" 930 878 1.44      
37 A" 871 823 0.17      
38 A" 771 729 43.49      
39 A" 691 653 13.77      
40 A" 596 563 8.17      
41 A" 424 401 0.08      
42 A" 407 384 0.00      
43 A" 197 187 0.24      
44 A" 154 145 0.01      
45 A" 59 56 2.78      

Unscaled Zero Point Vibrational Energy (zpe) 30508.6 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 28815.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.12298 0.04078 0.03080

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.658 2.191 0.000
C2 0.230 1.691 0.000
C3 0.000 0.213 0.000
O4 -0.710 2.468 0.000
C5 1.049 -0.713 0.000
C6 -1.323 -0.243 0.000
C7 0.774 -2.080 0.000
C8 -1.595 -1.607 0.000
C9 -0.546 -2.528 0.000
H10 1.646 3.279 0.000
H11 2.192 1.829 0.881
H12 2.192 1.829 -0.881
H13 2.078 -0.377 0.000
H14 -2.118 0.494 0.000
H15 1.590 -2.793 0.000
H16 -2.621 -1.954 0.000
H17 -0.757 -3.592 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17
C11.51292.58072.38452.96733.84884.36074.99975.20801.08801.09191.09192.60214.13944.98435.95746.2661
C21.51291.49571.22012.53972.48073.80933.76844.28972.12742.15522.15522.77382.63524.68564.62765.3736
C32.58071.49572.36451.39921.39992.41952.41932.79493.47952.86262.86262.16072.13623.40063.40113.8789
O42.38451.22012.36453.63552.78004.78384.16964.99912.49163.09973.09973.98432.42465.74224.81756.0599
C52.96732.53971.39923.63552.41821.39342.79012.41574.03642.92392.92391.08333.38862.14903.87403.3975
C63.84882.48071.39992.78002.41822.78761.38992.41364.60664.17504.17503.40461.08353.87152.14733.3958
C74.36073.80932.41954.78381.39342.78762.41541.39405.42864.25054.25052.14463.87081.08393.39772.1516
C84.99973.76842.41934.16962.79011.38992.41541.39625.86215.18845.18843.87332.16443.39831.08402.1546
C95.20804.28972.79494.99912.41572.41361.39401.39626.20665.22115.22113.39313.40632.15202.15351.0841
H101.08802.12743.47952.49164.03644.60665.42865.86216.20661.78191.78193.68144.68176.07206.75207.2781
H111.09192.15522.86263.09972.92394.17504.25055.18845.22111.78191.76242.37884.59714.74426.18546.2335
H121.09192.15522.86263.09972.92394.17504.25055.18845.22111.78191.76242.37884.59714.74426.18546.2335
H132.60212.77382.16073.98431.08333.40462.14463.87333.39313.68142.37882.37884.28562.46484.95734.2860
H144.13942.63522.13622.42463.38861.08353.87082.16443.40634.68174.59714.59714.28564.95472.49914.3060
H154.98434.68563.40065.74222.14903.87151.08393.39832.15206.07204.74424.74422.46484.95474.29402.4786
H165.95744.62763.40114.81753.87402.14733.39771.08402.15356.75206.18546.18544.95732.49914.29402.4816
H176.26615.37363.87896.05993.39753.39582.15162.15461.08417.27816.23356.23354.28604.30602.47862.4816

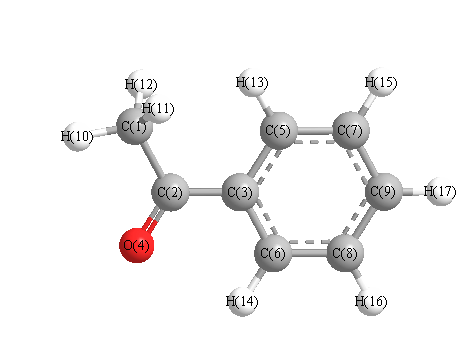
picture of acetophenone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 118.138 C1 C2 O4 121.125
C2 C1 H10 108.653 C2 C1 H11 110.629
C2 C1 H12 110.629 C2 C3 C5 122.609
C2 C3 C6 117.866 C3 C2 O4 120.737
C3 C5 C7 120.087 C3 C5 H13 120.469
C3 C6 C8 120.269 C3 C6 H14 118.115
C5 C3 C6 119.525 C5 C7 C9 120.144
C5 C7 H15 119.812 C6 C8 C9 120.059
C6 C8 H16 119.949 C7 C5 H13 119.444
C7 C9 C8 119.916 C7 C9 H17 119.996
C8 C6 H14 121.616 C8 C9 H17 120.088
C9 C7 H15 120.044 C9 C8 H16 119.993
H10 C1 H11 109.651 H10 C1 H12 109.651
H11 C1 H12 107.617
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability