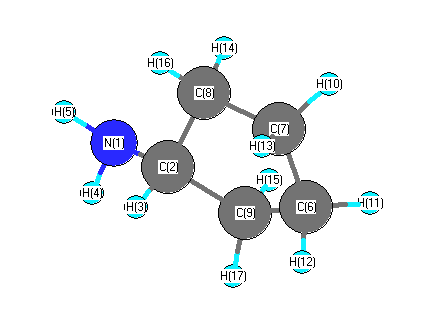
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3379 |
3261 |
6.29 |
|
|
|
| 2 |
A |
3135 |
3025 |
69.68 |
|
|
|
| 3 |
A |
3113 |
3003 |
7.94 |
|
|
|
| 4 |
A |
3084 |
2976 |
22.64 |
|
|
|
| 5 |
A |
3068 |
2960 |
28.62 |
|
|
|
| 6 |
A |
2924 |
2821 |
73.83 |
|
|
|
| 7 |
A |
1716 |
1656 |
17.12 |
|
|
|
| 8 |
A |
1575 |
1520 |
2.35 |
|
|
|
| 9 |
A |
1557 |
1502 |
5.95 |
|
|
|
| 10 |
A |
1412 |
1363 |
27.46 |
|
|
|
| 11 |
A |
1355 |
1307 |
0.31 |
|
|
|
| 12 |
A |
1341 |
1294 |
0.88 |
|
|
|
| 13 |
A |
1267 |
1222 |
1.77 |
|
|
|
| 14 |
A |
1219 |
1176 |
0.08 |
|
|
|
| 15 |
A |
1110 |
1071 |
4.60 |
|
|
|
| 16 |
A |
1040 |
1004 |
0.17 |
|
|
|
| 17 |
A |
990 |
955 |
3.01 |
|
|
|
| 18 |
A |
890 |
859 |
2.16 |
|
|
|
| 19 |
A |
865 |
835 |
0.15 |
|
|
|
| 20 |
A |
772 |
745 |
6.39 |
|
|
|
| 21 |
A |
744 |
718 |
167.24 |
|
|
|
| 22 |
A |
544 |
525 |
3.21 |
|
|
|
| 23 |
A |
462 |
446 |
15.19 |
|
|
|
| 24 |
A |
182 |
176 |
0.95 |
|
|
|
| 25 |
A |
3470 |
3348 |
1.16 |
|
|
|
| 26 |
A |
3119 |
3010 |
6.85 |
|
|
|
| 27 |
A |
3100 |
2991 |
19.63 |
|
|
|
| 28 |
A |
3070 |
2962 |
43.09 |
|
|
|
| 29 |
A |
3064 |
2956 |
15.05 |
|
|
|
| 30 |
A |
1550 |
1495 |
3.65 |
|
|
|
| 31 |
A |
1542 |
1488 |
0.69 |
|
|
|
| 32 |
A |
1412 |
1363 |
1.01 |
|
|
|
| 33 |
A |
1363 |
1315 |
0.52 |
|
|
|
| 34 |
A |
1334 |
1287 |
0.77 |
|
|
|
| 35 |
A |
1310 |
1264 |
2.71 |
|
|
|
| 36 |
A |
1245 |
1201 |
0.07 |
|
|
|
| 37 |
A |
1220 |
1177 |
1.58 |
|
|
|
| 38 |
A |
1036 |
999 |
0.16 |
|
|
|
| 39 |
A |
1007 |
972 |
0.19 |
|
|
|
| 40 |
A |
931 |
898 |
3.46 |
|
|
|
| 41 |
A |
831 |
802 |
0.00 |
|
|
|
| 42 |
A |
646 |
623 |
0.09 |
|
|
|
| 43 |
A |
350 |
338 |
13.62 |
|
|
|
| 44 |
A |
241 |
232 |
41.51 |
|
|
|
| 45 |
A |
49 |
47 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34816.5 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 33594.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.653 |
|
|
-1.049 |
| 2 |
C |
-0.060 |
|
|
0.405 |
| 3 |
H |
0.156 |
|
|
-0.030 |
| 4 |
H |
0.258 |
|
|
0.382 |
| 5 |
H |
0.258 |
|
|
0.382 |
| 6 |
C |
-0.379 |
|
|
-0.137 |
| 7 |
C |
-0.379 |
|
|
-0.137 |
| 8 |
C |
-0.360 |
|
|
-0.152 |
| 9 |
C |
-0.360 |
|
|
-0.152 |
| 10 |
H |
0.190 |
|
|
0.063 |
| 11 |
H |
0.190 |
|
|
0.063 |
| 12 |
H |
0.188 |
|
|
0.056 |
| 13 |
H |
0.188 |
|
|
0.056 |
| 14 |
H |
0.205 |
|
|
0.082 |
| 15 |
H |
0.205 |
|
|
0.082 |
| 16 |
H |
0.175 |
|
|
0.044 |
| 17 |
H |
0.175 |
|
|
0.044 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.976 |
0.873 |
0.000 |
1.310 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.969 |
0.874 |
0.000 |
1.305 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.854 |
0.751 |
0.000 |
| y |
0.751 |
-34.489 |
0.000 |
| z |
0.000 |
0.000 |
-37.786 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.716 |
0.751 |
0.000 |
| y |
0.751 |
5.331 |
0.000 |
| z |
0.000 |
0.000 |
0.385 |
|
| Polar |
|---|
| 3z2-r2 | 0.770 |
|---|
| x2-y2 | -7.364 |
|---|
| xy | 0.751 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.350 |
-0.758 |
0.000 |
| y |
-0.758 |
8.508 |
0.000 |
| z |
0.000 |
0.000 |
8.186 |
<r2> (average value of r
2) Å
2
| <r2> |
173.335 |
| (<r2>)1/2 |
13.166 |