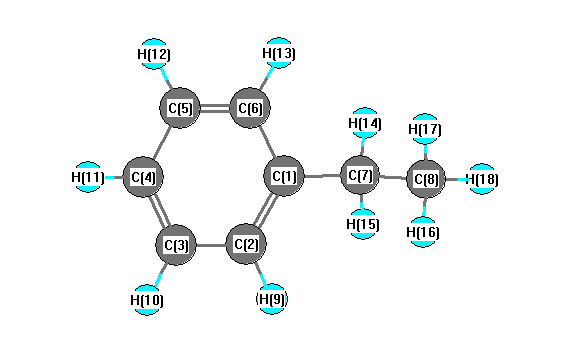
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3217 |
3104 |
13.99 |
|
|
|
| 2 |
A' |
3194 |
3081 |
5.89 |
|
|
|
| 3 |
A' |
3179 |
3068 |
7.78 |
|
|
|
| 4 |
A' |
3117 |
3007 |
39.63 |
|
|
|
| 5 |
A' |
3055 |
2948 |
21.48 |
|
|
|
| 6 |
A' |
3051 |
2944 |
28.81 |
|
|
|
| 7 |
A' |
1634 |
1577 |
4.36 |
|
|
|
| 8 |
A' |
1571 |
1515 |
5.59 |
|
|
|
| 9 |
A' |
1554 |
1500 |
8.07 |
|
|
|
| 10 |
A' |
1544 |
1490 |
8.24 |
|
|
|
| 11 |
A' |
1459 |
1408 |
3.73 |
|
|
|
| 12 |
A' |
1370 |
1322 |
1.01 |
|
|
|
| 13 |
A' |
1241 |
1197 |
0.09 |
|
|
|
| 14 |
A' |
1234 |
1191 |
0.34 |
|
|
|
| 15 |
A' |
1104 |
1065 |
13.99 |
|
|
|
| 16 |
A' |
1065 |
1028 |
2.29 |
|
|
|
| 17 |
A' |
1038 |
1002 |
0.57 |
|
|
|
| 18 |
A' |
1032 |
996 |
0.13 |
|
|
|
| 19 |
A' |
962 |
928 |
0.78 |
|
|
|
| 20 |
A' |
940 |
907 |
1.51 |
|
|
|
| 21 |
A' |
798 |
770 |
12.67 |
|
|
|
| 22 |
A' |
782 |
755 |
10.81 |
|
|
|
| 23 |
A' |
734 |
708 |
51.51 |
|
|
|
| 24 |
A' |
581 |
561 |
8.12 |
|
|
|
| 25 |
A' |
511 |
493 |
5.86 |
|
|
|
| 26 |
A' |
318 |
307 |
0.22 |
|
|
|
| 27 |
A' |
136 |
131 |
0.90 |
|
|
|
| 28 |
A" |
3202 |
3090 |
30.45 |
|
|
|
| 29 |
A" |
3180 |
3069 |
3.84 |
|
|
|
| 30 |
A" |
3128 |
3018 |
27.80 |
|
|
|
| 31 |
A" |
3090 |
2981 |
4.24 |
|
|
|
| 32 |
A" |
1618 |
1561 |
1.25 |
|
|
|
| 33 |
A" |
1564 |
1509 |
9.85 |
|
|
|
| 34 |
A" |
1512 |
1459 |
8.19 |
|
|
|
| 35 |
A" |
1411 |
1361 |
0.74 |
|
|
|
| 36 |
A" |
1338 |
1291 |
0.01 |
|
|
|
| 37 |
A" |
1291 |
1246 |
0.33 |
|
|
|
| 38 |
A" |
1230 |
1187 |
0.15 |
|
|
|
| 39 |
A" |
1146 |
1106 |
1.66 |
|
|
|
| 40 |
A" |
1076 |
1038 |
3.33 |
|
|
|
| 41 |
A" |
1002 |
967 |
0.00 |
|
|
|
| 42 |
A" |
871 |
840 |
0.01 |
|
|
|
| 43 |
A" |
811 |
783 |
1.37 |
|
|
|
| 44 |
A" |
661 |
638 |
0.07 |
|
|
|
| 45 |
A" |
424 |
410 |
0.00 |
|
|
|
| 46 |
A" |
358 |
345 |
0.12 |
|
|
|
| 47 |
A" |
227 |
219 |
0.02 |
|
|
|
| 48 |
A" |
50 |
48 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34803.3 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 33581.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.038 |
|
|
|
| 2 |
C |
-0.190 |
|
|
|
| 3 |
C |
-0.183 |
|
|
|
| 4 |
C |
-0.189 |
|
|
|
| 5 |
C |
-0.183 |
|
|
|
| 6 |
C |
-0.190 |
|
|
|
| 7 |
C |
-0.439 |
|
|
|
| 8 |
C |
-0.537 |
|
|
|
| 9 |
H |
0.178 |
|
|
|
| 10 |
H |
0.184 |
|
|
|
| 11 |
H |
0.183 |
|
|
|
| 12 |
H |
0.184 |
|
|
|
| 13 |
H |
0.178 |
|
|
|
| 14 |
H |
0.197 |
|
|
|
| 15 |
H |
0.197 |
|
|
|
| 16 |
H |
0.194 |
|
|
|
| 17 |
H |
0.194 |
|
|
|
| 18 |
H |
0.185 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.098 |
0.311 |
0.000 |
0.326 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-52.982 |
0.035 |
0.000 |
| y |
0.035 |
-44.431 |
0.000 |
| z |
0.000 |
0.000 |
-44.486 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.523 |
0.035 |
0.000 |
| y |
0.035 |
4.302 |
0.000 |
| z |
0.000 |
0.000 |
4.221 |
|
| Polar |
|---|
| 3z2-r2 | 8.442 |
|---|
| x2-y2 | -8.550 |
|---|
| xy | 0.035 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.348 |
0.644 |
0.000 |
| y |
0.644 |
14.764 |
0.000 |
| z |
0.000 |
0.000 |
12.150 |
<r2> (average value of r
2) Å
2
| <r2> |
290.832 |
| (<r2>)1/2 |
17.054 |