Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -5201.804805 |
| Energy at 298.15K | -5201.814967 |
| HF Energy | -5201.804805 |
| Nuclear repulsion energy | 409.619057 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3144 |
3033 |
0.00 |
|
|
|
| 2 |
Ag |
1535 |
1481 |
0.00 |
|
|
|
| 3 |
Ag |
1318 |
1272 |
0.00 |
|
|
|
| 4 |
Ag |
1052 |
1015 |
0.00 |
|
|
|
| 5 |
Ag |
663 |
640 |
0.00 |
|
|
|
| 6 |
Ag |
187 |
180 |
0.00 |
|
|
|
| 7 |
Au |
3230 |
3116 |
2.62 |
|
|
|
| 8 |
Au |
1143 |
1103 |
2.35 |
|
|
|
| 9 |
Au |
779 |
751 |
8.71 |
|
|
|
| 10 |
Au |
105 |
101 |
4.04 |
|
|
|
| 11 |
Bg |
3204 |
3092 |
0.00 |
|
|
|
| 12 |
Bg |
1338 |
1291 |
0.00 |
|
|
|
| 13 |
Bg |
937 |
904 |
0.00 |
|
|
|
| 14 |
Bu |
3153 |
3042 |
6.83 |
|
|
|
| 15 |
Bu |
1536 |
1482 |
15.38 |
|
|
|
| 16 |
Bu |
1255 |
1211 |
50.50 |
|
|
|
| 17 |
Bu |
598 |
577 |
71.40 |
|
|
|
| 18 |
Bu |
181 |
175 |
7.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12678.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 12233.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.502 |
0.564 |
0.000 |
| C2 |
-0.502 |
-0.564 |
0.000 |
| Br3 |
-0.502 |
2.309 |
0.000 |
| Br4 |
0.502 |
-2.309 |
0.000 |
| H5 |
1.119 |
0.567 |
0.895 |
| H6 |
1.119 |
0.567 |
-0.895 |
| H7 |
-1.119 |
-0.567 |
0.895 |
| H8 |
-1.119 |
-0.567 |
-0.895 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5102 | 2.0134 | 2.8734 | 1.0875 | 1.0875 | 2.1699 | 2.1699 |
C2 | 1.5102 | | 2.8734 | 2.0134 | 2.1699 | 2.1699 | 1.0875 | 1.0875 | Br3 | 2.0134 | 2.8734 | | 4.7265 | 2.5430 | 2.5430 | 3.0747 | 3.0747 | Br4 | 2.8734 | 2.0134 | 4.7265 | | 3.0747 | 3.0747 | 2.5430 | 2.5430 | H5 | 1.0875 | 2.1699 | 2.5430 | 3.0747 | | 1.7906 | 2.5090 | 3.0824 | H6 | 1.0875 | 2.1699 | 2.5430 | 3.0747 | 1.7906 | | 3.0824 | 2.5090 | H7 | 2.1699 | 1.0875 | 3.0747 | 2.5430 | 2.5090 | 3.0824 | | 1.7906 | H8 | 2.1699 | 1.0875 | 3.0747 | 2.5430 | 3.0824 | 2.5090 | 1.7906 | |
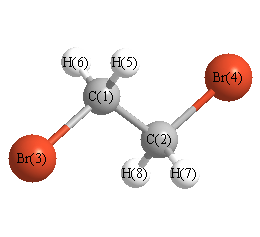 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.424 |
|
C1 |
C2 |
H7 |
112.276 |
| C1 |
C2 |
H8 |
112.276 |
|
C2 |
C1 |
Br3 |
108.424 |
| C2 |
C1 |
H5 |
112.276 |
|
C2 |
C1 |
H6 |
112.276 |
| Br3 |
C1 |
H5 |
106.319 |
|
Br3 |
C1 |
H6 |
106.319 |
| Br4 |
C2 |
H7 |
106.319 |
|
Br4 |
C2 |
H8 |
106.319 |
| H5 |
C1 |
H6 |
110.836 |
|
H7 |
C2 |
H8 |
110.836 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.576 |
|
|
|
| 2 |
C |
-0.576 |
|
|
|
| 3 |
Br |
0.041 |
|
|
|
| 4 |
Br |
0.041 |
|
|
|
| 5 |
H |
0.267 |
|
|
|
| 6 |
H |
0.267 |
|
|
|
| 7 |
H |
0.267 |
|
|
|
| 8 |
H |
0.267 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.358 |
1.450 |
0.000 |
| y |
1.450 |
-55.625 |
0.000 |
| z |
0.000 |
0.000 |
-48.947 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.928 |
1.450 |
0.000 |
| y |
1.450 |
-6.972 |
0.000 |
| z |
0.000 |
0.000 |
3.044 |
|
| Polar |
|---|
| 3z2-r2 | 6.088 |
|---|
| x2-y2 | 7.267 |
|---|
| xy | 1.450 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.004 |
-2.673 |
0.000 |
| y |
-2.673 |
11.448 |
0.000 |
| z |
0.000 |
0.000 |
3.629 |
<r2> (average value of r
2) Å
2
| <r2> |
439.126 |
| (<r2>)1/2 |
20.955 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -5201.800405 |
| Energy at 298.15K | |
| HF Energy | -5201.800405 |
| Nuclear repulsion energy | 443.490650 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3181 |
3070 |
0.02 |
107.05 |
0.74 |
0.85 |
| 2 |
A |
3119 |
3010 |
12.43 |
216.45 |
0.02 |
0.03 |
| 3 |
A |
1512 |
1459 |
0.67 |
9.10 |
0.70 |
0.82 |
| 4 |
A |
1350 |
1303 |
21.90 |
6.14 |
0.63 |
0.78 |
| 5 |
A |
1216 |
1173 |
2.52 |
29.68 |
0.74 |
0.85 |
| 6 |
A |
1035 |
999 |
3.15 |
4.74 |
0.62 |
0.76 |
| 7 |
A |
903 |
872 |
11.36 |
16.57 |
0.54 |
0.70 |
| 8 |
A |
548 |
529 |
7.95 |
16.82 |
0.13 |
0.22 |
| 9 |
A |
229 |
221 |
1.35 |
2.18 |
0.42 |
0.59 |
| 10 |
A |
78 |
75 |
0.25 |
2.17 |
0.74 |
0.85 |
| 11 |
B |
3197 |
3085 |
2.87 |
25.55 |
0.75 |
0.86 |
| 12 |
B |
3110 |
3001 |
1.89 |
65.70 |
0.75 |
0.86 |
| 13 |
B |
1516 |
1463 |
18.14 |
24.91 |
0.75 |
0.86 |
| 14 |
B |
1321 |
1274 |
61.01 |
1.80 |
0.75 |
0.86 |
| 15 |
B |
1165 |
1124 |
2.06 |
11.31 |
0.75 |
0.86 |
| 16 |
B |
849 |
819 |
25.84 |
1.55 |
0.75 |
0.86 |
| 17 |
B |
587 |
566 |
15.29 |
14.89 |
0.75 |
0.86 |
| 18 |
B |
356 |
343 |
6.25 |
4.43 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12635.7 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 12192.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.318 |
0.687 |
1.210 |
| C2 |
-0.318 |
-0.687 |
1.210 |
| Br3 |
-0.318 |
1.833 |
-0.300 |
| Br4 |
0.318 |
-1.833 |
-0.300 |
| H5 |
0.038 |
1.224 |
2.118 |
| H6 |
1.401 |
0.634 |
1.123 |
| H7 |
-0.038 |
-1.224 |
2.118 |
| H8 |
-1.401 |
-0.634 |
1.123 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5131 | 2.0000 | 2.9378 | 1.0916 | 1.0881 | 2.1456 | 2.1695 |
C2 | 1.5131 | | 2.9378 | 2.0000 | 2.1456 | 2.1695 | 1.0916 | 1.0881 | Br3 | 2.0000 | 2.9378 | | 3.7208 | 2.5192 | 2.5328 | 3.9085 | 3.0475 | Br4 | 2.9378 | 2.0000 | 3.7208 | | 3.9085 | 3.0475 | 2.5192 | 2.5328 | H5 | 1.0916 | 2.1456 | 2.5192 | 3.9085 | | 1.7879 | 2.4502 | 2.5531 | H6 | 1.0881 | 2.1695 | 2.5328 | 3.0475 | 1.7879 | | 2.5531 | 3.0759 | H7 | 2.1456 | 1.0916 | 3.9085 | 2.5192 | 2.4502 | 2.5531 | | 1.7879 | H8 | 2.1695 | 1.0881 | 3.0475 | 2.5328 | 2.5531 | 3.0759 | 1.7879 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.762 |
|
C1 |
C2 |
H7 |
109.860 |
| C1 |
C2 |
H8 |
111.989 |
|
C2 |
C1 |
Br3 |
112.762 |
| C2 |
C1 |
H5 |
109.860 |
|
C2 |
C1 |
H6 |
111.989 |
| Br3 |
C1 |
H5 |
105.339 |
|
Br3 |
C1 |
H6 |
106.434 |
| Br4 |
C2 |
H7 |
105.339 |
|
Br4 |
C2 |
H8 |
106.434 |
| H5 |
C1 |
H6 |
110.208 |
|
H7 |
C2 |
H8 |
110.208 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.586 |
|
|
|
| 2 |
C |
-0.586 |
|
|
|
| 3 |
Br |
0.058 |
|
|
|
| 4 |
Br |
0.058 |
|
|
|
| 5 |
H |
0.260 |
|
|
|
| 6 |
H |
0.268 |
|
|
|
| 7 |
H |
0.260 |
|
|
|
| 8 |
H |
0.268 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.608 |
2.608 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.695 |
0.858 |
0.000 |
| y |
0.858 |
-52.919 |
0.000 |
| z |
0.000 |
0.000 |
-44.965 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.247 |
0.858 |
0.000 |
| y |
0.858 |
-6.089 |
0.000 |
| z |
0.000 |
0.000 |
5.842 |
|
| Polar |
|---|
| 3z2-r2 | 11.684 |
|---|
| x2-y2 | 4.224 |
|---|
| xy | 0.858 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.178 |
-1.086 |
0.000 |
| y |
-1.086 |
7.368 |
0.000 |
| z |
0.000 |
0.000 |
6.872 |
<r2> (average value of r
2) Å
2
| <r2> |
322.775 |
| (<r2>)1/2 |
17.966 |
