Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -190.850469 |
| Energy at 298.15K | |
| HF Energy | -190.850469 |
| Nuclear repulsion energy | 102.454408 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3245 |
3131 |
6.69 |
60.82 |
0.44 |
0.61 |
| 2 |
A' |
3201 |
3089 |
1.45 |
81.53 |
0.40 |
0.57 |
| 3 |
A' |
3151 |
3041 |
4.19 |
64.11 |
0.11 |
0.19 |
| 4 |
A' |
2890 |
2789 |
104.37 |
75.55 |
0.29 |
0.45 |
| 5 |
A' |
1720 |
1660 |
125.50 |
55.75 |
0.36 |
0.52 |
| 6 |
A' |
1689 |
1630 |
7.65 |
7.77 |
0.14 |
0.24 |
| 7 |
A' |
1496 |
1443 |
10.98 |
11.51 |
0.44 |
0.61 |
| 8 |
A' |
1431 |
1381 |
4.62 |
17.02 |
0.52 |
0.69 |
| 9 |
A' |
1330 |
1283 |
1.83 |
9.92 |
0.37 |
0.54 |
| 10 |
A' |
1171 |
1130 |
25.22 |
11.13 |
0.62 |
0.77 |
| 11 |
A' |
933 |
900 |
33.60 |
0.71 |
0.13 |
0.23 |
| 12 |
A' |
569 |
549 |
4.90 |
6.00 |
0.34 |
0.51 |
| 13 |
A' |
324 |
313 |
7.41 |
0.72 |
0.32 |
0.49 |
| 14 |
A" |
1063 |
1026 |
7.17 |
4.86 |
0.75 |
0.86 |
| 15 |
A" |
1054 |
1017 |
6.92 |
3.22 |
0.75 |
0.86 |
| 16 |
A" |
1013 |
978 |
57.93 |
0.56 |
0.75 |
0.86 |
| 17 |
A" |
625 |
604 |
12.43 |
6.11 |
0.75 |
0.86 |
| 18 |
A" |
190 |
184 |
2.48 |
1.72 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13548.2 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 13072.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.144 |
-0.743 |
0.000 |
| C2 |
0.000 |
0.726 |
0.000 |
| C3 |
1.208 |
1.296 |
0.000 |
| O4 |
-1.219 |
-1.347 |
0.000 |
| H5 |
0.832 |
-1.270 |
0.000 |
| H6 |
-0.917 |
1.306 |
0.000 |
| H7 |
1.345 |
2.371 |
0.000 |
| H8 |
2.112 |
0.691 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4761 | 2.4459 | 1.2334 | 1.1095 | 2.1896 | 3.4512 | 2.6728 |
C2 | 1.4761 | | 1.3353 | 2.4050 | 2.1626 | 1.0847 | 2.1242 | 2.1122 | C3 | 2.4459 | 1.3353 | | 3.5879 | 2.5925 | 2.1248 | 1.0839 | 1.0877 | O4 | 1.2334 | 2.4050 | 3.5879 | | 2.0533 | 2.6696 | 4.5159 | 3.9049 | H5 | 1.1095 | 2.1626 | 2.5925 | 2.0533 | | 3.1134 | 3.6763 | 2.3408 | H6 | 2.1896 | 1.0847 | 2.1248 | 2.6696 | 3.1134 | | 2.4997 | 3.0907 | H7 | 3.4512 | 2.1242 | 1.0839 | 4.5159 | 3.6763 | 2.4997 | | 1.8472 | H8 | 2.6728 | 2.1122 | 1.0877 | 3.9049 | 2.3408 | 3.0907 | 1.8472 | |
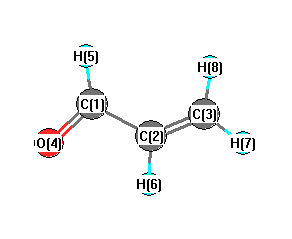 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.830 |
|
C1 |
C2 |
H6 |
116.696 |
| C2 |
C1 |
O4 |
124.911 |
|
C2 |
C1 |
H5 |
112.752 |
| C2 |
C3 |
H7 |
122.475 |
|
C2 |
C3 |
H8 |
120.977 |
| C3 |
C2 |
H6 |
122.474 |
|
O4 |
C1 |
H5 |
122.336 |
| H7 |
C3 |
H8 |
116.548 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.246 |
|
|
|
| 2 |
C |
-0.256 |
|
|
|
| 3 |
C |
-0.350 |
|
|
|
| 4 |
O |
-0.427 |
|
|
|
| 5 |
H |
0.154 |
|
|
|
| 6 |
H |
0.216 |
|
|
|
| 7 |
H |
0.214 |
|
|
|
| 8 |
H |
0.205 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.245 |
2.011 |
0.000 |
3.014 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.986 |
-2.407 |
0.000 |
| y |
-2.407 |
-23.835 |
0.000 |
| z |
0.000 |
0.000 |
-24.767 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.315 |
-2.407 |
0.000 |
| y |
-2.407 |
0.541 |
0.000 |
| z |
0.000 |
0.000 |
-0.856 |
|
| Polar |
|---|
| 3z2-r2 | -1.713 |
|---|
| x2-y2 | -0.151 |
|---|
| xy | -2.407 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.255 |
1.662 |
0.000 |
| y |
1.662 |
5.702 |
0.000 |
| z |
0.000 |
0.000 |
1.542 |
<r2> (average value of r
2) Å
2
| <r2> |
84.158 |
| (<r2>)1/2 |
9.174 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -190.850151 |
| Energy at 298.15K | |
| HF Energy | -190.850151 |
| Nuclear repulsion energy | 104.291940 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3255 |
3141 |
3.05 |
50.59 |
0.52 |
0.68 |
| 2 |
A' |
3198 |
3086 |
7.39 |
90.43 |
0.32 |
0.49 |
| 3 |
A' |
3162 |
3051 |
3.34 |
56.76 |
0.10 |
0.19 |
| 4 |
A' |
2940 |
2837 |
144.51 |
129.78 |
0.33 |
0.49 |
| 5 |
A' |
1739 |
1678 |
22.82 |
2.11 |
0.63 |
0.78 |
| 6 |
A' |
1651 |
1593 |
76.91 |
43.75 |
0.27 |
0.42 |
| 7 |
A' |
1479 |
1428 |
35.05 |
8.53 |
0.27 |
0.42 |
| 8 |
A' |
1449 |
1398 |
11.49 |
11.44 |
0.72 |
0.84 |
| 9 |
A' |
1356 |
1309 |
2.54 |
14.54 |
0.47 |
0.64 |
| 10 |
A' |
1105 |
1066 |
8.20 |
5.68 |
0.72 |
0.84 |
| 11 |
A' |
930 |
897 |
62.91 |
4.74 |
0.20 |
0.33 |
| 12 |
A' |
705 |
680 |
4.53 |
0.72 |
0.69 |
0.82 |
| 13 |
A' |
296 |
286 |
9.18 |
2.95 |
0.49 |
0.66 |
| 14 |
A" |
1079 |
1041 |
2.71 |
11.11 |
0.75 |
0.86 |
| 15 |
A" |
1047 |
1010 |
34.96 |
0.88 |
0.75 |
0.86 |
| 16 |
A" |
1033 |
996 |
23.52 |
0.03 |
0.75 |
0.86 |
| 17 |
A" |
592 |
572 |
13.66 |
7.55 |
0.75 |
0.86 |
| 18 |
A" |
244 |
235 |
7.35 |
1.14 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13630.3 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 13151.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.891 |
-0.273 |
0.000 |
| C2 |
0.000 |
0.912 |
0.000 |
| C3 |
1.326 |
0.753 |
0.000 |
| O4 |
-0.491 |
-1.443 |
0.000 |
| H5 |
-1.968 |
-0.026 |
0.000 |
| H6 |
-0.471 |
1.890 |
0.000 |
| H7 |
2.020 |
1.585 |
0.000 |
| H8 |
1.737 |
-0.253 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4831 | 2.4426 | 1.2359 | 1.1049 | 2.2034 | 3.4533 | 2.6280 |
C2 | 1.4831 | | 1.3354 | 2.4059 | 2.1802 | 1.0848 | 2.1289 | 2.0918 | C3 | 2.4426 | 1.3354 | | 2.8499 | 3.3845 | 2.1262 | 1.0837 | 1.0866 | O4 | 1.2359 | 2.4059 | 2.8499 | | 2.0462 | 3.3326 | 3.9336 | 2.5261 | H5 | 1.1049 | 2.1802 | 3.3845 | 2.0462 | | 2.4313 | 4.3009 | 3.7118 | H6 | 2.2034 | 1.0848 | 2.1262 | 3.3326 | 2.4313 | | 2.5093 | 3.0767 | H7 | 3.4533 | 2.1289 | 1.0837 | 3.9336 | 4.3009 | 2.5093 | | 1.8596 | H8 | 2.6280 | 2.0918 | 1.0866 | 2.5261 | 3.7118 | 3.0767 | 1.8596 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.050 |
|
C1 |
C2 |
H6 |
117.357 |
| C2 |
C1 |
O4 |
124.218 |
|
C2 |
C1 |
H5 |
113.996 |
| C2 |
C3 |
H7 |
122.958 |
|
C2 |
C3 |
H8 |
119.107 |
| C3 |
C2 |
H6 |
122.593 |
|
O4 |
C1 |
H5 |
121.786 |
| H7 |
C3 |
H8 |
117.935 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.241 |
|
|
|
| 2 |
C |
-0.285 |
|
|
|
| 3 |
C |
-0.319 |
|
|
|
| 4 |
O |
-0.426 |
|
|
|
| 5 |
H |
0.161 |
|
|
|
| 6 |
H |
0.197 |
|
|
|
| 7 |
H |
0.206 |
|
|
|
| 8 |
H |
0.225 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.275 |
2.484 |
0.000 |
2.499 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.170 |
-0.626 |
0.000 |
| y |
-0.626 |
-25.763 |
0.000 |
| z |
0.000 |
0.000 |
-24.744 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.084 |
-0.626 |
0.000 |
| y |
-0.626 |
-3.306 |
0.000 |
| z |
0.000 |
0.000 |
-1.778 |
|
| Polar |
|---|
| 3z2-r2 | -3.556 |
|---|
| x2-y2 | 5.593 |
|---|
| xy | -0.626 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.429 |
0.433 |
0.000 |
| y |
0.433 |
5.093 |
0.000 |
| z |
0.000 |
0.000 |
1.546 |
<r2> (average value of r
2) Å
2
| <r2> |
74.789 |
| (<r2>)1/2 |
8.648 |
