Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -610.368060 |
| Energy at 298.15K | -610.371318 |
| HF Energy | -610.368060 |
| Nuclear repulsion energy | 139.966313 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3220 |
3107 |
0.34 |
|
|
|
| 2 |
A |
3127 |
3017 |
2.09 |
|
|
|
| 3 |
A |
2974 |
2870 |
66.28 |
|
|
|
| 4 |
A |
1739 |
1678 |
87.23 |
|
|
|
| 5 |
A |
1497 |
1444 |
14.59 |
|
|
|
| 6 |
A |
1441 |
1391 |
3.44 |
|
|
|
| 7 |
A |
1263 |
1219 |
12.37 |
|
|
|
| 8 |
A |
1190 |
1148 |
7.95 |
|
|
|
| 9 |
A |
1036 |
999 |
22.84 |
|
|
|
| 10 |
A |
1006 |
971 |
15.83 |
|
|
|
| 11 |
A |
808 |
779 |
4.48 |
|
|
|
| 12 |
A |
656 |
633 |
28.59 |
|
|
|
| 13 |
A |
446 |
430 |
10.99 |
|
|
|
| 14 |
A |
289 |
279 |
9.26 |
|
|
|
| 15 |
A |
77 |
75 |
10.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10383.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 10019.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.059 |
0.781 |
0.094 |
| C2 |
1.180 |
-0.214 |
0.328 |
| Cl3 |
-1.551 |
-0.213 |
-0.082 |
| O4 |
2.235 |
-0.202 |
-0.298 |
| H5 |
-0.101 |
1.431 |
0.955 |
| H6 |
0.202 |
1.341 |
-0.827 |
| H7 |
0.952 |
-0.946 |
1.120 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5172 | 1.9009 | 2.4196 | 1.0907 | 1.0867 | 2.1983 |
C2 | 1.5172 | | 2.7622 | 1.2267 | 2.1774 | 2.1694 | 1.1025 | Cl3 | 1.9009 | 2.7622 | | 3.7924 | 2.4257 | 2.4577 | 2.8722 | O4 | 2.4196 | 1.2267 | 3.7924 | | 3.1136 | 2.6061 | 2.0523 | H5 | 1.0907 | 2.1774 | 2.4257 | 3.1136 | | 1.8098 | 2.6049 | H6 | 1.0867 | 2.1694 | 2.4577 | 2.6061 | 1.8098 | | 3.0951 | H7 | 2.1983 | 1.1025 | 2.8722 | 2.0523 | 2.6049 | 3.0951 | |
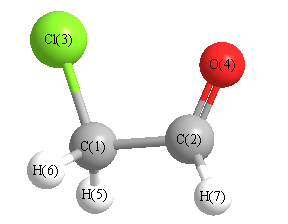 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
123.382 |
|
C1 |
C2 |
H7 |
113.152 |
| C2 |
C1 |
Cl3 |
107.293 |
|
C2 |
C1 |
H5 |
112.184 |
| C2 |
C1 |
H6 |
111.783 |
|
Cl3 |
C1 |
H5 |
105.111 |
| Cl3 |
C1 |
H6 |
107.556 |
|
O4 |
C2 |
H7 |
123.466 |
| H5 |
C1 |
H6 |
112.440 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.619 |
|
|
|
| 2 |
C |
0.280 |
|
|
|
| 3 |
Cl |
-0.027 |
|
|
|
| 4 |
O |
-0.385 |
|
|
|
| 5 |
H |
0.279 |
|
|
|
| 6 |
H |
0.281 |
|
|
|
| 7 |
H |
0.192 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.035 |
1.346 |
1.257 |
1.842 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.692 |
0.345 |
1.707 |
| y |
0.345 |
-28.518 |
-0.120 |
| z |
1.707 |
-0.120 |
-29.415 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.725 |
0.345 |
1.707 |
| y |
0.345 |
5.535 |
-0.120 |
| z |
1.707 |
-0.120 |
4.190 |
|
| Polar |
|---|
| 3z2-r2 | 8.379 |
|---|
| x2-y2 | -10.174 |
|---|
| xy | 0.345 |
|---|
| xz | 1.707 |
|---|
| yz | -0.120 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.423 |
0.644 |
-0.273 |
| y |
0.644 |
3.597 |
-0.245 |
| z |
-0.273 |
-0.245 |
3.146 |
<r2> (average value of r
2) Å
2
| <r2> |
124.476 |
| (<r2>)1/2 |
11.157 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -610.367392 |
| Energy at 298.15K | -610.370797 |
| HF Energy | -610.367392 |
| Nuclear repulsion energy | 143.934203 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3103 |
2994 |
6.40 |
|
|
|
| 2 |
A' |
2905 |
2803 |
84.78 |
|
|
|
| 3 |
A' |
1762 |
1700 |
72.06 |
|
|
|
| 4 |
A' |
1483 |
1431 |
34.28 |
|
|
|
| 5 |
A' |
1430 |
1380 |
13.42 |
|
|
|
| 6 |
A' |
1286 |
1241 |
17.23 |
|
|
|
| 7 |
A' |
911 |
879 |
7.13 |
|
|
|
| 8 |
A' |
750 |
723 |
4.04 |
|
|
|
| 9 |
A' |
588 |
567 |
41.98 |
|
|
|
| 10 |
A' |
214 |
206 |
1.73 |
|
|
|
| 11 |
A" |
3161 |
3050 |
0.02 |
|
|
|
| 12 |
A" |
1223 |
1180 |
1.88 |
|
|
|
| 13 |
A" |
1024 |
988 |
0.83 |
|
|
|
| 14 |
A" |
717 |
692 |
6.44 |
|
|
|
| 15 |
A" |
167 |
162 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10361.7 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 9998.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.981 |
0.000 |
| C2 |
1.371 |
0.337 |
0.000 |
| Cl3 |
-1.355 |
-0.313 |
0.000 |
| O4 |
1.616 |
-0.861 |
0.000 |
| H5 |
-0.139 |
1.593 |
0.894 |
| H6 |
-0.139 |
1.593 |
-0.894 |
| H7 |
2.165 |
1.111 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5147 | 1.8737 | 2.4501 | 1.0925 | 1.0925 | 2.1685 |
C2 | 1.5147 | | 2.8029 | 1.2223 | 2.1584 | 2.1584 | 1.1083 | Cl3 | 1.8737 | 2.8029 | | 3.0219 | 2.4311 | 2.4311 | 3.7970 | O4 | 2.4501 | 1.2223 | 3.0219 | | 3.1468 | 3.1468 | 2.0461 | H5 | 1.0925 | 2.1584 | 2.4311 | 3.1468 | | 1.7879 | 2.5180 | H6 | 1.0925 | 2.1584 | 2.4311 | 3.1468 | 1.7879 | | 2.5180 | H7 | 2.1685 | 1.1083 | 3.7970 | 2.0461 | 2.5180 | 2.5180 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
126.726 |
|
C1 |
C2 |
H7 |
110.568 |
| C2 |
C1 |
Cl3 |
111.186 |
|
C2 |
C1 |
H5 |
110.718 |
| C2 |
C1 |
H6 |
110.718 |
|
Cl3 |
C1 |
H5 |
107.134 |
| Cl3 |
C1 |
H6 |
107.134 |
|
O4 |
C2 |
H7 |
122.705 |
| H5 |
C1 |
H6 |
109.825 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.644 |
|
|
|
| 2 |
C |
0.275 |
|
|
|
| 3 |
Cl |
0.002 |
|
|
|
| 4 |
O |
-0.368 |
|
|
|
| 5 |
H |
0.278 |
|
|
|
| 6 |
H |
0.278 |
|
|
|
| 7 |
H |
0.180 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.006 |
3.664 |
0.000 |
3.800 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.973 |
3.020 |
0.000 |
| y |
3.020 |
-29.272 |
0.000 |
| z |
0.000 |
0.000 |
-29.448 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.613 |
3.020 |
0.000 |
| y |
3.020 |
1.939 |
0.000 |
| z |
0.000 |
0.000 |
1.674 |
|
| Polar |
|---|
| 3z2-r2 | 3.349 |
|---|
| x2-y2 | -3.702 |
|---|
| xy | 3.020 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.426 |
0.970 |
0.000 |
| y |
0.970 |
5.033 |
0.000 |
| z |
0.000 |
0.000 |
2.487 |
<r2> (average value of r
2) Å
2
| <r2> |
109.177 |
| (<r2>)1/2 |
10.449 |
