Jump to
S1C2
Vibrational Frequencies calculated at M06-2X/3-21G
Geometric Data calculated at M06-2X/3-21G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at M06-2X/3-21G
| | hartrees |
|---|
| Energy at 0K | -207.872095 |
| Energy at 298.15K | |
| HF Energy | -207.872095 |
| Nuclear repulsion energy | 118.541187 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3557 |
3410 |
42.11 |
|
|
|
| 2 |
A' |
3198 |
3065 |
8.24 |
|
|
|
| 3 |
A' |
3177 |
3046 |
1.75 |
|
|
|
| 4 |
A' |
3048 |
2922 |
4.69 |
|
|
|
| 5 |
A' |
1738 |
1666 |
4.94 |
|
|
|
| 6 |
A' |
1533 |
1470 |
15.17 |
|
|
|
| 7 |
A' |
1447 |
1388 |
28.52 |
|
|
|
| 8 |
A' |
1386 |
1329 |
0.32 |
|
|
|
| 9 |
A' |
1344 |
1288 |
72.55 |
|
|
|
| 10 |
A' |
1157 |
1109 |
18.16 |
|
|
|
| 11 |
A' |
956 |
917 |
94.11 |
|
|
|
| 12 |
A' |
919 |
881 |
0.29 |
|
|
|
| 13 |
A' |
669 |
641 |
13.08 |
|
|
|
| 14 |
A' |
320 |
307 |
2.88 |
|
|
|
| 15 |
A" |
3101 |
2973 |
4.98 |
|
|
|
| 16 |
A" |
1560 |
1496 |
17.36 |
|
|
|
| 17 |
A" |
1122 |
1076 |
0.14 |
|
|
|
| 18 |
A" |
916 |
878 |
15.01 |
|
|
|
| 19 |
A" |
492 |
472 |
40.92 |
|
|
|
| 20 |
A" |
395 |
379 |
126.89 |
|
|
|
| 21 |
A" |
67i |
64i |
0.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15984.1 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 15323.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.439 |
0.456 |
0.000 |
| C2 |
0.000 |
0.887 |
0.000 |
| N3 |
1.035 |
0.132 |
0.000 |
| O4 |
0.631 |
-1.284 |
0.000 |
| H5 |
1.514 |
-1.735 |
0.000 |
| H6 |
-1.494 |
-0.631 |
0.000 |
| H7 |
-1.949 |
0.851 |
0.885 |
| H8 |
-1.949 |
0.851 |
-0.885 |
| H9 |
0.225 |
1.950 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5027 | 2.4955 | 2.7042 | 3.6771 | 1.0886 | 1.0953 | 1.0953 | 2.2366 |
C2 | 1.5027 | | 1.2813 | 2.2610 | 3.0277 | 2.1302 | 2.1410 | 2.1410 | 1.0863 | N3 | 2.4955 | 1.2813 | | 1.4725 | 1.9272 | 2.6414 | 3.1946 | 3.1946 | 1.9904 | O4 | 2.7042 | 2.2610 | 1.4725 | | 0.9915 | 2.2225 | 3.4639 | 3.4639 | 3.2594 | H5 | 3.6771 | 3.0277 | 1.9272 | 0.9915 | | 3.2036 | 4.4118 | 4.4118 | 3.9038 | H6 | 1.0886 | 2.1302 | 2.6414 | 2.2225 | 3.2036 | | 1.7859 | 1.7859 | 3.1012 | H7 | 1.0953 | 2.1410 | 3.1946 | 3.4639 | 4.4118 | 1.7859 | | 1.7703 | 2.5917 | H8 | 1.0953 | 2.1410 | 3.1946 | 3.4639 | 4.4118 | 1.7859 | 1.7703 | | 2.5917 | H9 | 2.2366 | 1.0863 | 1.9904 | 3.2594 | 3.9038 | 3.1012 | 2.5917 | 2.5917 | |
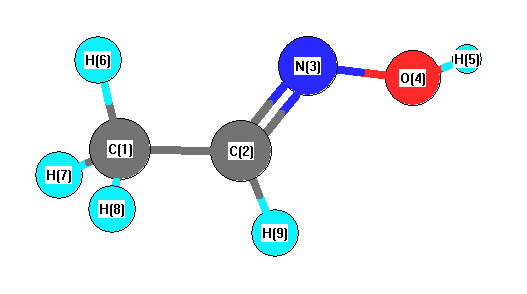 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.192 |
|
C1 |
C2 |
H9 |
118.632 |
| C2 |
C1 |
H6 |
109.545 |
|
C2 |
C1 |
H7 |
110.006 |
| C2 |
C1 |
H8 |
110.006 |
|
C2 |
N3 |
O4 |
110.191 |
| N3 |
C2 |
H9 |
114.176 |
|
N3 |
O4 |
H5 |
101.111 |
| H6 |
C1 |
H7 |
109.718 |
|
H6 |
C1 |
H8 |
109.718 |
| H7 |
C1 |
H8 |
107.825 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.652 |
|
|
|
| 2 |
C |
0.058 |
|
|
|
| 3 |
N |
-0.242 |
|
|
|
| 4 |
O |
-0.476 |
|
|
|
| 5 |
H |
0.383 |
|
|
|
| 6 |
H |
0.256 |
|
|
|
| 7 |
H |
0.220 |
|
|
|
| 8 |
H |
0.220 |
|
|
|
| 9 |
H |
0.232 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.513 |
0.301 |
0.000 |
0.595 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.659 |
-4.085 |
0.000 |
| y |
-4.085 |
-20.781 |
0.000 |
| z |
0.000 |
0.000 |
-24.990 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.226 |
-4.085 |
0.000 |
| y |
-4.085 |
3.043 |
0.000 |
| z |
0.000 |
0.000 |
-3.270 |
|
| Polar |
|---|
| 3z2-r2 | -6.540 |
|---|
| x2-y2 | -1.878 |
|---|
| xy | -4.085 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.059 |
-1.322 |
0.000 |
| y |
-1.322 |
5.510 |
0.000 |
| z |
0.000 |
0.000 |
2.402 |
<r2> (average value of r
2) Å
2
| <r2> |
79.039 |
| (<r2>)1/2 |
8.890 |
