Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP=FULL/3-21G
Geometric Data calculated at B2PLYP=FULL/3-21G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/3-21G
| | hartrees |
|---|
| Energy at 0K | -207.694581 |
| Energy at 298.15K | -207.700415 |
| HF Energy | -207.549475 |
| Nuclear repulsion energy | 117.760658 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3507 |
3339 |
21.58 |
|
|
|
| 2 |
A' |
3213 |
3059 |
3.16 |
|
|
|
| 3 |
A' |
3184 |
3031 |
11.47 |
|
|
|
| 4 |
A' |
3081 |
2933 |
9.85 |
|
|
|
| 5 |
A' |
1662 |
1582 |
2.08 |
|
|
|
| 6 |
A' |
1558 |
1483 |
13.97 |
|
|
|
| 7 |
A' |
1467 |
1397 |
20.50 |
|
|
|
| 8 |
A' |
1390 |
1324 |
0.53 |
|
|
|
| 9 |
A' |
1330 |
1266 |
65.63 |
|
|
|
| 10 |
A' |
1163 |
1107 |
20.14 |
|
|
|
| 11 |
A' |
924 |
880 |
44.48 |
|
|
|
| 12 |
A' |
873 |
831 |
43.03 |
|
|
|
| 13 |
A' |
660 |
629 |
13.66 |
|
|
|
| 14 |
A' |
318 |
303 |
2.38 |
|
|
|
| 15 |
A" |
3128 |
2978 |
11.95 |
|
|
|
| 16 |
A" |
1584 |
1508 |
13.86 |
|
|
|
| 17 |
A" |
1125 |
1071 |
0.10 |
|
|
|
| 18 |
A" |
885 |
843 |
14.57 |
|
|
|
| 19 |
A" |
478 |
455 |
19.57 |
|
|
|
| 20 |
A" |
336 |
320 |
129.69 |
|
|
|
| 21 |
A" |
13 |
13 |
1.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15938.8 cm
-1
Scaled (by 0.952) Zero Point Vibrational Energy (zpe) 15173.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.446 |
0.473 |
0.000 |
| C2 |
0.000 |
0.891 |
0.000 |
| N3 |
1.043 |
0.131 |
0.000 |
| O4 |
0.630 |
-1.304 |
0.000 |
| H5 |
1.524 |
-1.738 |
0.000 |
| H6 |
-1.510 |
-0.613 |
0.000 |
| H7 |
-1.953 |
0.871 |
0.884 |
| H8 |
-1.953 |
0.871 |
-0.884 |
| H9 |
0.230 |
1.951 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5050 | 2.5121 | 2.7322 | 3.7019 | 1.0876 | 1.0948 | 1.0948 | 2.2352 |
C2 | 1.5050 | | 1.2905 | 2.2834 | 3.0383 | 2.1312 | 2.1443 | 2.1443 | 1.0855 | N3 | 2.5121 | 1.2905 | | 1.4932 | 1.9294 | 2.6593 | 3.2106 | 3.2106 | 1.9940 | O4 | 2.7322 | 2.2834 | 1.4932 | | 0.9936 | 2.2489 | 3.4908 | 3.4908 | 3.2801 | H5 | 3.7019 | 3.0383 | 1.9294 | 0.9936 | | 3.2358 | 4.4360 | 4.4360 | 3.9095 | H6 | 1.0876 | 2.1312 | 2.6593 | 2.2489 | 3.2358 | | 1.7834 | 1.7834 | 3.0994 | H7 | 1.0948 | 2.1443 | 3.2106 | 3.4908 | 4.4360 | 1.7834 | | 1.7689 | 2.5919 | H8 | 1.0948 | 2.1443 | 3.2106 | 3.4908 | 4.4360 | 1.7834 | 1.7689 | | 2.5919 | H9 | 2.2352 | 1.0855 | 1.9940 | 3.2801 | 3.9095 | 3.0994 | 2.5919 | 2.5919 | |
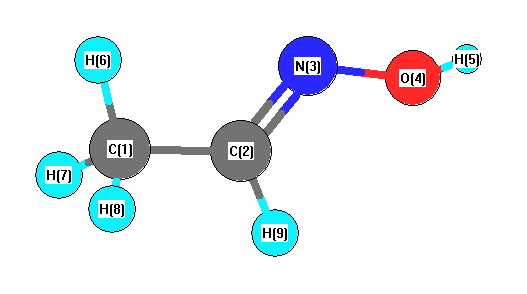 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.789 |
|
C1 |
C2 |
H9 |
118.375 |
| C2 |
C1 |
H6 |
109.531 |
|
C2 |
C1 |
H7 |
110.141 |
| C2 |
C1 |
H8 |
110.141 |
|
C2 |
N3 |
O4 |
110.017 |
| N3 |
C2 |
H9 |
113.836 |
|
N3 |
O4 |
H5 |
99.814 |
| H6 |
C1 |
H7 |
109.611 |
|
H6 |
C1 |
H8 |
109.611 |
| H7 |
C1 |
H8 |
107.782 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
