Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -207.974938 |
| Energy at 298.15K | -207.980869 |
| Nuclear repulsion energy | 115.100855 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3480 |
3358 |
18.59 |
|
|
|
| 2 |
A |
3149 |
3038 |
14.05 |
|
|
|
| 3 |
A |
3129 |
3019 |
2.58 |
|
|
|
| 4 |
A |
3047 |
2940 |
13.72 |
|
|
|
| 5 |
A |
1675 |
1616 |
2.67 |
|
|
|
| 6 |
A |
1541 |
1487 |
9.92 |
|
|
|
| 7 |
A |
1458 |
1407 |
7.63 |
|
|
|
| 8 |
A |
1412 |
1362 |
23.34 |
|
|
|
| 9 |
A |
1279 |
1234 |
51.47 |
|
|
|
| 10 |
A |
1156 |
1116 |
9.49 |
|
|
|
| 11 |
A |
990 |
955 |
78.67 |
|
|
|
| 12 |
A |
898 |
866 |
19.16 |
|
|
|
| 13 |
A |
541 |
522 |
15.25 |
|
|
|
| 14 |
A |
309 |
298 |
2.24 |
|
|
|
| 15 |
A |
3093 |
2984 |
13.47 |
|
|
|
| 16 |
A |
1550 |
1496 |
12.07 |
|
|
|
| 17 |
A |
1111 |
1072 |
0.02 |
|
|
|
| 18 |
A |
930 |
897 |
22.83 |
|
|
|
| 19 |
A |
363 |
350 |
149.28 |
|
|
|
| 20 |
A |
273 |
264 |
6.73 |
|
|
|
| 21 |
A |
207 |
199 |
0.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15794.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 15240.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.271 |
1.364 |
0.000 |
| C2 |
0.000 |
0.561 |
0.000 |
| N3 |
-0.052 |
-0.722 |
0.000 |
| O4 |
1.333 |
-1.257 |
0.000 |
| H5 |
1.146 |
-2.236 |
0.000 |
| H6 |
-2.134 |
0.694 |
0.000 |
| H7 |
-1.317 |
2.009 |
0.886 |
| H8 |
-1.317 |
2.009 |
-0.886 |
| H9 |
0.954 |
1.086 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5035 | 2.4157 | 3.6946 | 4.3364 | 1.0927 | 1.0970 | 1.0970 | 2.2424 |
C2 | 1.5035 | | 1.2836 | 2.2541 | 3.0225 | 2.1382 | 2.1493 | 2.1493 | 1.0890 | N3 | 2.4157 | 1.2836 | | 1.4850 | 1.9314 | 2.5175 | 3.1376 | 3.1376 | 2.0688 | O4 | 3.6946 | 2.2541 | 1.4850 | | 0.9962 | 3.9781 | 4.2987 | 4.2987 | 2.3732 | H5 | 4.3364 | 3.0225 | 1.9314 | 0.9962 | | 4.3985 | 4.9879 | 4.9879 | 3.3271 | H6 | 1.0927 | 2.1382 | 2.5175 | 3.9781 | 4.3985 | | 1.7839 | 1.7839 | 3.1129 | H7 | 1.0970 | 2.1493 | 3.1376 | 4.2987 | 4.9879 | 1.7839 | | 1.7718 | 2.6074 | H8 | 1.0970 | 2.1493 | 3.1376 | 4.2987 | 4.9879 | 1.7839 | 1.7718 | | 2.6074 | H9 | 2.2424 | 1.0890 | 2.0688 | 2.3732 | 3.3271 | 3.1129 | 2.6074 | 2.6074 | |
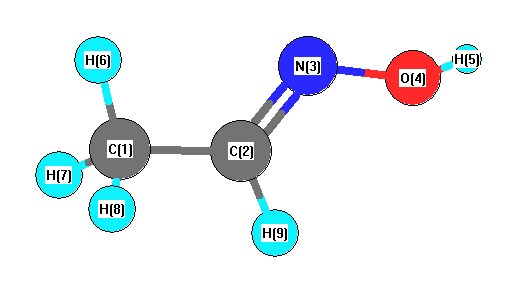 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
119.958 |
|
C1 |
C2 |
H9 |
118.884 |
| C2 |
C1 |
H6 |
109.877 |
|
C2 |
C1 |
H7 |
110.498 |
| C2 |
C1 |
H8 |
110.498 |
|
C2 |
N3 |
O4 |
108.795 |
| N3 |
C2 |
H9 |
121.158 |
|
N3 |
O4 |
H5 |
100.371 |
| H6 |
C1 |
H7 |
109.103 |
|
H6 |
C1 |
H8 |
109.103 |
| H7 |
C1 |
H8 |
107.714 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.441 |
|
|
-0.467 |
| 2 |
H |
0.362 |
|
|
0.343 |
| 3 |
N |
-0.248 |
|
|
-0.327 |
| 4 |
C |
0.089 |
|
|
-0.425 |
| 5 |
H |
0.207 |
|
|
0.411 |
| 6 |
C |
-0.615 |
|
|
0.144 |
| 7 |
H |
0.222 |
|
|
0.135 |
| 8 |
H |
0.212 |
|
|
0.135 |
| 9 |
H |
0.212 |
|
|
0.051 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.486 |
0.534 |
0.000 |
0.722 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.470 |
0.503 |
0.000 |
0.688 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.208 |
-2.083 |
0.000 |
| y |
-2.083 |
-18.102 |
0.000 |
| z |
0.000 |
0.000 |
-24.796 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.759 |
-2.083 |
0.000 |
| y |
-2.083 |
6.899 |
0.000 |
| z |
0.000 |
0.000 |
-3.141 |
|
| Polar |
|---|
| 3z2-r2 | -6.282 |
|---|
| x2-y2 | -7.105 |
|---|
| xy | -2.083 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.695 |
-1.476 |
0.000 |
| y |
-1.476 |
6.659 |
0.000 |
| z |
0.000 |
0.000 |
2.434 |
<r2> (average value of r
2) Å
2
| <r2> |
93.996 |
| (<r2>)1/2 |
9.695 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -207.976268 |
| Energy at 298.15K | -207.982164 |
| HF Energy | -207.976268 |
| Nuclear repulsion energy | 117.947702 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3490 |
3368 |
18.63 |
|
|
|
| 2 |
A' |
3189 |
3077 |
3.34 |
|
|
|
| 3 |
A' |
3155 |
3044 |
12.00 |
|
|
|
| 4 |
A' |
3052 |
2945 |
10.05 |
|
|
|
| 5 |
A' |
1682 |
1623 |
3.29 |
|
|
|
| 6 |
A' |
1538 |
1484 |
14.91 |
|
|
|
| 7 |
A' |
1448 |
1397 |
22.45 |
|
|
|
| 8 |
A' |
1379 |
1331 |
0.62 |
|
|
|
| 9 |
A' |
1334 |
1287 |
69.88 |
|
|
|
| 10 |
A' |
1154 |
1113 |
17.53 |
|
|
|
| 11 |
A' |
923 |
891 |
58.65 |
|
|
|
| 12 |
A' |
884 |
853 |
42.43 |
|
|
|
| 13 |
A' |
663 |
640 |
12.93 |
|
|
|
| 14 |
A' |
314 |
303 |
2.19 |
|
|
|
| 15 |
A" |
3093 |
2985 |
12.28 |
|
|
|
| 16 |
A" |
1562 |
1507 |
14.56 |
|
|
|
| 17 |
A" |
1107 |
1068 |
0.02 |
|
|
|
| 18 |
A" |
877 |
846 |
15.79 |
|
|
|
| 19 |
A" |
483 |
467 |
21.03 |
|
|
|
| 20 |
A" |
343 |
331 |
123.65 |
|
|
|
| 21 |
A" |
43 |
41 |
1.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15856.2 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 15299.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.445 |
0.474 |
0.000 |
| C2 |
0.000 |
0.886 |
0.000 |
| N3 |
1.038 |
0.128 |
0.000 |
| O4 |
0.634 |
-1.301 |
0.000 |
| H5 |
1.530 |
-1.735 |
0.000 |
| H6 |
-1.519 |
-0.612 |
0.000 |
| H7 |
-1.953 |
0.877 |
0.885 |
| H8 |
-1.953 |
0.877 |
-0.885 |
| H9 |
0.231 |
1.949 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5029 | 2.5077 | 2.7343 | 3.7058 | 1.0891 | 1.0971 | 1.0971 | 2.2326 |
C2 | 1.5029 | | 1.2857 | 2.2774 | 3.0346 | 2.1338 | 2.1446 | 2.1446 | 1.0876 | N3 | 2.5077 | 1.2857 | | 1.4852 | 1.9261 | 2.6624 | 3.2086 | 3.2086 | 1.9921 | O4 | 2.7343 | 2.2774 | 1.4852 | | 0.9955 | 2.2607 | 3.4963 | 3.4963 | 3.2751 | H5 | 3.7058 | 3.0346 | 1.9261 | 0.9955 | | 3.2494 | 4.4431 | 4.4431 | 3.9058 | H6 | 1.0891 | 2.1338 | 2.6624 | 2.2607 | 3.2494 | | 1.7862 | 1.7862 | 3.1020 | H7 | 1.0971 | 2.1446 | 3.2086 | 3.4963 | 4.4431 | 1.7862 | | 1.7698 | 2.5890 | H8 | 1.0971 | 2.1446 | 3.2086 | 3.4963 | 4.4431 | 1.7862 | 1.7698 | | 2.5890 | H9 | 2.2326 | 1.0876 | 1.9921 | 3.2751 | 3.9058 | 3.1020 | 2.5890 | 2.5890 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.954 |
|
C1 |
C2 |
H9 |
118.158 |
| C2 |
C1 |
H6 |
109.789 |
|
C2 |
C1 |
H7 |
110.165 |
| C2 |
C1 |
H8 |
110.165 |
|
C2 |
N3 |
O4 |
110.345 |
| N3 |
C2 |
H9 |
113.888 |
|
N3 |
O4 |
H5 |
99.992 |
| H6 |
C1 |
H7 |
109.580 |
|
H6 |
C1 |
H8 |
109.580 |
| H7 |
C1 |
H8 |
107.528 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.631 |
|
|
|
| 2 |
C |
0.077 |
|
|
|
| 3 |
N |
-0.247 |
|
|
|
| 4 |
O |
-0.444 |
|
|
|
| 5 |
H |
0.365 |
|
|
|
| 6 |
H |
0.243 |
|
|
|
| 7 |
H |
0.211 |
|
|
|
| 8 |
H |
0.211 |
|
|
|
| 9 |
H |
0.215 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.502 |
0.271 |
0.000 |
0.570 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.526 |
-3.919 |
0.000 |
| y |
-3.919 |
-20.793 |
0.000 |
| z |
0.000 |
0.000 |
-24.825 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.283 |
-3.919 |
0.000 |
| y |
-3.919 |
2.882 |
0.000 |
| z |
0.000 |
0.000 |
-3.166 |
|
| Polar |
|---|
| 3z2-r2 | -6.331 |
|---|
| x2-y2 | -1.733 |
|---|
| xy | -3.919 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.213 |
-1.386 |
0.000 |
| y |
-1.386 |
5.668 |
0.000 |
| z |
0.000 |
0.000 |
2.434 |
<r2> (average value of r
2) Å
2
| <r2> |
79.827 |
| (<r2>)1/2 |
8.935 |
