Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/3-21G
Geometric Data calculated at B3LYP/3-21G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -270.295092 |
| Energy at 298.15K | -270.305721 |
| Nuclear repulsion energy | 234.649213 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3160 |
3050 |
11.33 |
|
|
|
| 2 |
A |
3130 |
3020 |
46.22 |
|
|
|
| 3 |
A |
3114 |
3005 |
25.92 |
|
|
|
| 4 |
A |
3113 |
3004 |
16.28 |
|
|
|
| 5 |
A |
3104 |
2995 |
8.76 |
|
|
|
| 6 |
A |
3085 |
2977 |
1.44 |
|
|
|
| 7 |
A |
3067 |
2960 |
19.25 |
|
|
|
| 8 |
A |
3054 |
2947 |
9.62 |
|
|
|
| 9 |
A |
3049 |
2942 |
0.78 |
|
|
|
| 10 |
A |
3048 |
2941 |
21.33 |
|
|
|
| 11 |
A |
1750 |
1688 |
94.37 |
|
|
|
| 12 |
A |
1571 |
1516 |
8.68 |
|
|
|
| 13 |
A |
1566 |
1511 |
9.22 |
|
|
|
| 14 |
A |
1557 |
1502 |
4.39 |
|
|
|
| 15 |
A |
1544 |
1490 |
10.98 |
|
|
|
| 16 |
A |
1532 |
1479 |
1.82 |
|
|
|
| 17 |
A |
1520 |
1466 |
14.21 |
|
|
|
| 18 |
A |
1463 |
1411 |
3.42 |
|
|
|
| 19 |
A |
1429 |
1379 |
40.72 |
|
|
|
| 20 |
A |
1391 |
1343 |
4.80 |
|
|
|
| 21 |
A |
1367 |
1319 |
4.47 |
|
|
|
| 22 |
A |
1333 |
1286 |
14.44 |
|
|
|
| 23 |
A |
1304 |
1259 |
13.29 |
|
|
|
| 24 |
A |
1219 |
1176 |
71.38 |
|
|
|
| 25 |
A |
1148 |
1108 |
3.66 |
|
|
|
| 26 |
A |
1138 |
1098 |
23.20 |
|
|
|
| 27 |
A |
1053 |
1016 |
4.38 |
|
|
|
| 28 |
A |
1022 |
986 |
2.21 |
|
|
|
| 29 |
A |
982 |
947 |
14.05 |
|
|
|
| 30 |
A |
906 |
874 |
4.89 |
|
|
|
| 31 |
A |
880 |
849 |
5.20 |
|
|
|
| 32 |
A |
794 |
766 |
3.85 |
|
|
|
| 33 |
A |
767 |
740 |
0.93 |
|
|
|
| 34 |
A |
600 |
579 |
0.38 |
|
|
|
| 35 |
A |
520 |
502 |
14.11 |
|
|
|
| 36 |
A |
399 |
385 |
1.05 |
|
|
|
| 37 |
A |
323 |
311 |
0.27 |
|
|
|
| 38 |
A |
249 |
240 |
0.02 |
|
|
|
| 39 |
A |
190 |
184 |
0.21 |
|
|
|
| 40 |
A |
144 |
139 |
0.51 |
|
|
|
| 41 |
A |
91 |
88 |
0.86 |
|
|
|
| 42 |
A |
53 |
51 |
2.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31364.5 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30263.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.759 |
-0.292 |
-0.001 |
| C2 |
1.396 |
0.428 |
0.001 |
| C3 |
0.218 |
-0.560 |
0.001 |
| C4 |
-1.136 |
0.147 |
-0.000 |
| C5 |
-2.352 |
-0.773 |
-0.000 |
| O6 |
-1.229 |
1.378 |
-0.000 |
| H7 |
3.581 |
0.433 |
-0.000 |
| H8 |
2.865 |
-0.929 |
0.885 |
| H9 |
2.864 |
-0.926 |
-0.890 |
| H10 |
1.312 |
1.077 |
0.879 |
| H11 |
1.310 |
1.079 |
-0.876 |
| H12 |
0.266 |
-1.219 |
-0.879 |
| H13 |
0.265 |
-1.219 |
0.881 |
| H14 |
-3.264 |
-0.173 |
-0.000 |
| H15 |
-2.335 |
-1.421 |
-0.885 |
| H16 |
-2.335 |
-1.421 |
0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| C1 | | 1.5417 | 2.5559 | 3.9200 | 5.1339 | 4.3240 | 1.0955 | 1.0966 | 1.0966 | 2.1781 | 2.1782 | 2.8017 | 2.8034 | 6.0246 | 5.2921 | 5.2926 |
C2 | 1.5417 | | 1.5379 | 2.5475 | 3.9356 | 2.7917 | 2.1851 | 2.1868 | 2.1868 | 1.0956 | 1.0955 | 2.1829 | 2.1828 | 4.6986 | 4.2568 | 4.2565 | C3 | 2.5559 | 1.5379 | | 1.5274 | 2.5784 | 2.4187 | 3.5069 | 2.8160 | 2.8164 | 2.1567 | 2.1567 | 1.1003 | 1.1003 | 3.5032 | 2.8355 | 2.8350 | C4 | 3.9200 | 2.5475 | 1.5274 | | 1.5248 | 1.2341 | 4.7257 | 4.2372 | 4.2364 | 2.7625 | 2.7607 | 2.1459 | 2.1458 | 2.1520 | 2.1629 | 2.1631 | C5 | 5.1339 | 3.9356 | 2.5784 | 1.5248 | | 2.4261 | 6.0542 | 5.2945 | 5.2940 | 4.1978 | 4.1966 | 2.7972 | 2.7971 | 1.0916 | 1.0969 | 1.0969 | O6 | 4.3240 | 2.7917 | 2.4187 | 1.2341 | 2.4261 | | 4.9022 | 4.7825 | 4.7807 | 2.7061 | 2.7029 | 3.1228 | 3.1227 | 2.5585 | 3.1361 | 3.1366 | H7 | 1.0955 | 2.1851 | 3.5069 | 4.7257 | 6.0542 | 4.9022 | | 1.7747 | 1.7747 | 2.5174 | 2.5179 | 3.8069 | 3.8078 | 6.8718 | 6.2622 | 6.2624 | H8 | 1.0966 | 2.1868 | 2.8160 | 4.2372 | 5.2945 | 4.7825 | 1.7747 | | 1.7751 | 2.5371 | 3.0906 | 3.1555 | 2.6167 | 6.2392 | 5.5156 | 5.2237 | H9 | 1.0966 | 2.1868 | 2.8164 | 4.2364 | 5.2940 | 4.7807 | 1.7747 | 1.7751 | | 3.0905 | 2.5368 | 2.6153 | 3.1590 | 6.2383 | 5.2228 | 5.5161 | H10 | 2.1781 | 1.0956 | 2.1567 | 2.7625 | 4.1978 | 2.7061 | 2.5174 | 2.5371 | 3.0905 | | 1.7550 | 3.0758 | 2.5233 | 4.8247 | 4.7594 | 4.4205 | H11 | 2.1782 | 1.0955 | 2.1567 | 2.7607 | 4.1966 | 2.7029 | 2.5179 | 3.0906 | 2.5368 | 1.7550 | | 2.5250 | 3.0758 | 4.8231 | 4.4202 | 4.7581 | H12 | 2.8017 | 2.1829 | 1.1003 | 2.1459 | 2.7972 | 3.1228 | 3.8069 | 3.1555 | 2.6153 | 3.0758 | 2.5250 | | 1.7602 | 3.7850 | 2.6083 | 3.1488 | H13 | 2.8034 | 2.1828 | 1.1003 | 2.1458 | 2.7971 | 3.1227 | 3.8078 | 2.6167 | 3.1590 | 2.5233 | 3.0758 | 1.7602 | | 3.7847 | 3.1496 | 2.6078 | H14 | 6.0246 | 4.6986 | 3.5032 | 2.1520 | 1.0916 | 2.5585 | 6.8718 | 6.2392 | 6.2383 | 4.8247 | 4.8231 | 3.7850 | 3.7847 | | 1.7896 | 1.7898 | H15 | 5.2921 | 4.2568 | 2.8355 | 2.1629 | 1.0969 | 3.1361 | 6.2622 | 5.5156 | 5.2228 | 4.7594 | 4.4202 | 2.6083 | 3.1496 | 1.7896 | | 1.7700 | H16 | 5.2926 | 4.2565 | 2.8350 | 2.1631 | 1.0969 | 3.1366 | 6.2624 | 5.2237 | 5.5161 | 4.4205 | 4.7581 | 3.1488 | 2.6078 | 1.7898 | 1.7700 | |
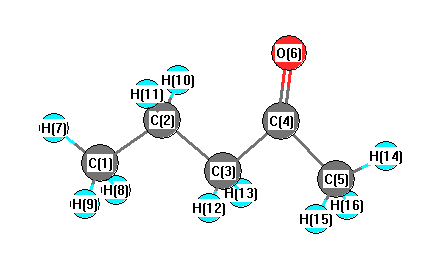 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.184 |
|
C1 |
C2 |
H10 |
110.205 |
| C1 |
C2 |
H11 |
110.213 |
|
C2 |
C1 |
H7 |
110.767 |
| C2 |
C1 |
H8 |
110.835 |
|
C2 |
C1 |
H9 |
110.833 |
| C2 |
C3 |
C4 |
112.416 |
|
C2 |
C3 |
H12 |
110.578 |
| C2 |
C3 |
H13 |
110.570 |
|
C3 |
C2 |
H10 |
108.803 |
| C3 |
C2 |
H11 |
108.805 |
|
C3 |
C4 |
C5 |
115.292 |
| C3 |
C4 |
O6 |
121.929 |
|
C4 |
C3 |
H12 |
108.406 |
| C4 |
C3 |
H13 |
108.399 |
|
C4 |
C5 |
H14 |
109.560 |
| C4 |
C5 |
H15 |
110.108 |
|
C4 |
C5 |
H16 |
110.120 |
| C5 |
C4 |
O6 |
122.779 |
|
H7 |
C1 |
H8 |
108.113 |
| H7 |
C1 |
H9 |
108.115 |
|
H8 |
C1 |
H9 |
108.066 |
| H10 |
C2 |
H11 |
106.448 |
|
H12 |
C3 |
H13 |
106.244 |
| H14 |
C5 |
H15 |
109.717 |
|
H14 |
C5 |
H16 |
109.733 |
| H15 |
C5 |
H16 |
107.576 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.549 |
|
|
-0.402 |
| 2 |
H |
0.188 |
|
|
0.110 |
| 3 |
H |
0.193 |
|
|
0.111 |
| 4 |
H |
0.192 |
|
|
0.096 |
| 5 |
C |
-0.379 |
|
|
0.228 |
| 6 |
H |
0.192 |
|
|
-0.018 |
| 7 |
H |
0.209 |
|
|
0.005 |
| 8 |
C |
-0.450 |
|
|
-0.435 |
| 9 |
H |
0.219 |
|
|
0.118 |
| 10 |
C |
-0.636 |
|
|
-0.641 |
| 11 |
H |
0.222 |
|
|
0.169 |
| 12 |
H |
0.216 |
|
|
0.172 |
| 13 |
C |
0.423 |
|
|
0.657 |
| 14 |
O |
-0.441 |
|
|
-0.455 |
| 15 |
H |
0.191 |
|
|
0.110 |
| 16 |
H |
0.210 |
|
|
0.174 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.465 |
2.229 |
0.499 |
2.714 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.461 |
2.234 |
0.506 |
2.717 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.120 |
-4.491 |
-1.071 |
| y |
-4.491 |
-40.341 |
-0.520 |
| z |
-1.071 |
-0.520 |
-37.519 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.191 |
-4.491 |
-1.071 |
| y |
-4.491 |
-1.521 |
-0.520 |
| z |
-1.071 |
-0.520 |
2.711 |
|
| Polar |
|---|
| 3z2-r2 | 5.423 |
|---|
| x2-y2 | 0.220 |
|---|
| xy | -4.491 |
|---|
| xz | -1.071 |
|---|
| yz | -0.520 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.854 |
0.090 |
0.187 |
| y |
0.090 |
7.669 |
0.161 |
| z |
0.187 |
0.161 |
6.430 |
<r2> (average value of r
2) Å
2
| <r2> |
229.083 |
| (<r2>)1/2 |
15.135 |
