Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -195.469981 |
| Energy at 298.15K | -195.480237 |
| Nuclear repulsion energy | 169.229274 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3228 |
3115 |
20.61 |
|
|
|
| 2 |
A |
3151 |
3040 |
4.30 |
|
|
|
| 3 |
A |
3138 |
3027 |
27.22 |
|
|
|
| 4 |
A |
3114 |
3005 |
53.98 |
|
|
|
| 5 |
A |
3111 |
3002 |
34.78 |
|
|
|
| 6 |
A |
3087 |
2979 |
3.68 |
|
|
|
| 7 |
A |
3069 |
2961 |
7.60 |
|
|
|
| 8 |
A |
3048 |
2941 |
27.96 |
|
|
|
| 9 |
A |
3045 |
2938 |
20.93 |
|
|
|
| 10 |
A |
3026 |
2920 |
15.59 |
|
|
|
| 11 |
A |
1718 |
1657 |
8.47 |
|
|
|
| 12 |
A |
1572 |
1517 |
6.84 |
|
|
|
| 13 |
A |
1567 |
1512 |
8.09 |
|
|
|
| 14 |
A |
1556 |
1501 |
2.06 |
|
|
|
| 15 |
A |
1540 |
1486 |
2.90 |
|
|
|
| 16 |
A |
1508 |
1455 |
1.50 |
|
|
|
| 17 |
A |
1461 |
1410 |
4.02 |
|
|
|
| 18 |
A |
1394 |
1346 |
1.36 |
|
|
|
| 19 |
A |
1372 |
1324 |
0.54 |
|
|
|
| 20 |
A |
1360 |
1312 |
0.03 |
|
|
|
| 21 |
A |
1342 |
1295 |
0.52 |
|
|
|
| 22 |
A |
1309 |
1263 |
0.14 |
|
|
|
| 23 |
A |
1223 |
1180 |
0.92 |
|
|
|
| 24 |
A |
1134 |
1095 |
2.98 |
|
|
|
| 25 |
A |
1071 |
1034 |
8.76 |
|
|
|
| 26 |
A |
1047 |
1011 |
9.14 |
|
|
|
| 27 |
A |
1026 |
990 |
0.60 |
|
|
|
| 28 |
A |
968 |
934 |
48.80 |
|
|
|
| 29 |
A |
948 |
915 |
0.85 |
|
|
|
| 30 |
A |
910 |
878 |
5.04 |
|
|
|
| 31 |
A |
895 |
864 |
5.65 |
|
|
|
| 32 |
A |
769 |
742 |
4.17 |
|
|
|
| 33 |
A |
650 |
627 |
10.53 |
|
|
|
| 34 |
A |
449 |
433 |
1.20 |
|
|
|
| 35 |
A |
387 |
373 |
0.18 |
|
|
|
| 36 |
A |
254 |
245 |
0.05 |
|
|
|
| 37 |
A |
239 |
231 |
0.16 |
|
|
|
| 38 |
A |
105 |
102 |
0.01 |
|
|
|
| 39 |
A |
92 |
89 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30441.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 29372.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.467 |
-0.222 |
-0.321 |
| C2 |
1.424 |
0.343 |
0.285 |
| C3 |
0.073 |
-0.309 |
0.467 |
| C4 |
-1.044 |
0.447 |
-0.302 |
| C5 |
-2.430 |
-0.195 |
-0.085 |
| H6 |
3.413 |
0.296 |
-0.442 |
| H7 |
2.413 |
-1.232 |
-0.719 |
| H8 |
1.510 |
1.359 |
0.671 |
| H9 |
0.112 |
-1.349 |
0.119 |
| H10 |
-0.190 |
-0.325 |
1.535 |
| H11 |
-1.065 |
1.493 |
0.031 |
| H12 |
-0.795 |
0.450 |
-1.370 |
| H13 |
-3.206 |
0.348 |
-0.636 |
| H14 |
-2.430 |
-1.236 |
-0.429 |
| H15 |
-2.696 |
-0.188 |
0.979 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3318 | 2.5214 | 3.5740 | 4.9026 | 1.0853 | 1.0868 | 2.0973 | 2.6475 | 3.2424 | 3.9417 | 3.4914 | 5.7098 | 5.0016 | 5.3239 |
C2 | 1.3318 | | 1.5112 | 2.5391 | 3.9093 | 2.1177 | 2.1137 | 1.0899 | 2.1478 | 2.1482 | 2.7534 | 2.7701 | 4.7208 | 4.2260 | 4.2119 | C3 | 2.5214 | 1.5112 | | 1.5526 | 2.5658 | 3.5135 | 2.7813 | 2.2107 | 1.0972 | 1.1000 | 2.1751 | 2.1687 | 3.5212 | 2.8157 | 2.8186 | C4 | 3.5740 | 2.5391 | 1.5526 | | 1.5433 | 4.4615 | 3.8662 | 2.8810 | 2.1769 | 2.1678 | 1.0978 | 1.0965 | 2.1898 | 2.1843 | 2.1849 | C5 | 4.9026 | 3.9093 | 2.5658 | 1.5433 | | 5.8744 | 4.9937 | 4.3025 | 2.7988 | 2.7673 | 2.1746 | 2.1778 | 1.0956 | 1.0963 | 1.0964 | H6 | 1.0853 | 2.1177 | 3.5135 | 4.4615 | 5.8744 | | 1.8467 | 2.4471 | 3.7305 | 4.1560 | 4.6588 | 4.3115 | 6.6217 | 6.0402 | 6.2904 | H7 | 1.0868 | 2.1137 | 2.7813 | 3.8662 | 4.9937 | 1.8467 | | 3.0758 | 2.4523 | 3.5609 | 4.4818 | 3.6808 | 5.8377 | 4.8519 | 5.4843 | H8 | 2.0973 | 1.0899 | 2.2107 | 2.8810 | 4.3025 | 2.4471 | 3.0758 | | 3.0969 | 2.5440 | 2.6563 | 3.2095 | 4.9969 | 4.8443 | 4.4920 | H9 | 2.6475 | 2.1478 | 1.0972 | 2.1769 | 2.7988 | 3.7305 | 2.4523 | 3.0969 | | 1.7728 | 3.0771 | 2.5054 | 3.8019 | 2.6026 | 3.1573 | H10 | 3.2424 | 2.1482 | 1.1000 | 2.1678 | 2.7673 | 4.1560 | 3.5609 | 2.5440 | 1.7728 | | 2.5163 | 3.0666 | 3.7762 | 3.1153 | 2.5705 | H11 | 3.9417 | 2.7534 | 2.1751 | 1.0978 | 2.1746 | 4.6588 | 4.4818 | 2.6563 | 3.0771 | 2.5163 | | 1.7670 | 2.5183 | 3.0862 | 2.5271 | H12 | 3.4914 | 2.7701 | 2.1687 | 1.0965 | 2.1778 | 4.3115 | 3.6808 | 3.2095 | 2.5054 | 3.0666 | 1.7670 | | 2.5224 | 2.5304 | 3.0885 | H13 | 5.7098 | 4.7208 | 3.5212 | 2.1898 | 1.0956 | 6.6217 | 5.8377 | 4.9969 | 3.8019 | 3.7762 | 2.5183 | 2.5224 | | 1.7758 | 1.7761 | H14 | 5.0016 | 4.2260 | 2.8157 | 2.1843 | 1.0963 | 6.0402 | 4.8519 | 4.8443 | 2.6026 | 3.1153 | 3.0862 | 2.5304 | 1.7758 | | 1.7754 | H15 | 5.3239 | 4.2119 | 2.8186 | 2.1849 | 1.0964 | 6.2904 | 5.4843 | 4.4920 | 3.1573 | 2.5705 | 2.5271 | 3.0885 | 1.7761 | 1.7754 | |
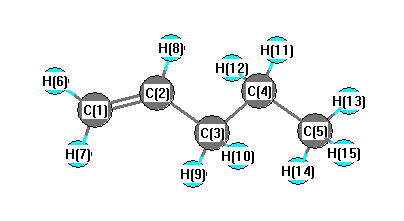 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.852 |
|
C1 |
C2 |
H8 |
119.681 |
| C2 |
C1 |
H6 |
122.025 |
|
C2 |
C1 |
H7 |
121.512 |
| C2 |
C3 |
C4 |
111.933 |
|
C2 |
C3 |
H9 |
109.838 |
| C2 |
C3 |
H10 |
109.702 |
|
C3 |
C2 |
H8 |
115.453 |
| C3 |
C4 |
C5 |
111.949 |
|
C3 |
C4 |
H11 |
109.100 |
| C3 |
C4 |
H12 |
108.678 |
|
C4 |
C3 |
H9 |
109.271 |
| C4 |
C3 |
H10 |
108.411 |
|
C4 |
C5 |
H13 |
111.028 |
| C4 |
C5 |
H14 |
110.542 |
|
C4 |
C5 |
H15 |
110.584 |
| C5 |
C4 |
H11 |
109.698 |
|
C5 |
C4 |
H12 |
110.023 |
| H6 |
C1 |
H7 |
116.463 |
|
H9 |
C3 |
H10 |
107.575 |
| H11 |
C4 |
H12 |
107.272 |
|
H13 |
C5 |
H14 |
108.222 |
| H13 |
C5 |
H15 |
108.243 |
|
H14 |
C5 |
H15 |
108.123 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.377 |
|
|
|
| 2 |
C |
-0.146 |
|
|
|
| 3 |
C |
-0.419 |
|
|
|
| 4 |
C |
-0.370 |
|
|
|
| 5 |
C |
-0.552 |
|
|
|
| 6 |
H |
0.181 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
| 8 |
H |
0.179 |
|
|
|
| 9 |
H |
0.190 |
|
|
|
| 10 |
H |
0.196 |
|
|
|
| 11 |
H |
0.186 |
|
|
|
| 12 |
H |
0.196 |
|
|
|
| 13 |
H |
0.188 |
|
|
|
| 14 |
H |
0.187 |
|
|
|
| 15 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.333 |
0.073 |
0.040 |
0.343 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.261 |
0.230 |
-0.704 |
| y |
0.230 |
-32.425 |
1.089 |
| z |
-0.704 |
1.089 |
-34.085 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.006 |
0.230 |
-0.704 |
| y |
0.230 |
1.248 |
1.089 |
| z |
-0.704 |
1.089 |
-1.242 |
|
| Polar |
|---|
| 3z2-r2 | -2.484 |
|---|
| x2-y2 | -0.836 |
|---|
| xy | 0.230 |
|---|
| xz | -0.704 |
|---|
| yz | 1.089 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.751 |
-0.319 |
-1.017 |
| y |
-0.319 |
6.744 |
0.700 |
| z |
-1.017 |
0.700 |
5.808 |
<r2> (average value of r
2) Å
2
| <r2> |
178.928 |
| (<r2>)1/2 |
13.376 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -195.470046 |
| Energy at 298.15K | -195.480390 |
| HF Energy | -195.470046 |
| Nuclear repulsion energy | 172.956045 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3228 |
3114 |
19.48 |
|
|
|
| 2 |
A |
3150 |
3040 |
4.39 |
|
|
|
| 3 |
A |
3134 |
3024 |
25.66 |
|
|
|
| 4 |
A |
3124 |
3015 |
31.57 |
|
|
|
| 5 |
A |
3109 |
3000 |
39.45 |
|
|
|
| 6 |
A |
3081 |
2973 |
19.15 |
|
|
|
| 7 |
A |
3071 |
2963 |
18.06 |
|
|
|
| 8 |
A |
3048 |
2941 |
17.98 |
|
|
|
| 9 |
A |
3043 |
2936 |
25.25 |
|
|
|
| 10 |
A |
3028 |
2922 |
21.60 |
|
|
|
| 11 |
A |
1716 |
1656 |
6.96 |
|
|
|
| 12 |
A |
1574 |
1519 |
4.58 |
|
|
|
| 13 |
A |
1565 |
1511 |
9.33 |
|
|
|
| 14 |
A |
1551 |
1497 |
2.21 |
|
|
|
| 15 |
A |
1539 |
1485 |
6.81 |
|
|
|
| 16 |
A |
1507 |
1454 |
2.16 |
|
|
|
| 17 |
A |
1460 |
1409 |
7.42 |
|
|
|
| 18 |
A |
1408 |
1359 |
2.43 |
|
|
|
| 19 |
A |
1383 |
1335 |
1.25 |
|
|
|
| 20 |
A |
1364 |
1316 |
0.67 |
|
|
|
| 21 |
A |
1324 |
1278 |
0.55 |
|
|
|
| 22 |
A |
1295 |
1250 |
1.90 |
|
|
|
| 23 |
A |
1218 |
1175 |
0.77 |
|
|
|
| 24 |
A |
1126 |
1087 |
4.66 |
|
|
|
| 25 |
A |
1073 |
1035 |
6.81 |
|
|
|
| 26 |
A |
1049 |
1012 |
8.15 |
|
|
|
| 27 |
A |
1037 |
1000 |
0.81 |
|
|
|
| 28 |
A |
969 |
935 |
55.40 |
|
|
|
| 29 |
A |
930 |
898 |
0.79 |
|
|
|
| 30 |
A |
913 |
881 |
1.04 |
|
|
|
| 31 |
A |
865 |
835 |
3.03 |
|
|
|
| 32 |
A |
789 |
761 |
5.43 |
|
|
|
| 33 |
A |
654 |
631 |
9.90 |
|
|
|
| 34 |
A |
442 |
426 |
0.46 |
|
|
|
| 35 |
A |
398 |
384 |
2.64 |
|
|
|
| 36 |
A |
306 |
295 |
0.14 |
|
|
|
| 37 |
A |
226 |
218 |
0.25 |
|
|
|
| 38 |
A |
126 |
121 |
0.02 |
|
|
|
| 39 |
A |
94 |
91 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30458.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 29389.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.073 |
0.585 |
0.112 |
| C2 |
-1.188 |
-0.273 |
-0.393 |
| C3 |
-0.063 |
-0.917 |
0.384 |
| C4 |
1.333 |
-0.522 |
-0.173 |
| C5 |
1.633 |
0.979 |
0.020 |
| H6 |
-2.859 |
1.024 |
-0.495 |
| H7 |
-2.047 |
0.879 |
1.158 |
| H8 |
-1.240 |
-0.545 |
-1.447 |
| H9 |
-0.166 |
-2.011 |
0.331 |
| H10 |
-0.128 |
-0.629 |
1.441 |
| H11 |
2.100 |
-1.121 |
0.333 |
| H12 |
1.376 |
-0.774 |
-1.241 |
| H13 |
2.607 |
1.243 |
-0.406 |
| H14 |
0.862 |
1.587 |
-0.463 |
| H15 |
1.644 |
1.230 |
1.088 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3321 | 2.5246 | 3.5929 | 3.7281 | 1.0854 | 1.0867 | 2.0978 | 3.2285 | 2.6504 | 4.5142 | 3.9462 | 4.7545 | 3.1543 | 3.8973 |
C2 | 1.3321 | | 1.5114 | 2.5426 | 3.1133 | 2.1179 | 2.1140 | 1.0902 | 2.1418 | 2.1476 | 3.4725 | 2.7464 | 4.0864 | 2.7687 | 3.5315 | C3 | 2.5246 | 1.5114 | | 1.5541 | 2.5693 | 3.5158 | 2.7864 | 2.2086 | 1.0997 | 1.0974 | 2.1733 | 2.1750 | 3.5239 | 2.8008 | 2.8322 | C4 | 3.5929 | 2.5426 | 1.5541 | | 1.5420 | 4.4798 | 3.8937 | 2.8716 | 2.1725 | 2.1796 | 1.0972 | 1.0981 | 2.1887 | 2.1800 | 2.1807 | C5 | 3.7281 | 3.1133 | 2.5693 | 1.5420 | | 4.5220 | 3.8533 | 3.5679 | 3.5028 | 2.7750 | 2.1732 | 2.1745 | 1.0956 | 1.0947 | 1.0966 | H6 | 1.0854 | 2.1179 | 3.5158 | 4.4798 | 4.5220 | | 1.8468 | 2.4472 | 4.1403 | 3.7334 | 5.4665 | 4.6610 | 5.4716 | 3.7640 | 4.7781 | H7 | 1.0867 | 2.1140 | 2.7864 | 3.8937 | 3.8533 | 1.8468 | | 3.0763 | 3.5459 | 2.4573 | 4.6779 | 4.4949 | 4.9234 | 3.4048 | 3.7091 | H8 | 2.0978 | 1.0902 | 2.2086 | 2.8716 | 3.5679 | 2.4472 | 3.0763 | | 2.5422 | 3.0959 | 3.8289 | 2.6341 | 4.3686 | 3.1518 | 4.2306 | H9 | 3.2285 | 2.1418 | 1.0997 | 2.1725 | 3.5028 | 4.1403 | 3.5459 | 2.5422 | | 1.7730 | 2.4353 | 2.5256 | 4.3383 | 3.8252 | 3.7889 | H10 | 2.6504 | 2.1476 | 1.0974 | 2.1796 | 2.7750 | 3.7334 | 2.4573 | 3.0959 | 1.7730 | | 2.5366 | 3.0780 | 3.7936 | 3.0845 | 2.5925 | H11 | 4.5142 | 3.4725 | 2.1733 | 1.0972 | 2.1732 | 5.4665 | 4.6779 | 3.8289 | 2.4353 | 2.5366 | | 1.7673 | 2.5275 | 3.0820 | 2.5108 | H12 | 3.9462 | 2.7464 | 2.1750 | 1.0981 | 2.1745 | 4.6610 | 4.4949 | 2.6341 | 2.5256 | 3.0780 | 1.7673 | | 2.5064 | 2.5384 | 3.0841 | H13 | 4.7545 | 4.0864 | 3.5239 | 2.1887 | 1.0956 | 5.4716 | 4.9234 | 4.3686 | 4.3383 | 3.7936 | 2.5275 | 2.5064 | | 1.7796 | 1.7767 | H14 | 3.1543 | 2.7687 | 2.8008 | 2.1800 | 1.0947 | 3.7640 | 3.4048 | 3.1518 | 3.8252 | 3.0845 | 3.0820 | 2.5384 | 1.7796 | | 1.7732 | H15 | 3.8973 | 3.5315 | 2.8322 | 2.1807 | 1.0966 | 4.7781 | 3.7091 | 4.2306 | 3.7889 | 2.5925 | 2.5108 | 3.0841 | 1.7767 | 1.7732 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.089 |
|
C1 |
C2 |
H8 |
119.671 |
| C2 |
C1 |
H6 |
122.008 |
|
C2 |
C1 |
H7 |
121.522 |
| C2 |
C3 |
C4 |
112.072 |
|
C2 |
C3 |
H9 |
109.210 |
| C2 |
C3 |
H10 |
109.791 |
|
C3 |
C2 |
H8 |
115.236 |
| C3 |
C4 |
C5 |
112.167 |
|
C3 |
C4 |
H11 |
108.889 |
| C3 |
C4 |
H12 |
108.967 |
|
C4 |
C3 |
H9 |
108.690 |
| C4 |
C3 |
H10 |
109.362 |
|
C4 |
C5 |
H13 |
111.031 |
| C4 |
C5 |
H14 |
110.394 |
|
C4 |
C5 |
H15 |
110.336 |
| C5 |
C4 |
H11 |
109.707 |
|
C5 |
C4 |
H12 |
109.758 |
| H6 |
C1 |
H7 |
116.470 |
|
H9 |
C3 |
H10 |
107.605 |
| H11 |
C4 |
H12 |
107.221 |
|
H13 |
C5 |
H14 |
108.681 |
| H13 |
C5 |
H15 |
108.280 |
|
H14 |
C5 |
H15 |
108.033 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.377 |
|
|
|
| 2 |
C |
-0.143 |
|
|
|
| 3 |
C |
-0.421 |
|
|
|
| 4 |
C |
-0.375 |
|
|
|
| 5 |
C |
-0.549 |
|
|
|
| 6 |
H |
0.181 |
|
|
|
| 7 |
H |
0.178 |
|
|
|
| 8 |
H |
0.179 |
|
|
|
| 9 |
H |
0.198 |
|
|
|
| 10 |
H |
0.190 |
|
|
|
| 11 |
H |
0.188 |
|
|
|
| 12 |
H |
0.185 |
|
|
|
| 13 |
H |
0.183 |
|
|
|
| 14 |
H |
0.200 |
|
|
|
| 15 |
H |
0.182 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.210 |
-0.169 |
-0.051 |
0.274 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.723 |
-1.298 |
0.422 |
| y |
-1.298 |
-33.735 |
0.528 |
| z |
0.422 |
0.528 |
-32.115 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.798 |
-1.298 |
0.422 |
| y |
-1.298 |
-0.816 |
0.528 |
| z |
0.422 |
0.528 |
1.614 |
|
| Polar |
|---|
| 3z2-r2 | 3.228 |
|---|
| x2-y2 | 0.013 |
|---|
| xy | -1.298 |
|---|
| xz | 0.422 |
|---|
| yz | 0.528 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.406 |
-1.460 |
-0.126 |
| y |
-1.460 |
6.824 |
0.516 |
| z |
-0.126 |
0.516 |
6.752 |
<r2> (average value of r
2) Å
2
| <r2> |
152.228 |
| (<r2>)1/2 |
12.338 |
