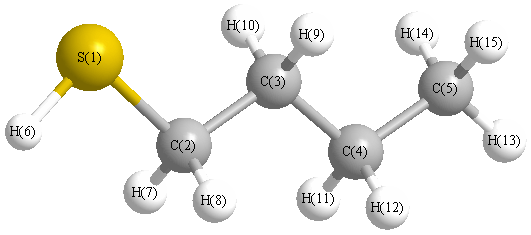
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3034 |
3017 |
37.94 |
|
|
|
| 2 |
A' |
3017 |
3000 |
22.77 |
|
|
|
| 3 |
A' |
2980 |
2964 |
24.42 |
|
|
|
| 4 |
A' |
2971 |
2955 |
19.91 |
|
|
|
| 5 |
A' |
2962 |
2946 |
12.53 |
|
|
|
| 6 |
A' |
2361 |
2348 |
50.05 |
|
|
|
| 7 |
A' |
1538 |
1529 |
6.14 |
|
|
|
| 8 |
A' |
1524 |
1516 |
1.76 |
|
|
|
| 9 |
A' |
1513 |
1505 |
0.82 |
|
|
|
| 10 |
A' |
1497 |
1489 |
2.52 |
|
|
|
| 11 |
A' |
1424 |
1416 |
1.99 |
|
|
|
| 12 |
A' |
1366 |
1358 |
3.83 |
|
|
|
| 13 |
A' |
1335 |
1328 |
7.94 |
|
|
|
| 14 |
A' |
1244 |
1237 |
35.26 |
|
|
|
| 15 |
A' |
1108 |
1102 |
5.18 |
|
|
|
| 16 |
A' |
1011 |
1005 |
3.62 |
|
|
|
| 17 |
A' |
975 |
970 |
1.74 |
|
|
|
| 18 |
A' |
891 |
887 |
1.74 |
|
|
|
| 19 |
A' |
792 |
787 |
1.99 |
|
|
|
| 20 |
A' |
653 |
650 |
9.88 |
|
|
|
| 21 |
A' |
381 |
379 |
1.48 |
|
|
|
| 22 |
A' |
301 |
300 |
2.27 |
|
|
|
| 23 |
A' |
149 |
148 |
2.07 |
|
|
|
| 24 |
A" |
3079 |
3062 |
26.51 |
|
|
|
| 25 |
A" |
3037 |
3020 |
54.94 |
|
|
|
| 26 |
A" |
3017 |
3000 |
3.27 |
|
|
|
| 27 |
A" |
2986 |
2970 |
2.95 |
|
|
|
| 28 |
A" |
1531 |
1523 |
7.45 |
|
|
|
| 29 |
A" |
1343 |
1336 |
0.04 |
|
|
|
| 30 |
A" |
1317 |
1310 |
0.58 |
|
|
|
| 31 |
A" |
1230 |
1223 |
0.67 |
|
|
|
| 32 |
A" |
1072 |
1066 |
0.09 |
|
|
|
| 33 |
A" |
925 |
919 |
2.38 |
|
|
|
| 34 |
A" |
790 |
786 |
0.02 |
|
|
|
| 35 |
A" |
744 |
740 |
6.78 |
|
|
|
| 36 |
A" |
237 |
236 |
|
|
|
|
| 37 |
A" |
159 |
158 |
18.95 |
|
|
|
| 38 |
A" |
111 |
111 |
4.93 |
|
|
|
| 39 |
A" |
97 |
97 |
3.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28351.4 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 28195.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.014 |
|
|
|
| 2 |
C |
-0.545 |
|
|
|
| 3 |
C |
-0.337 |
|
|
|
| 4 |
C |
-0.345 |
|
|
|
| 5 |
C |
-0.517 |
|
|
|
| 6 |
H |
0.064 |
|
|
|
| 7 |
H |
0.212 |
|
|
|
| 8 |
H |
0.212 |
|
|
|
| 9 |
H |
0.193 |
|
|
|
| 10 |
H |
0.193 |
|
|
|
| 11 |
H |
0.175 |
|
|
|
| 12 |
H |
0.175 |
|
|
|
| 13 |
H |
0.177 |
|
|
|
| 14 |
H |
0.178 |
|
|
|
| 15 |
H |
0.178 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.074 |
0.367 |
0.000 |
2.106 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.400 |
2.736 |
0.000 |
| y |
2.736 |
-38.168 |
0.000 |
| z |
0.000 |
0.000 |
-42.167 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.233 |
2.736 |
0.000 |
| y |
2.736 |
5.116 |
0.000 |
| z |
0.000 |
0.000 |
-0.883 |
|
| Polar |
|---|
| 3z2-r2 | -1.767 |
|---|
| x2-y2 | -6.232 |
|---|
| xy | 2.736 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.686 |
-1.624 |
0.000 |
| y |
-1.624 |
10.777 |
0.000 |
| z |
0.000 |
0.000 |
6.408 |
<r2> (average value of r
2) Å
2
| <r2> |
274.624 |
| (<r2>)1/2 |
16.572 |