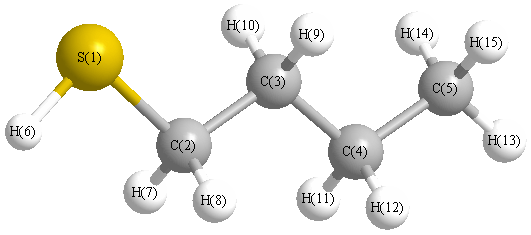
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3115 |
3005 |
33.79 |
|
|
|
| 2 |
A' |
3098 |
2989 |
20.35 |
|
|
|
| 3 |
A' |
3059 |
2951 |
21.47 |
|
|
|
| 4 |
A' |
3048 |
2941 |
19.13 |
|
|
|
| 5 |
A' |
3042 |
2935 |
11.60 |
|
|
|
| 6 |
A' |
2459 |
2373 |
43.63 |
|
|
|
| 7 |
A' |
1573 |
1517 |
8.24 |
|
|
|
| 8 |
A' |
1560 |
1505 |
2.81 |
|
|
|
| 9 |
A' |
1550 |
1495 |
0.75 |
|
|
|
| 10 |
A' |
1534 |
1480 |
3.49 |
|
|
|
| 11 |
A' |
1463 |
1412 |
3.55 |
|
|
|
| 12 |
A' |
1408 |
1358 |
3.92 |
|
|
|
| 13 |
A' |
1375 |
1326 |
10.76 |
|
|
|
| 14 |
A' |
1287 |
1242 |
29.86 |
|
|
|
| 15 |
A' |
1144 |
1104 |
6.04 |
|
|
|
| 16 |
A' |
1052 |
1015 |
2.74 |
|
|
|
| 17 |
A' |
1013 |
978 |
1.68 |
|
|
|
| 18 |
A' |
924 |
891 |
2.30 |
|
|
|
| 19 |
A' |
824 |
795 |
2.90 |
|
|
|
| 20 |
A' |
690 |
665 |
8.97 |
|
|
|
| 21 |
A' |
394 |
380 |
1.46 |
|
|
|
| 22 |
A' |
315 |
304 |
2.13 |
|
|
|
| 23 |
A' |
154 |
148 |
2.14 |
|
|
|
| 24 |
A" |
3161 |
3050 |
20.73 |
|
|
|
| 25 |
A" |
3117 |
3007 |
50.88 |
|
|
|
| 26 |
A" |
3097 |
2989 |
2.42 |
|
|
|
| 27 |
A" |
3068 |
2961 |
2.49 |
|
|
|
| 28 |
A" |
1567 |
1512 |
8.74 |
|
|
|
| 29 |
A" |
1378 |
1330 |
0.05 |
|
|
|
| 30 |
A" |
1353 |
1306 |
0.59 |
|
|
|
| 31 |
A" |
1265 |
1221 |
0.69 |
|
|
|
| 32 |
A" |
1106 |
1068 |
0.08 |
|
|
|
| 33 |
A" |
952 |
918 |
2.41 |
|
|
|
| 34 |
A" |
812 |
783 |
0.01 |
|
|
|
| 35 |
A" |
761 |
735 |
7.12 |
|
|
|
| 36 |
A" |
243 |
235 |
0.00 |
|
|
|
| 37 |
A" |
175 |
169 |
23.84 |
|
|
|
| 38 |
A" |
116 |
112 |
4.02 |
|
|
|
| 39 |
A" |
100 |
96 |
3.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29174.2 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 28150.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.000 |
|
|
|
| 2 |
C |
-0.592 |
|
|
|
| 3 |
C |
-0.367 |
|
|
|
| 4 |
C |
-0.376 |
|
|
|
| 5 |
C |
-0.551 |
|
|
|
| 6 |
H |
0.069 |
|
|
|
| 7 |
H |
0.227 |
|
|
|
| 8 |
H |
0.227 |
|
|
|
| 9 |
H |
0.208 |
|
|
|
| 10 |
H |
0.208 |
|
|
|
| 11 |
H |
0.189 |
|
|
|
| 12 |
H |
0.189 |
|
|
|
| 13 |
H |
0.190 |
|
|
|
| 14 |
H |
0.190 |
|
|
|
| 15 |
H |
0.190 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.106 |
0.304 |
0.000 |
2.128 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.517 |
2.711 |
0.000 |
| y |
2.711 |
-38.005 |
0.000 |
| z |
0.000 |
0.000 |
-42.088 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.471 |
2.711 |
0.000 |
| y |
2.711 |
5.298 |
0.000 |
| z |
0.000 |
0.000 |
-0.827 |
|
| Polar |
|---|
| 3z2-r2 | -1.654 |
|---|
| x2-y2 | -6.513 |
|---|
| xy | 2.711 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.473 |
-1.520 |
0.000 |
| y |
-1.520 |
10.369 |
0.000 |
| z |
0.000 |
0.000 |
6.312 |
<r2> (average value of r
2) Å
2
| <r2> |
269.938 |
| (<r2>)1/2 |
16.430 |