return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: MP4=FULL/3-21G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4=FULL/3-21G
 hartrees
Energy at 0K-211.688483
Energy at 298.15K-211.701153
HF Energy-211.134369
Nuclear repulsion energy185.519044
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/3-21G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3336 3265        
2 A' 3120 3054        
3 A' 3094 3028        
4 A' 3049 2985        
5 A' 3030 2965        
6 A' 2929 2867        
7 A' 1600 1566        
8 A' 1582 1548        
9 A' 1571 1538        
10 A' 1467 1435        
11 A' 1439 1408        
12 A' 1349 1320        
13 A' 1272 1245        
14 A' 1172 1147        
15 A' 1012 991        
16 A' 927 907        
17 A' 852 834        
18 A' 731 715        
19 A' 417 408        
20 A' 263 257        
21 A' 182 179        
22 A' 101 99        
23 A" 3120 3054        
24 A" 3093 3028        
25 A" 3047 2982        
26 A" 3029 2965        
27 A" 2925 2863        
28 A" 1591 1557        
29 A" 1580 1546        
30 A" 1574 1540        
31 A" 1514 1482        
32 A" 1464 1433        
33 A" 1387 1358        
34 A" 1329 1301        
35 A" 1145 1121        
36 A" 1114 1091        
37 A" 1068 1045        
38 A" 919 899        
39 A" 841 823        
40 A" 417 408        
41 A" 260 255        
42 A" 118 115        

Unscaled Zero Point Vibrational Energy (zpe) 33012.0 cm-1
Scaled (by 0.9788) Zero Point Vibrational Energy (zpe) 32312.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4=FULL/3-21G
ABC
0.56362 0.06816 0.06406

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4=FULL/3-21G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.017 -0.288 0.000
C2 0.017 0.543 1.248
C3 0.017 0.543 -1.248
C4 0.017 -0.389 2.486
C5 0.017 -0.389 -2.486
H6 -0.819 -0.894 0.000
H7 -0.838 1.249 1.303
H8 0.939 1.144 1.236
H9 -0.838 1.249 -1.303
H10 0.939 1.144 -1.236
H11 0.092 0.193 3.416
H12 -0.911 -0.979 2.520
H13 0.868 -1.078 2.415
H14 0.092 0.193 -3.416
H15 -0.911 -0.979 -2.520
H16 0.868 -1.078 -2.415

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.50001.50002.48802.48801.03252.18902.10452.18902.10453.45012.77282.67923.45012.77282.6792
C21.50002.49661.54953.84892.07921.10981.10042.78182.71682.19672.19012.17104.67764.16874.0952
C31.50002.49663.84891.54952.07922.78182.71681.10981.10044.67764.16874.09522.19672.19012.1710
C42.48801.54953.84894.97202.67102.19372.18264.21564.12931.09921.10051.09725.93065.12525.0215
C52.48803.84891.54954.97202.67104.21564.12932.19372.18265.93065.12525.02151.09921.10051.0972
H61.03252.07922.07922.67102.67102.50792.96172.50792.96173.69832.52292.95153.69832.52292.9515
H72.18901.10982.78182.19374.21562.50791.78112.60633.10062.53822.53973.09184.92414.42554.7060
H82.10451.10042.71682.18264.12932.96171.78113.10062.47142.52453.09532.51664.82244.69424.2742
H92.18902.78181.10984.21562.19372.50792.60633.10061.78114.92414.42554.70602.53822.53973.0918
H102.10452.71681.10044.12932.18262.96173.10062.47141.78114.82244.69424.27422.52453.09532.5166
H113.45012.19674.67761.09925.93063.69832.53822.52454.92414.82241.78421.79396.83106.13266.0173
H122.77282.19014.16871.10055.12522.52292.53973.09534.42554.69421.78421.78486.13265.03955.2463
H132.67922.17104.09521.09725.02152.95153.09182.51664.70604.27421.79391.78486.01735.24634.8294
H143.45014.67762.19675.93061.09923.69834.92414.82242.53822.52456.83106.13266.01731.78421.7939
H152.77284.16872.19015.12521.10052.52294.42554.69422.53973.09536.13265.03955.24631.78421.7848
H162.67924.09522.17105.02151.09722.95154.70604.27423.09182.51666.01735.24634.82941.79391.7848

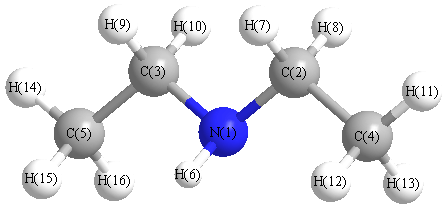
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 109.337 N1 C2 H7 113.176
N1 C2 H8 107.053 N1 C3 C5 109.337
N1 C3 H9 113.176 N1 C3 H10 107.053
C2 N1 C3 112.654 C2 N1 H6 108.977
C2 C4 H11 110.916 C2 C4 H12 110.324
C2 C4 H13 109.023 C3 N1 H6 108.977
C3 C5 H14 110.916 C3 C5 H15 110.324
C3 C5 H16 109.023 C4 C2 H7 110.057
C4 C2 H8 109.738 C5 C3 H8 149.698
C5 C3 H10 109.738 H7 C2 H8 107.385
H9 C3 H10 107.385 H11 C4 H12 108.410
H11 C4 H13 109.522 H12 C4 H13 108.608
H14 C5 H15 108.410 H14 C5 H16 109.522
H15 C5 H16 108.608
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability