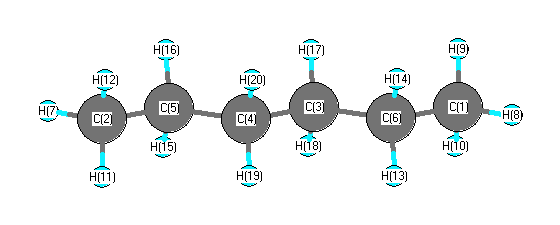
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3110 |
3000 |
0.00 |
|
|
|
| 2 |
Ag |
3045 |
2938 |
0.00 |
|
|
|
| 3 |
Ag |
3041 |
2934 |
0.00 |
|
|
|
| 4 |
Ag |
3025 |
2919 |
0.00 |
|
|
|
| 5 |
Ag |
1572 |
1517 |
0.00 |
|
|
|
| 6 |
Ag |
1554 |
1499 |
0.00 |
|
|
|
| 7 |
Ag |
1545 |
1491 |
0.00 |
|
|
|
| 8 |
Ag |
1461 |
1409 |
0.00 |
|
|
|
| 9 |
Ag |
1401 |
1352 |
0.00 |
|
|
|
| 10 |
Ag |
1376 |
1328 |
0.00 |
|
|
|
| 11 |
Ag |
1173 |
1132 |
0.00 |
|
|
|
| 12 |
Ag |
1055 |
1018 |
0.00 |
|
|
|
| 13 |
Ag |
990 |
955 |
0.00 |
|
|
|
| 14 |
Ag |
919 |
887 |
0.00 |
|
|
|
| 15 |
Ag |
367 |
354 |
0.00 |
|
|
|
| 16 |
Ag |
299 |
289 |
0.00 |
|
|
|
| 17 |
Au |
3113 |
3004 |
142.79 |
|
|
|
| 18 |
Au |
3088 |
2980 |
22.42 |
|
|
|
| 19 |
Au |
3058 |
2951 |
0.07 |
|
|
|
| 20 |
Au |
1568 |
1513 |
16.08 |
|
|
|
| 21 |
Au |
1376 |
1328 |
0.63 |
|
|
|
| 22 |
Au |
1293 |
1248 |
0.10 |
|
|
|
| 23 |
Au |
1062 |
1025 |
1.11 |
|
|
|
| 24 |
Au |
835 |
806 |
2.41 |
|
|
|
| 25 |
Au |
748 |
722 |
7.78 |
|
|
|
| 26 |
Au |
256 |
247 |
0.00 |
|
|
|
| 27 |
Au |
105 |
101 |
0.00 |
|
|
|
| 28 |
Au |
72 |
70 |
0.00 |
|
|
|
| 29 |
Bg |
3110 |
3001 |
0.00 |
|
|
|
| 30 |
Bg |
3074 |
2966 |
0.00 |
|
|
|
| 31 |
Bg |
3047 |
2940 |
0.00 |
|
|
|
| 32 |
Bg |
1568 |
1513 |
0.00 |
|
|
|
| 33 |
Bg |
1372 |
1324 |
0.00 |
|
|
|
| 34 |
Bg |
1350 |
1302 |
0.00 |
|
|
|
| 35 |
Bg |
1236 |
1192 |
0.00 |
|
|
|
| 36 |
Bg |
944 |
911 |
0.00 |
|
|
|
| 37 |
Bg |
767 |
740 |
0.00 |
|
|
|
| 38 |
Bg |
245 |
236 |
0.00 |
|
|
|
| 39 |
Bg |
149 |
144 |
0.00 |
|
|
|
| 40 |
Bu |
3109 |
3000 |
75.84 |
|
|
|
| 41 |
Bu |
3046 |
2939 |
118.51 |
|
|
|
| 42 |
Bu |
3043 |
2936 |
25.44 |
|
|
|
| 43 |
Bu |
3029 |
2923 |
4.34 |
|
|
|
| 44 |
Bu |
1575 |
1519 |
12.57 |
|
|
|
| 45 |
Bu |
1563 |
1508 |
2.00 |
|
|
|
| 46 |
Bu |
1545 |
1491 |
0.29 |
|
|
|
| 47 |
Bu |
1460 |
1408 |
8.34 |
|
|
|
| 48 |
Bu |
1404 |
1355 |
3.02 |
|
|
|
| 49 |
Bu |
1319 |
1272 |
0.11 |
|
|
|
| 50 |
Bu |
1098 |
1059 |
14.26 |
|
|
|
| 51 |
Bu |
1036 |
1000 |
0.13 |
|
|
|
| 52 |
Bu |
883 |
852 |
3.35 |
|
|
|
| 53 |
Bu |
473 |
456 |
0.16 |
|
|
|
| 54 |
Bu |
135 |
130 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42041.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 40565.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.551 |
|
|
-0.348 |
| 2 |
C |
-0.551 |
|
|
-0.366 |
| 3 |
C |
-0.356 |
|
|
-0.112 |
| 4 |
C |
-0.356 |
|
|
-0.155 |
| 5 |
C |
-0.377 |
|
|
0.155 |
| 6 |
C |
-0.377 |
|
|
0.110 |
| 7 |
H |
0.185 |
|
|
0.090 |
| 8 |
H |
0.185 |
|
|
0.089 |
| 9 |
H |
0.185 |
|
|
0.089 |
| 10 |
H |
0.185 |
|
|
0.089 |
| 11 |
H |
0.185 |
|
|
0.092 |
| 12 |
H |
0.185 |
|
|
0.092 |
| 13 |
H |
0.184 |
|
|
0.003 |
| 14 |
H |
0.184 |
|
|
0.003 |
| 15 |
H |
0.184 |
|
|
-0.008 |
| 16 |
H |
0.184 |
|
|
-0.008 |
| 17 |
H |
0.181 |
|
|
0.043 |
| 18 |
H |
0.181 |
|
|
0.043 |
| 19 |
H |
0.181 |
|
|
0.050 |
| 20 |
H |
0.181 |
|
|
0.050 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.006 |
0.002 |
0.000 |
0.006 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.681 |
-0.047 |
0.000 |
| y |
-0.047 |
-42.316 |
0.000 |
| z |
0.000 |
0.000 |
-41.011 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.018 |
-0.047 |
0.000 |
| y |
-0.047 |
-0.970 |
0.000 |
| z |
0.000 |
0.000 |
0.987 |
|
| Polar |
|---|
| 3z2-r2 | 1.975 |
|---|
| x2-y2 | 0.635 |
|---|
| xy | -0.047 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.062 |
1.151 |
0.000 |
| y |
1.151 |
10.379 |
0.000 |
| z |
0.000 |
0.000 |
8.246 |
<r2> (average value of r
2) Å
2
| <r2> |
321.418 |
| (<r2>)1/2 |
17.928 |