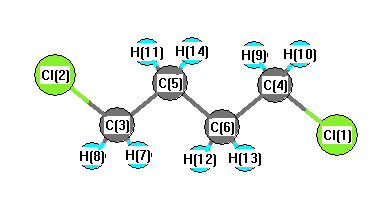
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3131 |
3021 |
0.00 |
|
|
|
| 2 |
Ag |
3070 |
2962 |
0.00 |
|
|
|
| 3 |
Ag |
1550 |
1495 |
0.00 |
|
|
|
| 4 |
Ag |
1535 |
1481 |
0.00 |
|
|
|
| 5 |
Ag |
1401 |
1352 |
0.00 |
|
|
|
| 6 |
Ag |
1298 |
1252 |
0.00 |
|
|
|
| 7 |
Ag |
1071 |
1034 |
0.00 |
|
|
|
| 8 |
Ag |
1000 |
965 |
0.00 |
|
|
|
| 9 |
Ag |
689 |
665 |
0.00 |
|
|
|
| 10 |
Ag |
328 |
316 |
0.00 |
|
|
|
| 11 |
Ag |
210 |
202 |
0.00 |
|
|
|
| 12 |
Au |
3204 |
3092 |
12.06 |
|
|
|
| 13 |
Au |
3128 |
3018 |
13.08 |
|
|
|
| 14 |
Au |
1359 |
1312 |
1.92 |
|
|
|
| 15 |
Au |
1142 |
1102 |
0.74 |
|
|
|
| 16 |
Au |
908 |
876 |
1.14 |
|
|
|
| 17 |
Au |
768 |
741 |
7.98 |
|
|
|
| 18 |
Au |
110 |
106 |
2.04 |
|
|
|
| 19 |
Au |
53 |
52 |
7.05 |
|
|
|
| 20 |
Bg |
3202 |
3090 |
0.00 |
|
|
|
| 21 |
Bg |
3105 |
2996 |
0.00 |
|
|
|
| 22 |
Bg |
1373 |
1324 |
0.00 |
|
|
|
| 23 |
Bg |
1281 |
1236 |
0.00 |
|
|
|
| 24 |
Bg |
1060 |
1023 |
0.00 |
|
|
|
| 25 |
Bg |
798 |
770 |
0.00 |
|
|
|
| 26 |
Bg |
146 |
141 |
0.00 |
|
|
|
| 27 |
Bu |
3132 |
3022 |
19.53 |
|
|
|
| 28 |
Bu |
3079 |
2971 |
12.33 |
|
|
|
| 29 |
Bu |
1563 |
1508 |
13.91 |
|
|
|
| 30 |
Bu |
1532 |
1479 |
3.45 |
|
|
|
| 31 |
Bu |
1370 |
1322 |
61.06 |
|
|
|
| 32 |
Bu |
1265 |
1221 |
20.11 |
|
|
|
| 33 |
Bu |
1017 |
981 |
29.10 |
|
|
|
| 34 |
Bu |
671 |
647 |
82.31 |
|
|
|
| 35 |
Bu |
399 |
385 |
27.63 |
|
|
|
| 36 |
Bu |
99 |
96 |
6.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25523.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 24627.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Cl |
-0.066 |
|
|
-0.217 |
| 2 |
Cl |
-0.066 |
|
|
-0.216 |
| 3 |
C |
-0.511 |
|
|
-0.218 |
| 4 |
C |
-0.511 |
|
|
-0.218 |
| 5 |
C |
-0.379 |
|
|
0.027 |
| 6 |
C |
-0.379 |
|
|
0.044 |
| 7 |
H |
0.255 |
|
|
0.156 |
| 8 |
H |
0.255 |
|
|
0.156 |
| 9 |
H |
0.255 |
|
|
0.155 |
| 10 |
H |
0.255 |
|
|
0.155 |
| 11 |
H |
0.223 |
|
|
0.046 |
| 12 |
H |
0.223 |
|
|
0.042 |
| 13 |
H |
0.223 |
|
|
0.042 |
| 14 |
H |
0.223 |
|
|
0.046 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.002 |
-0.003 |
0.000 |
0.004 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.336 |
7.561 |
0.000 |
| y |
7.561 |
-73.675 |
0.000 |
| z |
0.000 |
0.000 |
-50.708 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.855 |
7.561 |
0.000 |
| y |
7.561 |
-22.653 |
0.000 |
| z |
0.000 |
0.000 |
11.797 |
|
| Polar |
|---|
| 3z2-r2 | 23.595 |
|---|
| x2-y2 | 22.339 |
|---|
| xy | 7.561 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.979 |
-1.721 |
0.000 |
| y |
-1.721 |
13.498 |
0.000 |
| z |
0.000 |
0.000 |
6.022 |
<r2> (average value of r
2) Å
2
| <r2> |
539.426 |
| (<r2>)1/2 |
23.226 |