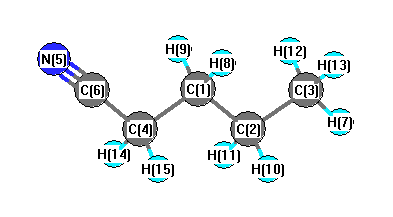
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3117 |
3007 |
29.92 |
|
|
|
| 2 |
A' |
3067 |
2960 |
29.54 |
|
|
|
| 3 |
A' |
3053 |
2946 |
0.86 |
|
|
|
| 4 |
A' |
3049 |
2942 |
18.07 |
|
|
|
| 5 |
A' |
3039 |
2932 |
14.04 |
|
|
|
| 6 |
A' |
2350 |
2267 |
4.64 |
|
|
|
| 7 |
A' |
1572 |
1517 |
8.74 |
|
|
|
| 8 |
A' |
1558 |
1503 |
3.92 |
|
|
|
| 9 |
A' |
1547 |
1493 |
0.59 |
|
|
|
| 10 |
A' |
1532 |
1478 |
4.63 |
|
|
|
| 11 |
A' |
1464 |
1412 |
5.37 |
|
|
|
| 12 |
A' |
1404 |
1355 |
0.38 |
|
|
|
| 13 |
A' |
1388 |
1340 |
3.91 |
|
|
|
| 14 |
A' |
1327 |
1280 |
0.94 |
|
|
|
| 15 |
A' |
1142 |
1102 |
8.80 |
|
|
|
| 16 |
A' |
1039 |
1003 |
0.19 |
|
|
|
| 17 |
A' |
1001 |
966 |
0.27 |
|
|
|
| 18 |
A' |
949 |
915 |
0.11 |
|
|
|
| 19 |
A' |
902 |
870 |
3.73 |
|
|
|
| 20 |
A' |
588 |
567 |
2.26 |
|
|
|
| 21 |
A' |
386 |
373 |
0.75 |
|
|
|
| 22 |
A' |
302 |
291 |
0.83 |
|
|
|
| 23 |
A' |
135 |
130 |
3.97 |
|
|
|
| 24 |
A" |
3122 |
3012 |
61.98 |
|
|
|
| 25 |
A" |
3109 |
3000 |
1.25 |
|
|
|
| 26 |
A" |
3085 |
2977 |
0.01 |
|
|
|
| 27 |
A" |
3066 |
2959 |
4.33 |
|
|
|
| 28 |
A" |
1567 |
1512 |
9.37 |
|
|
|
| 29 |
A" |
1372 |
1324 |
0.12 |
|
|
|
| 30 |
A" |
1362 |
1314 |
0.45 |
|
|
|
| 31 |
A" |
1289 |
1243 |
0.08 |
|
|
|
| 32 |
A" |
1183 |
1141 |
0.16 |
|
|
|
| 33 |
A" |
973 |
939 |
1.25 |
|
|
|
| 34 |
A" |
822 |
793 |
0.16 |
|
|
|
| 35 |
A" |
753 |
727 |
6.79 |
|
|
|
| 36 |
A" |
459 |
443 |
0.03 |
|
|
|
| 37 |
A" |
243 |
235 |
0.07 |
|
|
|
| 38 |
A" |
117 |
112 |
0.54 |
|
|
|
| 39 |
A" |
89 |
86 |
2.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29259.9 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 28232.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.348 |
|
|
0.093 |
| 2 |
C |
-0.383 |
|
|
0.053 |
| 3 |
C |
-0.553 |
|
|
-0.438 |
| 4 |
C |
-0.445 |
|
|
-0.507 |
| 5 |
N |
-0.485 |
|
|
-0.466 |
| 6 |
C |
0.345 |
|
|
0.465 |
| 7 |
H |
0.193 |
|
|
0.125 |
| 8 |
H |
0.209 |
|
|
0.032 |
| 9 |
H |
0.209 |
|
|
0.032 |
| 10 |
H |
0.191 |
|
|
0.027 |
| 11 |
H |
0.191 |
|
|
0.027 |
| 12 |
H |
0.192 |
|
|
0.113 |
| 13 |
H |
0.192 |
|
|
0.113 |
| 14 |
H |
0.246 |
|
|
0.165 |
| 15 |
H |
0.246 |
|
|
0.165 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.512 |
1.709 |
0.000 |
3.906 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
3.509 |
1.699 |
0.000 |
3.899 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.457 |
-7.482 |
0.000 |
| y |
-7.482 |
-41.410 |
0.000 |
| z |
0.000 |
0.000 |
-36.770 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.367 |
-7.482 |
0.000 |
| y |
-7.482 |
0.704 |
0.000 |
| z |
0.000 |
0.000 |
7.663 |
|
| Polar |
|---|
| 3z2-r2 | 15.326 |
|---|
| x2-y2 | -6.047 |
|---|
| xy | -7.482 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.891 |
1.687 |
0.000 |
| y |
1.687 |
7.924 |
0.000 |
| z |
0.000 |
0.000 |
6.206 |
<r2> (average value of r
2) Å
2
| <r2> |
259.780 |
| (<r2>)1/2 |
16.118 |