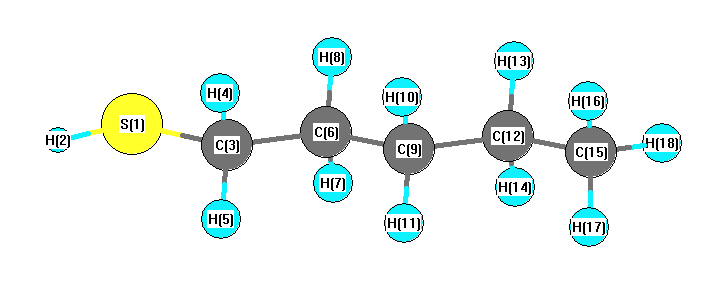
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3112 |
3003 |
35.04 |
|
|
|
| 2 |
A' |
3097 |
2989 |
21.85 |
|
|
|
| 3 |
A' |
3059 |
2952 |
21.98 |
|
|
|
| 4 |
A' |
3047 |
2940 |
35.71 |
|
|
|
| 5 |
A' |
3044 |
2937 |
14.21 |
|
|
|
| 6 |
A' |
3033 |
2926 |
3.42 |
|
|
|
| 7 |
A' |
2458 |
2372 |
44.16 |
|
|
|
| 8 |
A' |
1573 |
1518 |
8.68 |
|
|
|
| 9 |
A' |
1564 |
1509 |
3.23 |
|
|
|
| 10 |
A' |
1551 |
1497 |
0.55 |
|
|
|
| 11 |
A' |
1547 |
1492 |
0.76 |
|
|
|
| 12 |
A' |
1534 |
1480 |
3.35 |
|
|
|
| 13 |
A' |
1466 |
1415 |
3.99 |
|
|
|
| 14 |
A' |
1406 |
1357 |
0.47 |
|
|
|
| 15 |
A' |
1401 |
1352 |
9.32 |
|
|
|
| 16 |
A' |
1347 |
1300 |
13.07 |
|
|
|
| 17 |
A' |
1281 |
1236 |
22.50 |
|
|
|
| 18 |
A' |
1147 |
1107 |
9.28 |
|
|
|
| 19 |
A' |
1052 |
1015 |
2.79 |
|
|
|
| 20 |
A' |
1045 |
1009 |
0.86 |
|
|
|
| 21 |
A' |
1004 |
969 |
0.16 |
|
|
|
| 22 |
A' |
909 |
877 |
1.13 |
|
|
|
| 23 |
A' |
834 |
805 |
3.28 |
|
|
|
| 24 |
A' |
687 |
663 |
8.35 |
|
|
|
| 25 |
A' |
441 |
425 |
3.12 |
|
|
|
| 26 |
A' |
346 |
334 |
0.45 |
|
|
|
| 27 |
A' |
246 |
237 |
2.29 |
|
|
|
| 28 |
A' |
117 |
113 |
1.46 |
|
|
|
| 29 |
A" |
3161 |
3050 |
20.08 |
|
|
|
| 30 |
A" |
3115 |
3005 |
70.38 |
|
|
|
| 31 |
A" |
3101 |
2992 |
6.48 |
|
|
|
| 32 |
A" |
3078 |
2970 |
4.75 |
|
|
|
| 33 |
A" |
3057 |
2950 |
0.58 |
|
|
|
| 34 |
A" |
1570 |
1515 |
8.53 |
|
|
|
| 35 |
A" |
1376 |
1327 |
0.01 |
|
|
|
| 36 |
A" |
1375 |
1326 |
0.83 |
|
|
|
| 37 |
A" |
1328 |
1282 |
0.27 |
|
|
|
| 38 |
A" |
1255 |
1211 |
0.00 |
|
|
|
| 39 |
A" |
1115 |
1076 |
1.20 |
|
|
|
| 40 |
A" |
993 |
958 |
0.01 |
|
|
|
| 41 |
A" |
863 |
833 |
2.43 |
|
|
|
| 42 |
A" |
781 |
754 |
0.23 |
|
|
|
| 43 |
A" |
755 |
728 |
7.57 |
|
|
|
| 44 |
A" |
252 |
243 |
0.02 |
|
|
|
| 45 |
A" |
177 |
171 |
23.02 |
|
|
|
| 46 |
A" |
149 |
144 |
2.73 |
|
|
|
| 47 |
A" |
105 |
102 |
4.07 |
|
|
|
| 48 |
A" |
67 |
65 |
1.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35511.0 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 34264.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.000 |
|
|
-0.376 |
| 2 |
H |
0.068 |
|
|
0.220 |
| 3 |
C |
-0.592 |
|
|
-0.311 |
| 4 |
H |
0.227 |
|
|
0.153 |
| 5 |
H |
0.227 |
|
|
0.153 |
| 6 |
C |
-0.373 |
|
|
0.057 |
| 7 |
H |
0.208 |
|
|
0.070 |
| 8 |
H |
0.208 |
|
|
0.070 |
| 9 |
C |
-0.355 |
|
|
-0.242 |
| 10 |
H |
0.186 |
|
|
0.073 |
| 11 |
H |
0.186 |
|
|
0.073 |
| 12 |
C |
-0.377 |
|
|
0.148 |
| 13 |
H |
0.189 |
|
|
-0.013 |
| 14 |
H |
0.189 |
|
|
-0.013 |
| 15 |
C |
-0.552 |
|
|
-0.281 |
| 16 |
H |
0.186 |
|
|
0.072 |
| 17 |
H |
0.186 |
|
|
0.072 |
| 18 |
H |
0.189 |
|
|
0.073 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.403 |
-2.094 |
0.000 |
2.132 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.377 |
-2.130 |
0.000 |
2.163 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.943 |
4.229 |
0.000 |
| y |
4.229 |
-52.671 |
0.000 |
| z |
0.000 |
0.000 |
-48.640 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.712 |
4.229 |
0.000 |
| y |
4.229 |
-5.879 |
0.000 |
| z |
0.000 |
0.000 |
0.167 |
|
| Polar |
|---|
| 3z2-r2 | 0.333 |
|---|
| x2-y2 | 7.728 |
|---|
| xy | 4.229 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.362 |
-1.922 |
0.000 |
| y |
-1.922 |
9.943 |
0.000 |
| z |
0.000 |
0.000 |
7.572 |
<r2> (average value of r
2) Å
2
| <r2> |
421.487 |
| (<r2>)1/2 |
20.530 |