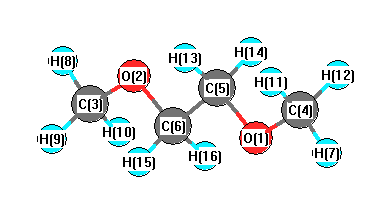
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3129 |
3019 |
0.00 |
|
|
|
| 2 |
Ag |
3010 |
2904 |
0.00 |
|
|
|
| 3 |
Ag |
2990 |
2885 |
0.00 |
|
|
|
| 4 |
Ag |
1592 |
1536 |
0.00 |
|
|
|
| 5 |
Ag |
1573 |
1518 |
0.00 |
|
|
|
| 6 |
Ag |
1514 |
1460 |
0.00 |
|
|
|
| 7 |
Ag |
1441 |
1390 |
0.00 |
|
|
|
| 8 |
Ag |
1213 |
1171 |
0.00 |
|
|
|
| 9 |
Ag |
1105 |
1066 |
0.00 |
|
|
|
| 10 |
Ag |
1065 |
1027 |
0.00 |
|
|
|
| 11 |
Ag |
954 |
920 |
0.00 |
|
|
|
| 12 |
Ag |
380 |
366 |
0.00 |
|
|
|
| 13 |
Ag |
319 |
308 |
0.00 |
|
|
|
| 14 |
Au |
3061 |
2954 |
111.50 |
|
|
|
| 15 |
Au |
3030 |
2923 |
131.57 |
|
|
|
| 16 |
Au |
1550 |
1496 |
14.21 |
|
|
|
| 17 |
Au |
1223 |
1180 |
7.10 |
|
|
|
| 18 |
Au |
1162 |
1121 |
0.49 |
|
|
|
| 19 |
Au |
884 |
853 |
0.01 |
|
|
|
| 20 |
Au |
239 |
231 |
10.39 |
|
|
|
| 21 |
Au |
103 |
100 |
12.44 |
|
|
|
| 22 |
Au |
64 |
62 |
3.26 |
|
|
|
| 23 |
Bg |
3031 |
2924 |
0.00 |
|
|
|
| 24 |
Bg |
3029 |
2923 |
0.00 |
|
|
|
| 25 |
Bg |
1550 |
1496 |
0.00 |
|
|
|
| 26 |
Bg |
1333 |
1287 |
0.00 |
|
|
|
| 27 |
Bg |
1180 |
1139 |
0.00 |
|
|
|
| 28 |
Bg |
1142 |
1102 |
0.00 |
|
|
|
| 29 |
Bg |
229 |
221 |
0.00 |
|
|
|
| 30 |
Bg |
122 |
118 |
0.00 |
|
|
|
| 31 |
Bu |
3129 |
3019 |
56.28 |
|
|
|
| 32 |
Bu |
3016 |
2910 |
80.03 |
|
|
|
| 33 |
Bu |
2990 |
2885 |
95.92 |
|
|
|
| 34 |
Bu |
1601 |
1545 |
2.47 |
|
|
|
| 35 |
Bu |
1577 |
1521 |
25.91 |
|
|
|
| 36 |
Bu |
1508 |
1456 |
7.29 |
|
|
|
| 37 |
Bu |
1378 |
1329 |
24.91 |
|
|
|
| 38 |
Bu |
1191 |
1149 |
44.51 |
|
|
|
| 39 |
Bu |
1116 |
1077 |
236.73 |
|
|
|
| 40 |
Bu |
900 |
869 |
35.43 |
|
|
|
| 41 |
Bu |
484 |
467 |
6.98 |
|
|
|
| 42 |
Bu |
130 |
126 |
6.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31117.7 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30025.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.492 |
|
|
-0.391 |
| 2 |
O |
-0.492 |
|
|
-0.388 |
| 3 |
C |
-0.308 |
|
|
-0.023 |
| 4 |
C |
-0.308 |
|
|
-0.030 |
| 5 |
C |
-0.128 |
|
|
0.135 |
| 6 |
C |
-0.128 |
|
|
0.103 |
| 7 |
H |
0.204 |
|
|
0.093 |
| 8 |
H |
0.204 |
|
|
0.091 |
| 9 |
H |
0.175 |
|
|
0.056 |
| 10 |
H |
0.175 |
|
|
0.056 |
| 11 |
H |
0.175 |
|
|
0.057 |
| 12 |
H |
0.175 |
|
|
0.057 |
| 13 |
H |
0.187 |
|
|
0.042 |
| 14 |
H |
0.187 |
|
|
0.042 |
| 15 |
H |
0.187 |
|
|
0.049 |
| 16 |
H |
0.187 |
|
|
0.049 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.007 |
-0.007 |
0.000 |
0.010 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.618 |
2.757 |
0.000 |
| y |
2.757 |
-28.600 |
0.000 |
| z |
0.000 |
0.000 |
-38.182 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.227 |
2.757 |
0.000 |
| y |
2.757 |
10.800 |
0.000 |
| z |
0.000 |
0.000 |
-3.573 |
|
| Polar |
|---|
| 3z2-r2 | -7.146 |
|---|
| x2-y2 | -12.019 |
|---|
| xy | 2.757 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.308 |
1.537 |
0.000 |
| y |
1.537 |
8.544 |
0.000 |
| z |
0.000 |
0.000 |
6.130 |
<r2> (average value of r
2) Å
2
| <r2> |
276.857 |
| (<r2>)1/2 |
16.639 |