Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -310.592920 |
| Energy at 298.15K | -310.608437 |
| HF Energy | -310.592920 |
| Nuclear repulsion energy | 311.354035 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3117 |
3007 |
45.55 |
|
|
|
| 2 |
A1 |
3071 |
2963 |
31.31 |
|
|
|
| 3 |
A1 |
3047 |
2940 |
14.28 |
|
|
|
| 4 |
A1 |
2979 |
2875 |
70.94 |
|
|
|
| 5 |
A1 |
1597 |
1541 |
0.59 |
|
|
|
| 6 |
A1 |
1572 |
1516 |
8.66 |
|
|
|
| 7 |
A1 |
1558 |
1504 |
1.97 |
|
|
|
| 8 |
A1 |
1474 |
1422 |
0.66 |
|
|
|
| 9 |
A1 |
1454 |
1403 |
4.22 |
|
|
|
| 10 |
A1 |
1370 |
1321 |
0.03 |
|
|
|
| 11 |
A1 |
1175 |
1134 |
0.04 |
|
|
|
| 12 |
A1 |
1037 |
1001 |
0.65 |
|
|
|
| 13 |
A1 |
947 |
914 |
10.70 |
|
|
|
| 14 |
A1 |
931 |
899 |
12.68 |
|
|
|
| 15 |
A1 |
426 |
412 |
0.69 |
|
|
|
| 16 |
A1 |
314 |
303 |
0.08 |
|
|
|
| 17 |
A1 |
100 |
97 |
0.23 |
|
|
|
| 18 |
A2 |
3122 |
3012 |
0.00 |
|
|
|
| 19 |
A2 |
3093 |
2984 |
0.00 |
|
|
|
| 20 |
A2 |
2998 |
2893 |
0.00 |
|
|
|
| 21 |
A2 |
1569 |
1514 |
0.00 |
|
|
|
| 22 |
A2 |
1356 |
1308 |
0.00 |
|
|
|
| 23 |
A2 |
1291 |
1246 |
0.00 |
|
|
|
| 24 |
A2 |
1184 |
1143 |
0.00 |
|
|
|
| 25 |
A2 |
935 |
902 |
0.00 |
|
|
|
| 26 |
A2 |
797 |
769 |
0.00 |
|
|
|
| 27 |
A2 |
238 |
230 |
0.00 |
|
|
|
| 28 |
A2 |
138 |
133 |
0.00 |
|
|
|
| 29 |
A2 |
79 |
76 |
0.00 |
|
|
|
| 30 |
B1 |
3122 |
3013 |
111.02 |
|
|
|
| 31 |
B1 |
3093 |
2984 |
2.89 |
|
|
|
| 32 |
B1 |
3000 |
2894 |
105.04 |
|
|
|
| 33 |
B1 |
1569 |
1514 |
18.03 |
|
|
|
| 34 |
B1 |
1355 |
1308 |
0.02 |
|
|
|
| 35 |
B1 |
1292 |
1247 |
1.06 |
|
|
|
| 36 |
B1 |
1210 |
1168 |
2.69 |
|
|
|
| 37 |
B1 |
949 |
915 |
4.02 |
|
|
|
| 38 |
B1 |
798 |
770 |
7.23 |
|
|
|
| 39 |
B1 |
237 |
229 |
0.10 |
|
|
|
| 40 |
B1 |
157 |
152 |
6.85 |
|
|
|
| 41 |
B1 |
50 |
48 |
0.47 |
|
|
|
| 42 |
B2 |
3117 |
3007 |
17.97 |
|
|
|
| 43 |
B2 |
3070 |
2962 |
4.71 |
|
|
|
| 44 |
B2 |
3047 |
2940 |
33.62 |
|
|
|
| 45 |
B2 |
2969 |
2865 |
12.43 |
|
|
|
| 46 |
B2 |
1581 |
1526 |
11.34 |
|
|
|
| 47 |
B2 |
1570 |
1515 |
0.16 |
|
|
|
| 48 |
B2 |
1556 |
1502 |
4.08 |
|
|
|
| 49 |
B2 |
1464 |
1413 |
13.29 |
|
|
|
| 50 |
B2 |
1404 |
1355 |
31.94 |
|
|
|
| 51 |
B2 |
1353 |
1305 |
17.77 |
|
|
|
| 52 |
B2 |
1142 |
1102 |
13.35 |
|
|
|
| 53 |
B2 |
1103 |
1065 |
181.78 |
|
|
|
| 54 |
B2 |
1029 |
993 |
0.16 |
|
|
|
| 55 |
B2 |
901 |
869 |
4.23 |
|
|
|
| 56 |
B2 |
502 |
484 |
7.44 |
|
|
|
| 57 |
B2 |
247 |
238 |
1.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42926.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 41419.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.370 |
| C2 |
0.000 |
1.214 |
-0.434 |
| C3 |
0.000 |
-1.214 |
-0.434 |
| C4 |
0.000 |
2.393 |
0.542 |
| C5 |
0.000 |
-2.393 |
0.542 |
| C6 |
0.000 |
3.746 |
-0.197 |
| C7 |
0.000 |
-3.746 |
-0.197 |
| H8 |
0.890 |
1.256 |
-1.083 |
| H9 |
-0.890 |
1.256 |
-1.083 |
| H10 |
-0.890 |
-1.256 |
-1.083 |
| H11 |
0.890 |
-1.256 |
-1.083 |
| H12 |
0.883 |
2.304 |
1.184 |
| H13 |
-0.883 |
2.304 |
1.184 |
| H14 |
-0.883 |
-2.304 |
1.184 |
| H15 |
0.883 |
-2.304 |
1.184 |
| H16 |
0.000 |
4.579 |
0.515 |
| H17 |
-0.888 |
3.843 |
-0.832 |
| H18 |
0.888 |
3.843 |
-0.832 |
| H19 |
0.000 |
-4.579 |
0.515 |
| H20 |
0.888 |
-3.843 |
-0.832 |
| H21 |
-0.888 |
-3.843 |
-0.832 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
H21 |
| O1 | | 1.4561 | 1.4561 | 2.3992 | 2.3992 | 3.7888 | 3.7888 | 2.1173 | 2.1173 | 2.1173 | 2.1173 | 2.5985 | 2.5985 | 2.5985 | 2.5985 | 4.5808 | 4.1233 | 4.1233 | 4.5808 | 4.1233 | 4.1233 |
C2 | 1.4561 | | 2.4271 | 1.5312 | 3.7365 | 2.5437 | 4.9654 | 1.1025 | 1.1025 | 2.7042 | 2.7042 | 2.1420 | 2.1420 | 3.9717 | 3.9717 | 3.4963 | 2.8035 | 2.8035 | 5.8694 | 5.1491 | 5.1491 | C3 | 1.4561 | 2.4271 | | 3.7365 | 1.5312 | 4.9654 | 2.5437 | 2.7042 | 2.7042 | 1.1025 | 1.1025 | 3.9717 | 3.9717 | 2.1420 | 2.1420 | 5.8694 | 5.1491 | 5.1491 | 3.4963 | 2.8035 | 2.8035 | C4 | 2.3992 | 1.5312 | 3.7365 | | 4.7862 | 1.5415 | 6.1835 | 2.1742 | 2.1742 | 4.0927 | 4.0927 | 1.0954 | 1.0954 | 4.8227 | 4.8227 | 2.1856 | 2.1862 | 2.1862 | 6.9717 | 6.4469 | 6.4469 | C5 | 2.3992 | 3.7365 | 1.5312 | 4.7862 | | 6.1835 | 1.5415 | 4.0927 | 4.0927 | 2.1742 | 2.1742 | 4.8227 | 4.8227 | 1.0954 | 1.0954 | 6.9717 | 6.4469 | 6.4469 | 2.1856 | 2.1862 | 2.1862 | C6 | 3.7888 | 2.5437 | 4.9654 | 1.5415 | 6.1835 | | 7.4923 | 2.7892 | 2.7892 | 5.1576 | 5.1576 | 2.1826 | 2.1826 | 6.2686 | 6.2686 | 1.0949 | 1.0965 | 1.0965 | 8.3550 | 7.6671 | 7.6671 | C7 | 3.7888 | 4.9654 | 2.5437 | 6.1835 | 1.5415 | 7.4923 | | 5.1576 | 5.1576 | 2.7892 | 2.7892 | 6.2686 | 6.2686 | 2.1826 | 2.1826 | 8.3550 | 7.6671 | 7.6671 | 1.0949 | 1.0965 | 1.0965 | H8 | 2.1173 | 1.1025 | 2.7042 | 2.1742 | 4.0927 | 2.7892 | 5.1576 | | 1.7807 | 3.0791 | 2.5120 | 2.4975 | 3.0634 | 4.5784 | 4.2208 | 3.7928 | 3.1490 | 2.5988 | 6.1146 | 5.1049 | 5.4057 | H9 | 2.1173 | 1.1025 | 2.7042 | 2.1742 | 4.0927 | 2.7892 | 5.1576 | 1.7807 | | 2.5120 | 3.0791 | 3.0634 | 2.4975 | 4.2208 | 4.5784 | 3.7928 | 2.5988 | 3.1490 | 6.1146 | 5.4057 | 5.1049 | H10 | 2.1173 | 2.7042 | 1.1025 | 4.0927 | 2.1742 | 5.1576 | 2.7892 | 3.0791 | 2.5120 | | 1.7807 | 4.5784 | 4.2208 | 2.4975 | 3.0634 | 6.1146 | 5.1049 | 5.4057 | 3.7928 | 3.1490 | 2.5988 | H11 | 2.1173 | 2.7042 | 1.1025 | 4.0927 | 2.1742 | 5.1576 | 2.7892 | 2.5120 | 3.0791 | 1.7807 | | 4.2208 | 4.5784 | 3.0634 | 2.4975 | 6.1146 | 5.4057 | 5.1049 | 3.7928 | 2.5988 | 3.1490 | H12 | 2.5985 | 2.1420 | 3.9717 | 1.0954 | 4.8227 | 2.1826 | 6.2686 | 2.4975 | 3.0634 | 4.5784 | 4.2208 | | 1.7670 | 4.9360 | 4.6089 | 2.5297 | 3.0934 | 2.5359 | 6.9716 | 6.4693 | 6.7075 | H13 | 2.5985 | 2.1420 | 3.9717 | 1.0954 | 4.8227 | 2.1826 | 6.2686 | 3.0634 | 2.4975 | 4.2208 | 4.5784 | 1.7670 | | 4.6089 | 4.9360 | 2.5297 | 2.5359 | 3.0934 | 6.9716 | 6.7075 | 6.4693 | H14 | 2.5985 | 3.9717 | 2.1420 | 4.8227 | 1.0954 | 6.2686 | 2.1826 | 4.5784 | 4.2208 | 2.4975 | 3.0634 | 4.9360 | 4.6089 | | 1.7670 | 6.9716 | 6.4693 | 6.7075 | 2.5297 | 3.0934 | 2.5359 | H15 | 2.5985 | 3.9717 | 2.1420 | 4.8227 | 1.0954 | 6.2686 | 2.1826 | 4.2208 | 4.5784 | 3.0634 | 2.4975 | 4.6089 | 4.9360 | 1.7670 | | 6.9716 | 6.7075 | 6.4693 | 2.5297 | 2.5359 | 3.0934 | H16 | 4.5808 | 3.4963 | 5.8694 | 2.1856 | 6.9717 | 1.0949 | 8.3550 | 3.7928 | 3.7928 | 6.1146 | 6.1146 | 2.5297 | 2.5297 | 6.9716 | 6.9716 | | 1.7735 | 1.7735 | 9.1571 | 8.5744 | 8.5744 | H17 | 4.1233 | 2.8035 | 5.1491 | 2.1862 | 6.4469 | 1.0965 | 7.6671 | 3.1490 | 2.5988 | 5.1049 | 5.4057 | 3.0934 | 2.5359 | 6.4693 | 6.7075 | 1.7735 | | 1.7759 | 8.5744 | 7.8879 | 7.6854 | H18 | 4.1233 | 2.8035 | 5.1491 | 2.1862 | 6.4469 | 1.0965 | 7.6671 | 2.5988 | 3.1490 | 5.4057 | 5.1049 | 2.5359 | 3.0934 | 6.7075 | 6.4693 | 1.7735 | 1.7759 | | 8.5744 | 7.6854 | 7.8879 | H19 | 4.5808 | 5.8694 | 3.4963 | 6.9717 | 2.1856 | 8.3550 | 1.0949 | 6.1146 | 6.1146 | 3.7928 | 3.7928 | 6.9716 | 6.9716 | 2.5297 | 2.5297 | 9.1571 | 8.5744 | 8.5744 | | 1.7735 | 1.7735 | H20 | 4.1233 | 5.1491 | 2.8035 | 6.4469 | 2.1862 | 7.6671 | 1.0965 | 5.1049 | 5.4057 | 3.1490 | 2.5988 | 6.4693 | 6.7075 | 3.0934 | 2.5359 | 8.5744 | 7.8879 | 7.6854 | 1.7735 | | 1.7759 | H21 | 4.1233 | 5.1491 | 2.8035 | 6.4469 | 2.1862 | 7.6671 | 1.0965 | 5.4057 | 5.1049 | 2.5988 | 3.1490 | 6.7075 | 6.4693 | 2.5359 | 3.0934 | 8.5744 | 7.6854 | 7.8879 | 1.7735 | 1.7759 | |
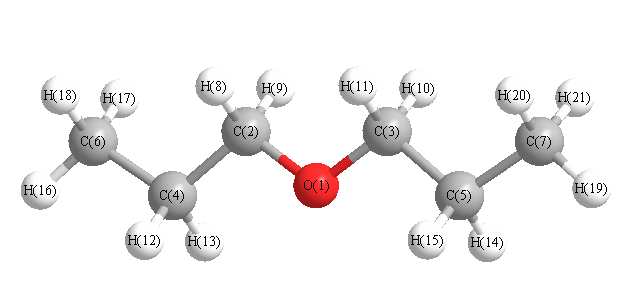 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
106.834 |
|
O1 |
C2 |
H8 |
110.935 |
| O1 |
C2 |
H9 |
110.935 |
|
O1 |
C3 |
C5 |
106.834 |
| O1 |
C3 |
H10 |
110.935 |
|
O1 |
C3 |
H11 |
110.935 |
| C2 |
O1 |
C3 |
112.905 |
|
C2 |
C4 |
C6 |
111.751 |
| C2 |
C4 |
H12 |
108.126 |
|
C2 |
C4 |
H13 |
108.126 |
| C3 |
C5 |
C7 |
111.751 |
|
C3 |
C5 |
H14 |
108.126 |
| C3 |
C5 |
H15 |
108.126 |
|
C4 |
C2 |
H8 |
110.222 |
| C4 |
C2 |
H9 |
110.222 |
|
C4 |
C6 |
H16 |
110.854 |
| C4 |
C6 |
H17 |
110.806 |
|
C4 |
C6 |
H18 |
110.806 |
| C5 |
C3 |
H10 |
110.222 |
|
C5 |
C3 |
H11 |
110.222 |
| C5 |
C7 |
H19 |
110.854 |
|
C5 |
C7 |
H20 |
110.806 |
| C5 |
C7 |
H21 |
110.806 |
|
C6 |
C4 |
H12 |
110.587 |
| C6 |
C4 |
H13 |
110.587 |
|
C7 |
C5 |
H14 |
110.587 |
| C7 |
C5 |
H15 |
110.587 |
|
H8 |
C2 |
H9 |
107.715 |
| H10 |
C3 |
H11 |
107.715 |
|
H12 |
C4 |
H13 |
107.517 |
| H14 |
C5 |
H15 |
107.517 |
|
H16 |
C6 |
H17 |
108.052 |
| H16 |
C6 |
H18 |
108.052 |
|
H17 |
C6 |
H18 |
108.158 |
| H19 |
C7 |
H20 |
108.052 |
|
H19 |
C7 |
H21 |
108.052 |
| H20 |
C7 |
H21 |
108.158 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.494 |
|
|
|
| 2 |
C |
-0.093 |
|
|
|
| 3 |
C |
-0.093 |
|
|
|
| 4 |
C |
-0.405 |
|
|
|
| 5 |
C |
-0.405 |
|
|
|
| 6 |
C |
-0.563 |
|
|
|
| 7 |
C |
-0.563 |
|
|
|
| 8 |
H |
0.168 |
|
|
|
| 9 |
H |
0.168 |
|
|
|
| 10 |
H |
0.168 |
|
|
|
| 11 |
H |
0.168 |
|
|
|
| 12 |
H |
0.203 |
|
|
|
| 13 |
H |
0.203 |
|
|
|
| 14 |
H |
0.203 |
|
|
|
| 15 |
H |
0.203 |
|
|
|
| 16 |
H |
0.194 |
|
|
|
| 17 |
H |
0.187 |
|
|
|
| 18 |
H |
0.187 |
|
|
|
| 19 |
H |
0.194 |
|
|
|
| 20 |
H |
0.187 |
|
|
|
| 21 |
H |
0.187 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.132 |
1.132 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.007 |
0.000 |
0.000 |
| y |
0.000 |
-43.135 |
0.000 |
| z |
0.000 |
0.000 |
-46.708 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.085 |
0.000 |
0.000 |
| y |
0.000 |
3.222 |
0.000 |
| z |
0.000 |
0.000 |
-2.137 |
|
| Polar |
|---|
| 3z2-r2 | -4.274 |
|---|
| x2-y2 | -2.872 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.524 |
0.000 |
0.000 |
| y |
0.000 |
12.457 |
0.000 |
| z |
0.000 |
0.000 |
8.722 |
<r2> (average value of r
2) Å
2
| <r2> |
442.023 |
| (<r2>)1/2 |
21.024 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -310.595725 |
| Energy at 298.15K | -310.611611 |
| HF Energy | -310.595725 |
| Nuclear repulsion energy | 324.272302 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3151 |
3040 |
3.94 |
|
|
|
| 2 |
A |
3116 |
3007 |
63.68 |
|
|
|
| 3 |
A |
3096 |
2987 |
0.27 |
|
|
|
| 4 |
A |
3062 |
2954 |
3.41 |
|
|
|
| 5 |
A |
3052 |
2945 |
3.97 |
|
|
|
| 6 |
A |
3013 |
2907 |
2.96 |
|
|
|
| 7 |
A |
2988 |
2883 |
79.56 |
|
|
|
| 8 |
A |
1592 |
1536 |
2.59 |
|
|
|
| 9 |
A |
1582 |
1526 |
6.18 |
|
|
|
| 10 |
A |
1561 |
1506 |
9.73 |
|
|
|
| 11 |
A |
1541 |
1487 |
1.25 |
|
|
|
| 12 |
A |
1469 |
1417 |
6.29 |
|
|
|
| 13 |
A |
1440 |
1390 |
3.55 |
|
|
|
| 14 |
A |
1410 |
1360 |
0.29 |
|
|
|
| 15 |
A |
1346 |
1299 |
0.56 |
|
|
|
| 16 |
A |
1280 |
1235 |
0.00 |
|
|
|
| 17 |
A |
1190 |
1148 |
1.61 |
|
|
|
| 18 |
A |
1146 |
1106 |
7.20 |
|
|
|
| 19 |
A |
1057 |
1020 |
2.55 |
|
|
|
| 20 |
A |
965 |
932 |
1.97 |
|
|
|
| 21 |
A |
924 |
892 |
10.67 |
|
|
|
| 22 |
A |
891 |
860 |
0.67 |
|
|
|
| 23 |
A |
766 |
739 |
1.53 |
|
|
|
| 24 |
A |
480 |
463 |
0.39 |
|
|
|
| 25 |
A |
349 |
336 |
0.88 |
|
|
|
| 26 |
A |
276 |
266 |
1.13 |
|
|
|
| 27 |
A |
175 |
168 |
0.09 |
|
|
|
| 28 |
A |
126 |
122 |
0.00 |
|
|
|
| 29 |
A |
55 |
54 |
0.00 |
|
|
|
| 30 |
B |
3151 |
3040 |
34.33 |
|
|
|
| 31 |
B |
3116 |
3007 |
28.25 |
|
|
|
| 32 |
B |
3096 |
2987 |
27.43 |
|
|
|
| 33 |
B |
3062 |
2954 |
57.45 |
|
|
|
| 34 |
B |
3052 |
2945 |
47.62 |
|
|
|
| 35 |
B |
3014 |
2908 |
95.58 |
|
|
|
| 36 |
B |
2978 |
2874 |
8.99 |
|
|
|
| 37 |
B |
1581 |
1525 |
7.00 |
|
|
|
| 38 |
B |
1573 |
1518 |
7.72 |
|
|
|
| 39 |
B |
1561 |
1506 |
7.17 |
|
|
|
| 40 |
B |
1541 |
1487 |
7.19 |
|
|
|
| 41 |
B |
1450 |
1399 |
19.96 |
|
|
|
| 42 |
B |
1409 |
1360 |
3.04 |
|
|
|
| 43 |
B |
1393 |
1344 |
49.42 |
|
|
|
| 44 |
B |
1335 |
1288 |
0.13 |
|
|
|
| 45 |
B |
1279 |
1234 |
3.28 |
|
|
|
| 46 |
B |
1193 |
1151 |
2.80 |
|
|
|
| 47 |
B |
1144 |
1104 |
10.86 |
|
|
|
| 48 |
B |
1089 |
1050 |
139.68 |
|
|
|
| 49 |
B |
1025 |
989 |
53.83 |
|
|
|
| 50 |
B |
965 |
932 |
0.22 |
|
|
|
| 51 |
B |
873 |
842 |
0.74 |
|
|
|
| 52 |
B |
810 |
782 |
7.89 |
|
|
|
| 53 |
B |
500 |
483 |
1.16 |
|
|
|
| 54 |
B |
327 |
315 |
1.67 |
|
|
|
| 55 |
B |
248 |
239 |
5.92 |
|
|
|
| 56 |
B |
187 |
180 |
5.03 |
|
|
|
| 57 |
B |
72 |
69 |
1.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 43058.2 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 41546.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.126 |
| C2 |
0.054 |
1.218 |
0.928 |
| C3 |
-0.054 |
-1.218 |
0.928 |
| C4 |
0.000 |
2.388 |
-0.060 |
| C5 |
0.000 |
-2.388 |
-0.060 |
| C6 |
-1.297 |
2.343 |
-0.893 |
| C7 |
1.297 |
-2.343 |
-0.893 |
| H8 |
-0.805 |
1.265 |
1.618 |
| H9 |
0.977 |
1.247 |
1.528 |
| H10 |
0.805 |
-1.265 |
1.618 |
| H11 |
-0.977 |
-1.247 |
1.528 |
| H12 |
0.869 |
2.311 |
-0.723 |
| H13 |
0.077 |
3.331 |
0.496 |
| H14 |
-0.869 |
-2.311 |
-0.723 |
| H15 |
-0.077 |
-3.331 |
0.496 |
| H16 |
-1.297 |
3.112 |
-1.673 |
| H17 |
-1.379 |
1.356 |
-1.353 |
| H18 |
-2.172 |
2.502 |
-0.250 |
| H19 |
1.297 |
-3.112 |
-1.673 |
| H20 |
1.379 |
-1.356 |
-1.353 |
| H21 |
2.172 |
-2.502 |
-0.250 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
H21 |
| O1 | | 1.4591 | 1.4591 | 2.3954 | 2.3954 | 2.8656 | 2.8656 | 2.1151 | 2.1148 | 2.1151 | 2.1148 | 2.6112 | 3.3518 | 2.6112 | 3.3518 | 3.8210 | 2.4351 | 3.3344 | 3.8210 | 2.4351 | 3.3344 |
C2 | 1.4591 | | 2.4376 | 1.5330 | 3.7392 | 2.5314 | 4.1884 | 1.1020 | 1.1010 | 2.6841 | 2.7374 | 2.1422 | 2.1569 | 4.0040 | 4.5706 | 3.4896 | 2.6978 | 2.8270 | 5.2014 | 3.6861 | 4.4397 | C3 | 1.4591 | 2.4376 | | 3.7392 | 1.5330 | 4.1884 | 2.5314 | 2.6841 | 2.7374 | 1.1020 | 1.1010 | 4.0040 | 4.5706 | 2.1422 | 2.1569 | 5.2014 | 3.6861 | 4.4397 | 3.4896 | 2.6978 | 2.8270 | C4 | 2.3954 | 1.5330 | 3.7392 | | 4.7763 | 1.5421 | 4.9761 | 2.1736 | 2.1859 | 4.1001 | 4.0850 | 1.0958 | 1.0967 | 4.8248 | 5.7461 | 2.1924 | 2.1540 | 2.1834 | 5.8762 | 4.1944 | 5.3539 | C5 | 2.3954 | 3.7392 | 1.5330 | 4.7763 | | 4.9761 | 1.5421 | 4.1001 | 4.0850 | 2.1736 | 2.1859 | 4.8248 | 5.7461 | 1.0958 | 1.0967 | 5.8762 | 4.1944 | 5.3539 | 2.1924 | 2.1540 | 2.1834 | C6 | 2.8656 | 2.5314 | 4.1884 | 1.5421 | 4.9761 | | 5.3566 | 2.7763 | 3.4974 | 4.8726 | 4.3415 | 2.1730 | 2.1887 | 4.6771 | 5.9672 | 1.0949 | 1.0925 | 1.0968 | 6.0904 | 4.5890 | 5.9934 | C7 | 2.8656 | 4.1884 | 2.5314 | 4.9761 | 1.5421 | 5.3566 | | 4.8726 | 4.3415 | 2.7763 | 3.4974 | 4.6771 | 5.9672 | 2.1730 | 2.1887 | 6.0904 | 4.5890 | 5.9934 | 1.0949 | 1.0925 | 1.0968 | H8 | 2.1151 | 1.1020 | 2.6841 | 2.1736 | 4.1001 | 2.7763 | 4.8726 | | 1.7838 | 2.9990 | 2.5194 | 3.0621 | 2.5106 | 4.2751 | 4.7865 | 3.8052 | 3.0277 | 2.6247 | 5.8654 | 4.5242 | 5.1519 | H9 | 2.1148 | 1.1010 | 2.7374 | 2.1859 | 4.0850 | 3.4974 | 4.3415 | 1.7838 | | 2.5194 | 3.1674 | 2.4923 | 2.4934 | 4.5970 | 4.8090 | 4.3464 | 3.7232 | 3.8278 | 5.4166 | 3.9034 | 4.3175 | H10 | 2.1151 | 2.6841 | 1.1020 | 4.1001 | 2.1736 | 4.8726 | 2.7763 | 2.9990 | 2.5194 | | 1.7838 | 4.2751 | 4.7865 | 3.0621 | 2.5106 | 5.8654 | 4.5242 | 5.1519 | 3.8052 | 3.0277 | 2.6247 | H11 | 2.1148 | 2.7374 | 1.1010 | 4.0850 | 2.1859 | 4.3415 | 3.4974 | 2.5194 | 3.1674 | 1.7838 | | 4.5970 | 4.8090 | 2.4923 | 2.4934 | 5.4166 | 3.9034 | 4.3175 | 4.3464 | 3.7232 | 3.8278 | H12 | 2.6112 | 2.1422 | 4.0040 | 1.0958 | 4.8248 | 2.1730 | 4.6771 | 3.0621 | 2.4923 | 4.2751 | 4.5970 | | 1.7752 | 4.9383 | 5.8489 | 2.4969 | 2.5225 | 3.0835 | 5.5219 | 3.7558 | 5.0085 | H13 | 3.3518 | 2.1569 | 4.5706 | 1.0967 | 5.7461 | 2.1887 | 5.9672 | 2.5106 | 2.4934 | 4.7865 | 4.8090 | 1.7752 | | 5.8489 | 6.6628 | 2.5764 | 3.0720 | 2.5105 | 6.9058 | 5.2036 | 6.2417 | H14 | 2.6112 | 4.0040 | 2.1422 | 4.8248 | 1.0958 | 4.6771 | 2.1730 | 4.2751 | 4.5970 | 3.0621 | 2.4923 | 4.9383 | 5.8489 | | 1.7752 | 5.5219 | 3.7558 | 5.0085 | 2.4969 | 2.5225 | 3.0835 | H15 | 3.3518 | 4.5706 | 2.1569 | 5.7461 | 1.0967 | 5.9672 | 2.1887 | 4.7865 | 4.8090 | 2.5106 | 2.4934 | 5.8489 | 6.6628 | 1.7752 | | 6.9058 | 5.2036 | 6.2417 | 2.5764 | 3.0720 | 2.5105 | H16 | 3.8210 | 3.4896 | 5.2014 | 2.1924 | 5.8762 | 1.0949 | 6.0904 | 3.8052 | 4.3464 | 5.8654 | 5.4166 | 2.4969 | 2.5764 | 5.5219 | 6.9058 | | 1.7862 | 1.7777 | 6.7422 | 5.2176 | 6.7503 | H17 | 2.4351 | 2.6978 | 3.6861 | 2.1540 | 4.1944 | 1.0925 | 4.5890 | 3.0277 | 3.7232 | 4.5242 | 3.9034 | 2.5225 | 3.0720 | 3.7558 | 5.2036 | 1.7862 | | 1.7770 | 5.2176 | 3.8681 | 5.3580 | H18 | 3.3344 | 2.8270 | 4.4397 | 2.1834 | 5.3539 | 1.0968 | 5.9934 | 2.6247 | 3.8278 | 5.1519 | 4.3175 | 3.0835 | 2.5105 | 5.0085 | 6.2417 | 1.7777 | 1.7770 | | 6.7503 | 5.3580 | 6.6261 | H19 | 3.8210 | 5.2014 | 3.4896 | 5.8762 | 2.1924 | 6.0904 | 1.0949 | 5.8654 | 5.4166 | 3.8052 | 4.3464 | 5.5219 | 6.9058 | 2.4969 | 2.5764 | 6.7422 | 5.2176 | 6.7503 | | 1.7862 | 1.7777 | H20 | 2.4351 | 3.6861 | 2.6978 | 4.1944 | 2.1540 | 4.5890 | 1.0925 | 4.5242 | 3.9034 | 3.0277 | 3.7232 | 3.7558 | 5.2036 | 2.5225 | 3.0720 | 5.2176 | 3.8681 | 5.3580 | 1.7862 | | 1.7770 | H21 | 3.3344 | 4.4397 | 2.8270 | 5.3539 | 2.1834 | 5.9934 | 1.0968 | 5.1519 | 4.3175 | 2.6247 | 3.8278 | 5.0085 | 6.2417 | 3.0835 | 2.5105 | 6.7503 | 5.3580 | 6.6261 | 1.7777 | 1.7770 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
106.341 |
|
O1 |
C2 |
H8 |
110.578 |
| O1 |
C2 |
H9 |
110.611 |
|
O1 |
C3 |
C5 |
106.341 |
| O1 |
C3 |
H10 |
110.578 |
|
O1 |
C3 |
H11 |
110.611 |
| C2 |
O1 |
C3 |
113.293 |
|
C2 |
C4 |
C6 |
110.811 |
| C2 |
C4 |
H12 |
107.995 |
|
C2 |
C4 |
H13 |
109.085 |
| C3 |
C5 |
C7 |
110.811 |
|
C3 |
C5 |
H14 |
107.995 |
| C3 |
C5 |
H15 |
109.085 |
|
C4 |
C2 |
H8 |
110.078 |
| C4 |
C2 |
H9 |
111.107 |
|
C4 |
C6 |
H16 |
111.358 |
| C4 |
C6 |
H17 |
108.483 |
|
C4 |
C6 |
H18 |
110.529 |
| C5 |
C3 |
H10 |
110.078 |
|
C5 |
C3 |
H11 |
111.107 |
| C5 |
C7 |
H19 |
111.358 |
|
C5 |
C7 |
H20 |
108.483 |
| C5 |
C7 |
H21 |
110.529 |
|
C6 |
C4 |
H12 |
109.772 |
| C6 |
C4 |
H13 |
110.958 |
|
C7 |
C5 |
H14 |
109.772 |
| C7 |
C5 |
H15 |
110.958 |
|
H8 |
C2 |
H9 |
108.141 |
| H10 |
C3 |
H11 |
108.141 |
|
H12 |
C4 |
H13 |
108.131 |
| H14 |
C5 |
H15 |
108.131 |
|
H16 |
C6 |
H17 |
109.493 |
| H16 |
C6 |
H18 |
108.410 |
|
H17 |
C6 |
H18 |
108.522 |
| H19 |
C7 |
H20 |
109.493 |
|
H19 |
C7 |
H21 |
108.410 |
| H20 |
C7 |
H21 |
108.522 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.485 |
|
|
|
| 2 |
C |
-0.100 |
|
|
|
| 3 |
C |
-0.100 |
|
|
|
| 4 |
C |
-0.407 |
|
|
|
| 5 |
C |
-0.407 |
|
|
|
| 6 |
C |
-0.551 |
|
|
|
| 7 |
C |
-0.551 |
|
|
|
| 8 |
H |
0.170 |
|
|
|
| 9 |
H |
0.174 |
|
|
|
| 10 |
H |
0.170 |
|
|
|
| 11 |
H |
0.174 |
|
|
|
| 12 |
H |
0.203 |
|
|
|
| 13 |
H |
0.184 |
|
|
|
| 14 |
H |
0.203 |
|
|
|
| 15 |
H |
0.184 |
|
|
|
| 16 |
H |
0.181 |
|
|
|
| 17 |
H |
0.213 |
|
|
|
| 18 |
H |
0.177 |
|
|
|
| 19 |
H |
0.181 |
|
|
|
| 20 |
H |
0.213 |
|
|
|
| 21 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.250 |
1.250 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.759 |
0.515 |
0.000 |
| y |
0.515 |
-44.156 |
0.000 |
| z |
0.000 |
0.000 |
-45.200 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.081 |
0.515 |
0.000 |
| y |
0.515 |
1.823 |
0.000 |
| z |
0.000 |
0.000 |
0.258 |
|
| Polar |
|---|
| 3z2-r2 | 0.516 |
|---|
| x2-y2 | -2.603 |
|---|
| xy | 0.515 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.960 |
-0.504 |
0.000 |
| y |
-0.504 |
11.245 |
0.000 |
| z |
0.000 |
0.000 |
9.117 |
<r2> (average value of r
2) Å
2
| <r2> |
337.651 |
| (<r2>)1/2 |
18.375 |
