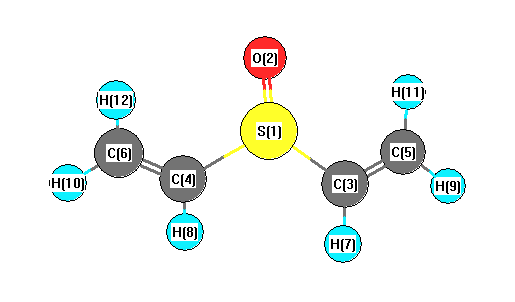
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3175 |
3158 |
1.77 |
|
|
|
| 2 |
A' |
3142 |
3125 |
0.74 |
|
|
|
| 3 |
A' |
2995 |
2979 |
5.66 |
|
|
|
| 4 |
A' |
1621 |
1612 |
8.77 |
|
|
|
| 5 |
A' |
1400 |
1392 |
33.46 |
|
|
|
| 6 |
A' |
1223 |
1216 |
18.22 |
|
|
|
| 7 |
A' |
1016 |
1010 |
54.47 |
|
|
|
| 8 |
A' |
958 |
952 |
0.49 |
|
|
|
| 9 |
A' |
926 |
921 |
0.26 |
|
|
|
| 10 |
A' |
831 |
827 |
21.34 |
|
|
|
| 11 |
A' |
655 |
651 |
72.56 |
|
|
|
| 12 |
A' |
521 |
518 |
0.02 |
|
|
|
| 13 |
A' |
443 |
440 |
2.82 |
|
|
|
| 14 |
A' |
293 |
291 |
3.01 |
|
|
|
| 15 |
A' |
215 |
213 |
1.78 |
|
|
|
| 16 |
A' |
109 |
108 |
0.42 |
|
|
|
| 17 |
A" |
3173 |
3156 |
3.68 |
|
|
|
| 18 |
A" |
3141 |
3124 |
0.08 |
|
|
|
| 19 |
A" |
2992 |
2976 |
16.52 |
|
|
|
| 20 |
A" |
1614 |
1605 |
22.11 |
|
|
|
| 21 |
A" |
1394 |
1386 |
7.31 |
|
|
|
| 22 |
A" |
1203 |
1196 |
46.71 |
|
|
|
| 23 |
A" |
1004 |
999 |
46.71 |
|
|
|
| 24 |
A" |
949 |
944 |
7.82 |
|
|
|
| 25 |
A" |
920 |
915 |
0.22 |
|
|
|
| 26 |
A" |
570 |
567 |
23.55 |
|
|
|
| 27 |
A" |
470 |
468 |
3.27 |
|
|
|
| 28 |
A" |
414 |
412 |
0.19 |
|
|
|
| 29 |
A" |
266 |
265 |
10.56 |
|
|
|
| 30 |
A" |
169 |
168 |
1.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18900.0 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 18796.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.811 |
|
|
|
| 2 |
O |
-0.511 |
|
|
|
| 3 |
C |
-0.436 |
|
|
|
| 4 |
C |
-0.436 |
|
|
|
| 5 |
C |
-0.322 |
|
|
|
| 6 |
C |
-0.322 |
|
|
|
| 7 |
H |
0.199 |
|
|
|
| 8 |
H |
0.199 |
|
|
|
| 9 |
H |
0.193 |
|
|
|
| 10 |
H |
0.193 |
|
|
|
| 11 |
H |
0.215 |
|
|
|
| 12 |
H |
0.215 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.531 |
-1.713 |
0.000 |
3.057 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.724 |
-0.504 |
0.000 |
| y |
-0.504 |
-41.086 |
0.000 |
| z |
0.000 |
0.000 |
-39.222 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.570 |
-0.504 |
0.000 |
| y |
-0.504 |
1.387 |
0.000 |
| z |
0.000 |
0.000 |
4.183 |
|
| Polar |
|---|
| 3z2-r2 | 8.367 |
|---|
| x2-y2 | -4.638 |
|---|
| xy | -0.504 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.899 |
0.916 |
0.000 |
| y |
0.916 |
8.763 |
0.000 |
| z |
0.000 |
0.000 |
11.147 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |