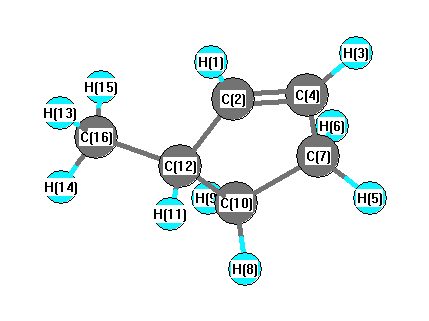
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3226 |
3113 |
21.73 |
|
|
|
| 2 |
A |
3196 |
3084 |
5.51 |
|
|
|
| 3 |
A |
3127 |
3017 |
40.10 |
|
|
|
| 4 |
A |
3112 |
3003 |
21.52 |
|
|
|
| 5 |
A |
3103 |
2994 |
28.80 |
|
|
|
| 6 |
A |
3085 |
2977 |
30.75 |
|
|
|
| 7 |
A |
3067 |
2960 |
14.74 |
|
|
|
| 8 |
A |
3043 |
2936 |
21.42 |
|
|
|
| 9 |
A |
3038 |
2932 |
38.22 |
|
|
|
| 10 |
A |
3019 |
2913 |
21.44 |
|
|
|
| 11 |
A |
1680 |
1621 |
0.30 |
|
|
|
| 12 |
A |
1566 |
1511 |
6.03 |
|
|
|
| 13 |
A |
1564 |
1509 |
4.30 |
|
|
|
| 14 |
A |
1552 |
1498 |
2.78 |
|
|
|
| 15 |
A |
1538 |
1484 |
1.51 |
|
|
|
| 16 |
A |
1460 |
1409 |
4.25 |
|
|
|
| 17 |
A |
1395 |
1347 |
5.14 |
|
|
|
| 18 |
A |
1365 |
1317 |
2.31 |
|
|
|
| 19 |
A |
1351 |
1304 |
0.15 |
|
|
|
| 20 |
A |
1341 |
1294 |
4.55 |
|
|
|
| 21 |
A |
1332 |
1286 |
0.59 |
|
|
|
| 22 |
A |
1255 |
1211 |
1.01 |
|
|
|
| 23 |
A |
1229 |
1186 |
0.65 |
|
|
|
| 24 |
A |
1170 |
1129 |
0.30 |
|
|
|
| 25 |
A |
1143 |
1103 |
3.79 |
|
|
|
| 26 |
A |
1126 |
1086 |
6.36 |
|
|
|
| 27 |
A |
1067 |
1030 |
0.93 |
|
|
|
| 28 |
A |
1010 |
974 |
0.47 |
|
|
|
| 29 |
A |
989 |
954 |
0.01 |
|
|
|
| 30 |
A |
977 |
942 |
3.25 |
|
|
|
| 31 |
A |
940 |
907 |
0.63 |
|
|
|
| 32 |
A |
894 |
863 |
3.15 |
|
|
|
| 33 |
A |
852 |
822 |
1.26 |
|
|
|
| 34 |
A |
827 |
798 |
2.68 |
|
|
|
| 35 |
A |
774 |
747 |
0.24 |
|
|
|
| 36 |
A |
748 |
722 |
38.67 |
|
|
|
| 37 |
A |
604 |
583 |
7.10 |
|
|
|
| 38 |
A |
499 |
481 |
3.17 |
|
|
|
| 39 |
A |
346 |
334 |
0.11 |
|
|
|
| 40 |
A |
284 |
274 |
0.17 |
|
|
|
| 41 |
A |
241 |
232 |
0.06 |
|
|
|
| 42 |
A |
94 |
90 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32114.4 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30987.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.173 |
|
|
0.174 |
| 2 |
C |
-0.133 |
|
|
-0.370 |
| 3 |
H |
0.175 |
|
|
0.132 |
| 4 |
C |
-0.167 |
|
|
-0.142 |
| 5 |
H |
0.199 |
|
|
0.061 |
| 6 |
H |
0.193 |
|
|
0.064 |
| 7 |
C |
-0.415 |
|
|
-0.087 |
| 8 |
H |
0.186 |
|
|
0.044 |
| 9 |
H |
0.197 |
|
|
0.071 |
| 10 |
C |
-0.358 |
|
|
-0.168 |
| 11 |
H |
0.197 |
|
|
-0.002 |
| 12 |
C |
-0.269 |
|
|
0.357 |
| 13 |
H |
0.183 |
|
|
0.129 |
| 14 |
H |
0.185 |
|
|
0.110 |
| 15 |
H |
0.190 |
|
|
0.143 |
| 16 |
C |
-0.535 |
|
|
-0.517 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.015 |
-0.132 |
0.076 |
0.153 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.005 |
-0.113 |
0.061 |
0.129 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.319 |
-0.219 |
-0.669 |
| y |
-0.219 |
-37.369 |
-0.165 |
| z |
-0.669 |
-0.165 |
-39.190 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.961 |
-0.219 |
-0.669 |
| y |
-0.219 |
0.885 |
-0.165 |
| z |
-0.669 |
-0.165 |
-1.846 |
|
| Polar |
|---|
| 3z2-r2 | -3.692 |
|---|
| x2-y2 | 0.050 |
|---|
| xy | -0.219 |
|---|
| xz | -0.669 |
|---|
| yz | -0.165 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.305 |
-0.369 |
-0.589 |
| y |
-0.369 |
8.286 |
-0.095 |
| z |
-0.589 |
-0.095 |
6.356 |
<r2> (average value of r
2) Å
2
| <r2> |
165.832 |
| (<r2>)1/2 |
12.878 |