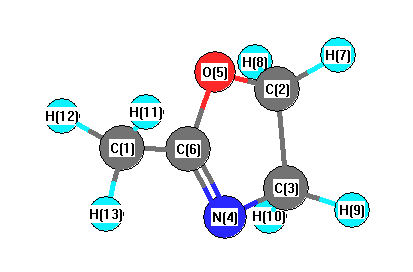
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3179 |
3068 |
3.94 |
|
|
|
| 2 |
A |
3097 |
2988 |
32.11 |
|
|
|
| 3 |
A |
3086 |
2978 |
30.45 |
|
|
|
| 4 |
A |
3071 |
2963 |
6.12 |
|
|
|
| 5 |
A |
1703 |
1643 |
91.05 |
|
|
|
| 6 |
A |
1572 |
1517 |
0.65 |
|
|
|
| 7 |
A |
1550 |
1495 |
1.47 |
|
|
|
| 8 |
A |
1538 |
1484 |
14.54 |
|
|
|
| 9 |
A |
1457 |
1406 |
17.37 |
|
|
|
| 10 |
A |
1363 |
1315 |
12.70 |
|
|
|
| 11 |
A |
1324 |
1277 |
2.16 |
|
|
|
| 12 |
A |
1229 |
1186 |
103.26 |
|
|
|
| 13 |
A |
1083 |
1045 |
2.41 |
|
|
|
| 14 |
A |
948 |
914 |
45.76 |
|
|
|
| 15 |
A |
917 |
884 |
2.54 |
|
|
|
| 16 |
A |
889 |
858 |
46.82 |
|
|
|
| 17 |
A |
861 |
831 |
21.51 |
|
|
|
| 18 |
A |
742 |
715 |
9.78 |
|
|
|
| 19 |
A |
649 |
626 |
2.99 |
|
|
|
| 20 |
A |
326 |
315 |
5.38 |
|
|
|
| 21 |
A |
3152 |
3041 |
37.66 |
|
|
|
| 22 |
A |
3123 |
3013 |
1.03 |
|
|
|
| 23 |
A |
3120 |
3011 |
7.14 |
|
|
|
| 24 |
A |
1543 |
1489 |
10.78 |
|
|
|
| 25 |
A |
1277 |
1232 |
0.13 |
|
|
|
| 26 |
A |
1201 |
1159 |
0.20 |
|
|
|
| 27 |
A |
1117 |
1077 |
4.10 |
|
|
|
| 28 |
A |
1093 |
1055 |
1.81 |
|
|
|
| 29 |
A |
833 |
804 |
0.01 |
|
|
|
| 30 |
A |
574 |
553 |
4.94 |
|
|
|
| 31 |
A |
234 |
226 |
13.12 |
|
|
|
| 32 |
A |
139 |
134 |
1.83 |
|
|
|
| 33 |
A |
123 |
119 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24054.4 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 23210.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.608 |
|
|
0.069 |
| 2 |
C |
-0.185 |
|
|
0.069 |
| 3 |
C |
-0.305 |
|
|
0.030 |
| 4 |
N |
-0.482 |
|
|
0.041 |
| 5 |
O |
-0.485 |
|
|
0.041 |
| 6 |
C |
0.523 |
|
|
0.166 |
| 7 |
H |
0.212 |
|
|
-0.652 |
| 8 |
H |
0.212 |
|
|
-0.441 |
| 9 |
H |
0.219 |
|
|
0.812 |
| 10 |
H |
0.219 |
|
|
0.207 |
| 11 |
H |
0.225 |
|
|
0.204 |
| 12 |
H |
0.225 |
|
|
0.207 |
| 13 |
H |
0.229 |
|
|
-0.752 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.671 |
-1.097 |
0.000 |
1.286 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-1.037 |
-0.841 |
0.001 |
1.335 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.168 |
2.188 |
0.000 |
| y |
2.188 |
-28.852 |
0.000 |
| z |
0.000 |
0.000 |
-35.217 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.133 |
2.188 |
0.000 |
| y |
2.188 |
9.341 |
0.000 |
| z |
0.000 |
0.000 |
-0.208 |
|
| Polar |
|---|
| 3z2-r2 | -0.415 |
|---|
| x2-y2 | -12.316 |
|---|
| xy | 2.188 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.675 |
-0.346 |
0.000 |
| y |
-0.346 |
6.670 |
-0.001 |
| z |
0.000 |
-0.001 |
5.022 |
<r2> (average value of r
2) Å
2
| <r2> |
152.121 |
| (<r2>)1/2 |
12.334 |