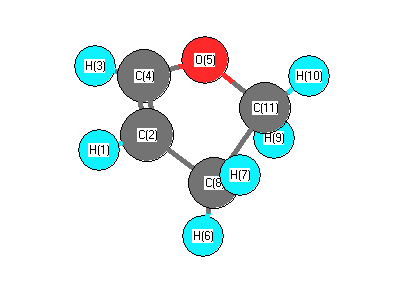
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3287 |
3172 |
5.02 |
|
|
|
| 2 |
A |
3258 |
3143 |
1.25 |
|
|
|
| 3 |
A |
3141 |
3031 |
36.13 |
|
|
|
| 4 |
A |
3091 |
2983 |
39.19 |
|
|
|
| 5 |
A |
3084 |
2976 |
10.52 |
|
|
|
| 6 |
A |
3056 |
2948 |
31.41 |
|
|
|
| 7 |
A |
1675 |
1616 |
36.71 |
|
|
|
| 8 |
A |
1570 |
1515 |
0.36 |
|
|
|
| 9 |
A |
1546 |
1492 |
0.33 |
|
|
|
| 10 |
A |
1397 |
1348 |
1.82 |
|
|
|
| 11 |
A |
1331 |
1285 |
10.09 |
|
|
|
| 12 |
A |
1306 |
1260 |
13.23 |
|
|
|
| 13 |
A |
1281 |
1236 |
0.67 |
|
|
|
| 14 |
A |
1206 |
1164 |
1.67 |
|
|
|
| 15 |
A |
1179 |
1137 |
20.79 |
|
|
|
| 16 |
A |
1081 |
1043 |
0.18 |
|
|
|
| 17 |
A |
1046 |
1009 |
78.55 |
|
|
|
| 18 |
A |
984 |
950 |
1.33 |
|
|
|
| 19 |
A |
946 |
913 |
2.55 |
|
|
|
| 20 |
A |
910 |
878 |
19.19 |
|
|
|
| 21 |
A |
892 |
861 |
30.44 |
|
|
|
| 22 |
A |
840 |
810 |
0.78 |
|
|
|
| 23 |
A |
836 |
807 |
1.29 |
|
|
|
| 24 |
A |
735 |
710 |
46.63 |
|
|
|
| 25 |
A |
703 |
679 |
3.30 |
|
|
|
| 26 |
A |
456 |
440 |
26.84 |
|
|
|
| 27 |
A |
69 |
67 |
1.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20454.5 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 19736.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.181 |
|
|
0.150 |
| 2 |
C |
-0.261 |
|
|
-0.355 |
| 3 |
H |
0.214 |
|
|
0.150 |
| 4 |
C |
0.114 |
|
|
0.032 |
| 5 |
O |
-0.484 |
|
|
-0.315 |
| 6 |
H |
0.207 |
|
|
-0.008 |
| 7 |
H |
0.207 |
|
|
-0.008 |
| 8 |
C |
-0.437 |
|
|
0.183 |
| 9 |
H |
0.210 |
|
|
0.062 |
| 10 |
H |
0.210 |
|
|
0.062 |
| 11 |
C |
-0.161 |
|
|
0.049 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.809 |
1.155 |
0.000 |
1.410 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.841 |
1.103 |
0.000 |
1.387 |
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.582 |
0.149 |
-0.001 |
| y |
0.149 |
6.644 |
-0.001 |
| z |
-0.001 |
-0.001 |
3.714 |
<r2> (average value of r
2) Å
2
| <r2> |
96.438 |
| (<r2>)1/2 |
9.820 |