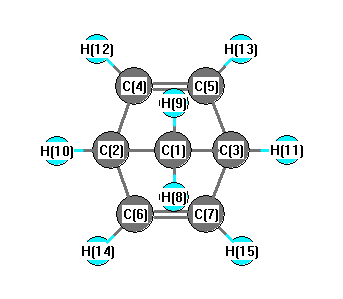
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3262 |
3147 |
6.51 |
|
|
|
| 2 |
A1 |
3166 |
3055 |
7.97 |
|
|
|
| 3 |
A1 |
3086 |
2978 |
32.60 |
|
|
|
| 4 |
A1 |
1649 |
1591 |
0.09 |
|
|
|
| 5 |
A1 |
1530 |
1477 |
0.80 |
|
|
|
| 6 |
A1 |
1264 |
1220 |
4.15 |
|
|
|
| 7 |
A1 |
1153 |
1113 |
0.25 |
|
|
|
| 8 |
A1 |
958 |
924 |
3.73 |
|
|
|
| 9 |
A1 |
882 |
851 |
13.85 |
|
|
|
| 10 |
A1 |
800 |
772 |
6.04 |
|
|
|
| 11 |
A1 |
766 |
739 |
52.02 |
|
|
|
| 12 |
A1 |
438 |
423 |
6.44 |
|
|
|
| 13 |
A2 |
3229 |
3116 |
0.00 |
|
|
|
| 14 |
A2 |
1315 |
1269 |
0.00 |
|
|
|
| 15 |
A2 |
1288 |
1243 |
0.00 |
|
|
|
| 16 |
A2 |
1167 |
1126 |
0.00 |
|
|
|
| 17 |
A2 |
956 |
922 |
0.00 |
|
|
|
| 18 |
A2 |
890 |
859 |
0.00 |
|
|
|
| 19 |
A2 |
741 |
715 |
0.00 |
|
|
|
| 20 |
A2 |
458 |
442 |
0.00 |
|
|
|
| 21 |
B1 |
3258 |
3143 |
20.41 |
|
|
|
| 22 |
B1 |
3152 |
3042 |
28.74 |
|
|
|
| 23 |
B1 |
1612 |
1556 |
3.28 |
|
|
|
| 24 |
B1 |
1245 |
1202 |
3.32 |
|
|
|
| 25 |
B1 |
1113 |
1074 |
0.14 |
|
|
|
| 26 |
B1 |
1047 |
1011 |
1.26 |
|
|
|
| 27 |
B1 |
877 |
846 |
3.35 |
|
|
|
| 28 |
B1 |
689 |
664 |
38.01 |
|
|
|
| 29 |
B1 |
508 |
490 |
7.28 |
|
|
|
| 30 |
B2 |
3231 |
3118 |
8.68 |
|
|
|
| 31 |
B2 |
3162 |
3051 |
40.62 |
|
|
|
| 32 |
B2 |
1358 |
1311 |
24.89 |
|
|
|
| 33 |
B2 |
1309 |
1263 |
0.75 |
|
|
|
| 34 |
B2 |
1183 |
1141 |
0.80 |
|
|
|
| 35 |
B2 |
989 |
955 |
2.02 |
|
|
|
| 36 |
B2 |
967 |
933 |
2.09 |
|
|
|
| 37 |
B2 |
890 |
859 |
0.49 |
|
|
|
| 38 |
B2 |
802 |
774 |
6.78 |
|
|
|
| 39 |
B2 |
554 |
534 |
1.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28471.4 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 27472.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.251 |
|
|
|
| 2 |
C |
-0.391 |
|
|
|
| 3 |
C |
-0.391 |
|
|
|
| 4 |
C |
-0.101 |
|
|
|
| 5 |
C |
-0.101 |
|
|
|
| 6 |
C |
-0.101 |
|
|
|
| 7 |
C |
-0.101 |
|
|
|
| 8 |
H |
0.193 |
|
|
|
| 9 |
H |
0.193 |
|
|
|
| 10 |
H |
0.168 |
|
|
|
| 11 |
H |
0.168 |
|
|
|
| 12 |
H |
0.179 |
|
|
|
| 13 |
H |
0.179 |
|
|
|
| 14 |
H |
0.179 |
|
|
|
| 15 |
H |
0.179 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.010 |
0.010 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.837 |
0.000 |
0.000 |
| y |
0.000 |
-38.784 |
0.000 |
| z |
0.000 |
0.000 |
-42.736 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.077 |
0.000 |
0.000 |
| y |
0.000 |
3.503 |
0.000 |
| z |
0.000 |
0.000 |
-2.425 |
|
| Polar |
|---|
| 3z2-r2 | -4.851 |
|---|
| x2-y2 | -3.053 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.425 |
0.000 |
0.000 |
| y |
0.000 |
10.136 |
0.000 |
| z |
0.000 |
0.000 |
6.940 |
<r2> (average value of r
2) Å
2
| <r2> |
153.926 |
| (<r2>)1/2 |
12.407 |