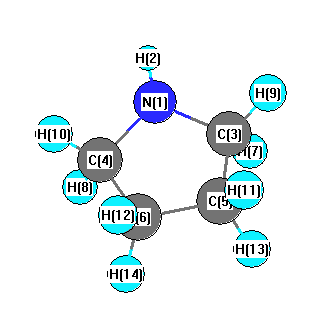
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3466 |
3344 |
0.99 |
|
|
|
| 2 |
A |
3141 |
3031 |
43.74 |
|
|
|
| 3 |
A |
3125 |
3015 |
17.11 |
|
|
|
| 4 |
A |
3091 |
2982 |
35.85 |
|
|
|
| 5 |
A |
3078 |
2970 |
3.49 |
|
|
|
| 6 |
A |
3077 |
2969 |
48.03 |
|
|
|
| 7 |
A |
3055 |
2948 |
46.17 |
|
|
|
| 8 |
A |
2996 |
2891 |
74.61 |
|
|
|
| 9 |
A |
2936 |
2833 |
97.58 |
|
|
|
| 10 |
A |
1595 |
1539 |
1.68 |
|
|
|
| 11 |
A |
1576 |
1521 |
4.20 |
|
|
|
| 12 |
A |
1552 |
1498 |
3.80 |
|
|
|
| 13 |
A |
1539 |
1485 |
4.50 |
|
|
|
| 14 |
A |
1463 |
1412 |
3.42 |
|
|
|
| 15 |
A |
1407 |
1357 |
0.28 |
|
|
|
| 16 |
A |
1376 |
1328 |
1.58 |
|
|
|
| 17 |
A |
1360 |
1313 |
3.07 |
|
|
|
| 18 |
A |
1328 |
1281 |
20.31 |
|
|
|
| 19 |
A |
1270 |
1226 |
2.91 |
|
|
|
| 20 |
A |
1267 |
1223 |
5.90 |
|
|
|
| 21 |
A |
1215 |
1173 |
2.89 |
|
|
|
| 22 |
A |
1198 |
1156 |
0.36 |
|
|
|
| 23 |
A |
1150 |
1110 |
13.88 |
|
|
|
| 24 |
A |
1092 |
1053 |
16.44 |
|
|
|
| 25 |
A |
1061 |
1024 |
2.76 |
|
|
|
| 26 |
A |
1005 |
970 |
3.39 |
|
|
|
| 27 |
A |
925 |
893 |
1.59 |
|
|
|
| 28 |
A |
918 |
886 |
0.84 |
|
|
|
| 29 |
A |
897 |
866 |
1.59 |
|
|
|
| 30 |
A |
884 |
853 |
2.81 |
|
|
|
| 31 |
A |
836 |
807 |
0.78 |
|
|
|
| 32 |
A |
756 |
730 |
31.89 |
|
|
|
| 33 |
A |
592 |
572 |
2.69 |
|
|
|
| 34 |
A |
497 |
479 |
115.48 |
|
|
|
| 35 |
A |
271 |
261 |
1.77 |
|
|
|
| 36 |
A |
47 |
45 |
2.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28520.7 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 27519.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.572 |
|
|
0.129 |
| 2 |
H |
0.266 |
|
|
0.080 |
| 3 |
C |
-0.202 |
|
|
-0.122 |
| 4 |
C |
-0.205 |
|
|
-0.159 |
| 5 |
C |
-0.400 |
|
|
-0.668 |
| 6 |
C |
-0.376 |
|
|
0.023 |
| 7 |
H |
0.168 |
|
|
0.036 |
| 8 |
H |
0.157 |
|
|
0.035 |
| 9 |
H |
0.193 |
|
|
0.049 |
| 10 |
H |
0.194 |
|
|
0.076 |
| 11 |
H |
0.194 |
|
|
0.051 |
| 12 |
H |
0.206 |
|
|
0.081 |
| 13 |
H |
0.193 |
|
|
0.057 |
| 14 |
H |
0.184 |
|
|
0.334 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.096 |
0.149 |
1.055 |
1.070 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.004 |
-0.141 |
0.991 |
1.001 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.335 |
-0.135 |
0.028 |
| y |
-0.135 |
-30.963 |
-1.939 |
| z |
0.028 |
-1.939 |
-33.336 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.815 |
-0.135 |
0.028 |
| y |
-0.135 |
1.372 |
-1.939 |
| z |
0.028 |
-1.939 |
-2.187 |
|
| Polar |
|---|
| 3z2-r2 | -4.374 |
|---|
| x2-y2 | -0.372 |
|---|
| xy | -0.135 |
|---|
| xz | 0.028 |
|---|
| yz | -1.939 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.187 |
-0.000 |
-0.000 |
| y |
-0.000 |
6.720 |
0.017 |
| z |
-0.000 |
0.017 |
5.911 |
<r2> (average value of r
2) Å
2
| <r2> |
110.839 |
| (<r2>)1/2 |
10.528 |