Jump to
S1C2
Energy calculated at B97D3/3-21G
| | hartrees |
|---|
| Energy at 0K | -1189.880190 |
| Energy at 298.15K | -1189.882083 |
| HF Energy | -1189.880190 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2401 |
2360 |
0.23 |
|
|
|
| 2 |
A |
771 |
758 |
0.06 |
|
|
|
| 3 |
A |
390 |
384 |
0.86 |
|
|
|
| 4 |
A |
275 |
271 |
40.21 |
|
|
|
| 5 |
A |
145 |
143 |
0.06 |
|
|
|
| 6 |
B |
2402 |
2361 |
52.51 |
|
|
|
| 7 |
B |
769 |
756 |
1.47 |
|
|
|
| 8 |
B |
365 |
359 |
37.27 |
|
|
|
| 9 |
B |
294 |
289 |
26.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3906.0 cm
-1
Scaled (by 0.983) Zero Point Vibrational Energy (zpe) 3839.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/3-21G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.954 |
| S2 |
0.000 |
1.827 |
-0.439 |
| S3 |
0.000 |
-1.827 |
-0.439 |
| H4 |
-1.372 |
1.869 |
-0.610 |
| H5 |
1.372 |
-1.869 |
-0.610 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.2976 | 2.2976 | 2.7976 | 2.7976 |
S2 | 2.2976 | | 3.6535 | 1.3837 | 3.9465 | S3 | 2.2976 | 3.6535 | | 3.9465 | 1.3837 | H4 | 2.7976 | 1.3837 | 3.9465 | | 4.6382 | H5 | 2.7976 | 3.9465 | 1.3837 | 4.6382 | |
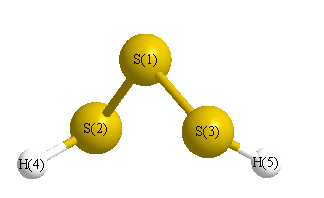 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
95.714 |
|
S1 |
S3 |
H5 |
95.714 |
| S2 |
S1 |
S3 |
105.326 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.045 |
|
|
|
| 2 |
S |
-0.093 |
|
|
|
| 3 |
S |
-0.093 |
|
|
|
| 4 |
H |
0.115 |
|
|
|
| 5 |
H |
0.115 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.632 |
0.632 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.472 |
-5.295 |
0.000 |
| y |
-5.295 |
-40.915 |
0.000 |
| z |
0.000 |
0.000 |
-42.383 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.177 |
-5.295 |
0.000 |
| y |
-5.295 |
-0.487 |
0.000 |
| z |
0.000 |
0.000 |
-2.690 |
|
| Polar |
|---|
| 3z2-r2 | -5.379 |
|---|
| x2-y2 | 2.442 |
|---|
| xy | -5.295 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.204 |
-0.832 |
0.000 |
| y |
-0.832 |
13.493 |
0.000 |
| z |
0.000 |
0.000 |
5.907 |
<r2> (average value of r
2) Å
2
| <r2> |
164.384 |
| (<r2>)1/2 |
12.821 |
Jump to
S1C1
Energy calculated at B97D3/3-21G
| | hartrees |
|---|
| Energy at 0K | -1189.879138 |
| Energy at 298.15K | -1189.880939 |
| HF Energy | -1189.879138 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2407 |
2366 |
67.34 |
|
|
|
| 2 |
A' |
762 |
749 |
2.37 |
|
|
|
| 3 |
A' |
389 |
383 |
0.71 |
|
|
|
| 4 |
A' |
268 |
263 |
30.08 |
|
|
|
| 5 |
A' |
146 |
144 |
0.07 |
|
|
|
| 6 |
A" |
2406 |
2365 |
0.00 |
|
|
|
| 7 |
A" |
757 |
744 |
0.38 |
|
|
|
| 8 |
A" |
366 |
360 |
43.62 |
|
|
|
| 9 |
A" |
236 |
232 |
10.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3867.9 cm
-1
Scaled (by 0.983) Zero Point Vibrational Energy (zpe) 3802.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.055 |
0.947 |
0.000 |
| S2 |
-0.055 |
-0.440 |
1.832 |
| S3 |
-0.055 |
-0.440 |
-1.832 |
| H4 |
1.323 |
-0.537 |
1.935 |
| H5 |
1.323 |
-0.537 |
-1.935 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.2977 | 2.2977 | 2.8007 | 2.8007 |
S2 | 2.2977 | | 3.6639 | 1.3852 | 4.0119 | S3 | 2.2977 | 3.6639 | | 4.0119 | 1.3852 | H4 | 2.8007 | 1.3852 | 4.0119 | | 3.8692 | H5 | 2.8007 | 4.0119 | 1.3852 | 3.8692 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
95.822 |
|
S1 |
S3 |
H5 |
95.822 |
| S2 |
S1 |
S3 |
105.746 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.045 |
|
|
|
| 2 |
S |
-0.082 |
|
|
|
| 3 |
S |
-0.082 |
|
|
|
| 4 |
H |
0.105 |
|
|
|
| 5 |
H |
0.105 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.563 |
-0.487 |
0.000 |
2.608 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.832 |
-1.656 |
0.000 |
| y |
-1.656 |
-42.596 |
0.000 |
| z |
0.000 |
0.000 |
-40.467 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.700 |
-1.656 |
0.000 |
| y |
-1.656 |
-2.947 |
0.000 |
| z |
0.000 |
0.000 |
0.247 |
|
| Polar |
|---|
| 3z2-r2 | 0.494 |
|---|
| x2-y2 | 3.764 |
|---|
| xy | -1.656 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.189 |
-0.374 |
0.000 |
| y |
-0.374 |
5.823 |
0.000 |
| z |
0.000 |
0.000 |
13.628 |
<r2> (average value of r
2) Å
2
| <r2> |
165.021 |
| (<r2>)1/2 |
12.846 |
