Jump to
S1C2
S1C3
Vibrational Frequencies calculated at MP4=FULL/3-21G
Geometric Data calculated at MP4=FULL/3-21G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at MP4=FULL/3-21G
Geometric Data calculated at MP4=FULL/3-21G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP4=FULL/3-21G
| | hartrees |
|---|
| Energy at 0K | -208.398872 |
| Energy at 298.15K | -208.408146 |
| HF Energy | -207.936113 |
| Nuclear repulsion energy | 132.781892 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3480 |
3407 |
|
|
|
|
| 2 |
A |
3379 |
3308 |
|
|
|
|
| 3 |
A |
3346 |
3275 |
|
|
|
|
| 4 |
A |
3094 |
3029 |
|
|
|
|
| 5 |
A |
3081 |
3016 |
|
|
|
|
| 6 |
A |
3000 |
2936 |
|
|
|
|
| 7 |
A |
2956 |
2893 |
|
|
|
|
| 8 |
A |
1735 |
1698 |
|
|
|
|
| 9 |
A |
1597 |
1563 |
|
|
|
|
| 10 |
A |
1564 |
1531 |
|
|
|
|
| 11 |
A |
1486 |
1455 |
|
|
|
|
| 12 |
A |
1431 |
1401 |
|
|
|
|
| 13 |
A |
1413 |
1383 |
|
|
|
|
| 14 |
A |
1351 |
1322 |
|
|
|
|
| 15 |
A |
1276 |
1249 |
|
|
|
|
| 16 |
A |
1196 |
1171 |
|
|
|
|
| 17 |
A |
1074 |
1052 |
|
|
|
|
| 18 |
A |
1018 |
997 |
|
|
|
|
| 19 |
A |
1005 |
984 |
|
|
|
|
| 20 |
A |
930 |
911 |
|
|
|
|
| 21 |
A |
851 |
833 |
|
|
|
|
| 22 |
A |
811 |
794 |
|
|
|
|
| 23 |
A |
699 |
684 |
|
|
|
|
| 24 |
A |
539 |
527 |
|
|
|
|
| 25 |
A |
337 |
330 |
|
|
|
|
| 26 |
A |
297 |
291 |
|
|
|
|
| 27 |
A |
205 |
201 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21575.9 cm
-1
Scaled (by 0.9788) Zero Point Vibrational Energy (zpe) 21118.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.332 |
-0.574 |
0.130 |
| C2 |
-0.635 |
0.705 |
-0.272 |
| C3 |
0.816 |
0.589 |
0.266 |
| O4 |
1.367 |
-0.698 |
-0.170 |
| H5 |
-1.630 |
-0.533 |
1.112 |
| H6 |
-2.147 |
-0.765 |
-0.463 |
| H7 |
-1.126 |
1.627 |
0.085 |
| H8 |
-0.597 |
0.720 |
-1.369 |
| H9 |
1.449 |
1.387 |
-0.143 |
| H10 |
0.798 |
0.685 |
1.368 |
| H11 |
0.559 |
-1.292 |
-0.101 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.5109 | 2.4470 | 2.7186 | 1.0277 | 1.0254 | 2.2108 | 2.1116 | 3.4138 | 2.7669 | 2.0366 |
C2 | 1.5109 | | 1.5523 | 2.4469 | 2.1071 | 2.1176 | 1.1033 | 1.0977 | 2.1964 | 2.1780 | 2.3336 | C3 | 2.4470 | 1.5523 | | 1.4661 | 2.8217 | 3.3388 | 2.2095 | 2.1651 | 1.0972 | 1.1066 | 1.9337 | O4 | 2.7186 | 2.4469 | 1.4661 | | 3.2639 | 3.5268 | 3.4180 | 2.7028 | 2.0868 | 2.1451 | 1.0048 | H5 | 1.0277 | 2.1071 | 2.8217 | 3.2639 | | 1.6739 | 2.4449 | 2.9654 | 3.8398 | 2.7285 | 2.6155 | H6 | 1.0254 | 2.1176 | 3.3388 | 3.5268 | 1.6739 | | 2.6579 | 2.3297 | 4.2029 | 3.7587 | 2.7810 | H7 | 2.2108 | 1.1033 | 2.2095 | 3.4180 | 2.4449 | 2.6579 | | 1.7929 | 2.5957 | 2.4970 | 3.3755 | H8 | 2.1116 | 1.0977 | 2.1651 | 2.7028 | 2.9654 | 2.3297 | 1.7929 | | 2.4766 | 3.0722 | 2.6446 | H9 | 3.4138 | 2.1964 | 1.0972 | 2.0868 | 3.8398 | 4.2029 | 2.5957 | 2.4766 | | 1.7892 | 2.8232 | H10 | 2.7669 | 2.1780 | 1.1066 | 2.1451 | 2.7285 | 3.7587 | 2.4970 | 3.0722 | 1.7892 | | 2.4743 | H11 | 2.0366 | 2.3336 | 1.9337 | 1.0048 | 2.6155 | 2.7810 | 3.3755 | 2.6446 | 2.8232 | 2.4743 | |
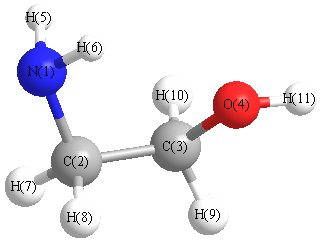 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
106.030 |
|
N1 |
C2 |
H7 |
114.592 |
| N1 |
C2 |
H8 |
107.016 |
|
C2 |
N1 |
H5 |
110.764 |
| C2 |
N1 |
H6 |
111.786 |
|
C2 |
C3 |
O4 |
108.289 |
| C2 |
C3 |
H9 |
110.820 |
|
C2 |
C3 |
H10 |
108.836 |
| C3 |
C2 |
H7 |
111.491 |
|
C3 |
C2 |
H8 |
108.351 |
| C3 |
O4 |
H11 |
101.351 |
|
O4 |
C3 |
H9 |
108.132 |
| O4 |
C3 |
H10 |
112.227 |
|
H5 |
N1 |
H6 |
109.241 |
| H7 |
C2 |
H8 |
109.096 |
|
H9 |
C3 |
H10 |
108.551 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
