Jump to
S1C2
S1C3
Energy calculated at mPW1PW91/3-21G
| | hartrees |
|---|
| Energy at 0K | -689.234191 |
| Energy at 298.15K | -689.233677 |
| HF Energy | -689.234191 |
| Nuclear repulsion energy | 61.003713 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at mPW1PW91/3-21G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.305 |
| C2 |
0.000 |
0.000 |
-1.273 |
| N3 |
0.000 |
0.000 |
-2.451 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.5787 | 3.7563 |
C2 | 2.5787 | | 1.1777 | N3 | 3.7563 | 1.1777 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.675 |
|
|
|
| 2 |
C |
-0.207 |
|
|
|
| 3 |
N |
-0.468 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
12.279 |
12.279 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.656 |
0.000 |
0.000 |
| y |
0.000 |
-23.656 |
0.000 |
| z |
0.000 |
0.000 |
-29.006 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.675 |
0.000 |
0.000 |
| y |
0.000 |
2.675 |
0.000 |
| z |
0.000 |
0.000 |
-5.350 |
|
| Polar |
|---|
| 3z2-r2 | -10.701 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.035 |
0.000 |
0.000 |
| y |
0.000 |
3.035 |
0.000 |
| z |
0.000 |
0.000 |
5.587 |
<r2> (average value of r
2) Å
2
| <r2> |
100.042 |
| (<r2>)1/2 |
10.002 |
Jump to
S1C1
S1C3
Energy calculated at mPW1PW91/3-21G
| | hartrees |
|---|
| Energy at 0K | -689.236412 |
| Energy at 298.15K | |
| HF Energy | -689.236412 |
| Nuclear repulsion energy | 64.294895 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at mPW1PW91/3-21G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.212 |
| C2 |
0.000 |
0.000 |
-2.410 |
| N3 |
0.000 |
0.000 |
-1.225 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.6217 | 2.4366 |
C2 | 3.6217 | | 1.1851 | N3 | 2.4366 | 1.1851 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.678 |
|
|
|
| 2 |
C |
-0.016 |
|
|
|
| 3 |
N |
-0.662 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
12.406 |
12.406 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.619 |
0.000 |
0.000 |
| y |
0.000 |
-23.619 |
0.000 |
| z |
0.000 |
0.000 |
-31.939 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.160 |
0.000 |
0.000 |
| y |
0.000 |
4.160 |
0.000 |
| z |
0.000 |
0.000 |
-8.320 |
|
| Polar |
|---|
| 3z2-r2 | -16.640 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.907 |
0.000 |
0.000 |
| y |
0.000 |
2.907 |
0.000 |
| z |
0.000 |
0.000 |
5.783 |
<r2> (average value of r
2) Å
2
| <r2> |
89.733 |
| (<r2>)1/2 |
9.473 |
Jump to
S1C1
S1C2
Energy calculated at mPW1PW91/3-21G
| | hartrees |
|---|
| Energy at 0K | -689.239441 |
| Energy at 298.15K | -689.239256 |
| HF Energy | -689.239441 |
| Nuclear repulsion energy | 67.885806 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at mPW1PW91/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.055 |
0.000 |
| C2 |
0.631 |
-1.647 |
0.000 |
| N3 |
-0.541 |
-1.452 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.7754 | 2.5650 |
C2 | 2.7754 | | 1.1879 | N3 | 2.5650 | 1.1879 | |
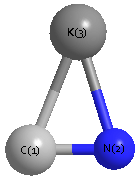 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
67.391 |
|
K1 |
N3 |
C2 |
87.298 |
| C2 |
K1 |
N3 |
25.311 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.626 |
|
|
|
| 2 |
C |
-0.082 |
|
|
|
| 3 |
N |
-0.544 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.434 |
10.717 |
0.000 |
10.726 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.167 |
1.044 |
0.000 |
| y |
1.044 |
-25.924 |
0.000 |
| z |
0.000 |
0.000 |
-23.911 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.249 |
1.044 |
0.000 |
| y |
1.044 |
0.115 |
0.000 |
| z |
0.000 |
0.000 |
3.135 |
|
| Polar |
|---|
| 3z2-r2 | 6.269 |
|---|
| x2-y2 | -2.243 |
|---|
| xy | 1.044 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.073 |
-0.132 |
0.000 |
| y |
-0.132 |
3.448 |
-0.000 |
| z |
0.000 |
-0.000 |
3.050 |
<r2> (average value of r
2) Å
2
| <r2> |
72.877 |
| (<r2>)1/2 |
8.537 |
