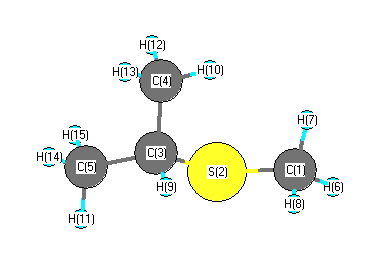
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3098 |
3081 |
6.32 |
|
|
|
| 2 |
A |
3088 |
3071 |
15.92 |
|
|
|
| 3 |
A |
3056 |
3039 |
35.82 |
|
|
|
| 4 |
A |
3051 |
3034 |
15.12 |
|
|
|
| 5 |
A |
3032 |
3015 |
27.87 |
|
|
|
| 6 |
A |
3023 |
3007 |
14.50 |
|
|
|
| 7 |
A |
3004 |
2987 |
3.86 |
|
|
|
| 8 |
A |
3002 |
2985 |
21.32 |
|
|
|
| 9 |
A |
2973 |
2957 |
24.99 |
|
|
|
| 10 |
A |
2963 |
2947 |
22.07 |
|
|
|
| 11 |
A |
1536 |
1528 |
7.80 |
|
|
|
| 12 |
A |
1529 |
1520 |
6.67 |
|
|
|
| 13 |
A |
1517 |
1509 |
3.47 |
|
|
|
| 14 |
A |
1516 |
1508 |
5.91 |
|
|
|
| 15 |
A |
1507 |
1498 |
8.88 |
|
|
|
| 16 |
A |
1501 |
1493 |
9.44 |
|
|
|
| 17 |
A |
1431 |
1423 |
5.09 |
|
|
|
| 18 |
A |
1415 |
1407 |
9.34 |
|
|
|
| 19 |
A |
1345 |
1338 |
4.21 |
|
|
|
| 20 |
A |
1340 |
1333 |
1.60 |
|
|
|
| 21 |
A |
1252 |
1246 |
34.96 |
|
|
|
| 22 |
A |
1176 |
1169 |
24.80 |
|
|
|
| 23 |
A |
1101 |
1095 |
1.67 |
|
|
|
| 24 |
A |
1066 |
1060 |
4.55 |
|
|
|
| 25 |
A |
967 |
961 |
8.38 |
|
|
|
| 26 |
A |
957 |
952 |
5.52 |
|
|
|
| 27 |
A |
944 |
939 |
5.21 |
|
|
|
| 28 |
A |
941 |
935 |
0.80 |
|
|
|
| 29 |
A |
845 |
841 |
2.34 |
|
|
|
| 30 |
A |
616 |
612 |
2.86 |
|
|
|
| 31 |
A |
567 |
564 |
4.57 |
|
|
|
| 32 |
A |
407 |
405 |
1.37 |
|
|
|
| 33 |
A |
323 |
321 |
1.30 |
|
|
|
| 34 |
A |
311 |
309 |
1.02 |
|
|
|
| 35 |
A |
247 |
246 |
0.07 |
|
|
|
| 36 |
A |
227 |
226 |
0.02 |
|
|
|
| 37 |
A |
183 |
182 |
0.64 |
|
|
|
| 38 |
A |
141 |
140 |
0.46 |
|
|
|
| 39 |
A |
65 |
64 |
1.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28629.7 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 28472.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.746 |
|
|
|
| 2 |
S |
0.197 |
|
|
|
| 3 |
C |
-0.409 |
|
|
|
| 4 |
C |
-0.500 |
|
|
|
| 5 |
C |
-0.506 |
|
|
|
| 6 |
H |
0.218 |
|
|
|
| 7 |
H |
0.210 |
|
|
|
| 8 |
H |
0.207 |
|
|
|
| 9 |
H |
0.206 |
|
|
|
| 10 |
H |
0.183 |
|
|
|
| 11 |
H |
0.195 |
|
|
|
| 12 |
H |
0.193 |
|
|
|
| 13 |
H |
0.178 |
|
|
|
| 14 |
H |
0.183 |
|
|
|
| 15 |
H |
0.192 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.440 |
1.502 |
1.091 |
1.908 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.123 |
1.601 |
0.733 |
| y |
1.601 |
-42.174 |
-0.103 |
| z |
0.733 |
-0.103 |
-41.616 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.773 |
1.601 |
0.733 |
| y |
1.601 |
-2.804 |
-0.103 |
| z |
0.733 |
-0.103 |
-1.968 |
|
| Polar |
|---|
| 3z2-r2 | -3.937 |
|---|
| x2-y2 | 5.051 |
|---|
| xy | 1.601 |
|---|
| xz | 0.733 |
|---|
| yz | -0.103 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
193.215 |
| (<r2>)1/2 |
13.900 |